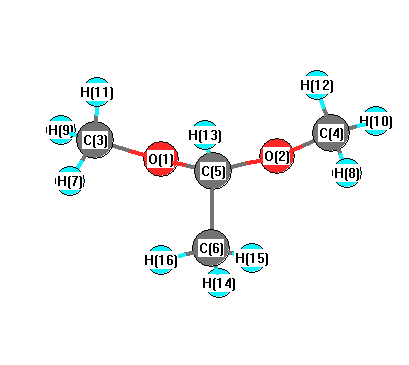
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3257 |
2952 |
52.66 |
|
|
|
| 2 |
A' |
3251 |
2946 |
20.85 |
|
|
|
| 3 |
A' |
3183 |
2885 |
122.24 |
|
|
|
| 4 |
A' |
3173 |
2876 |
16.96 |
|
|
|
| 5 |
A' |
3139 |
2845 |
85.79 |
|
|
|
| 6 |
A' |
3091 |
2801 |
35.52 |
|
|
|
| 7 |
A' |
1632 |
1479 |
3.76 |
|
|
|
| 8 |
A' |
1627 |
1474 |
3.08 |
|
|
|
| 9 |
A' |
1614 |
1463 |
12.85 |
|
|
|
| 10 |
A' |
1599 |
1450 |
3.20 |
|
|
|
| 11 |
A' |
1547 |
1402 |
58.48 |
|
|
|
| 12 |
A' |
1503 |
1362 |
11.23 |
|
|
|
| 13 |
A' |
1387 |
1257 |
36.09 |
|
|
|
| 14 |
A' |
1293 |
1172 |
51.54 |
|
|
|
| 15 |
A' |
1281 |
1161 |
11.10 |
|
|
|
| 16 |
A' |
1224 |
1110 |
3.32 |
|
|
|
| 17 |
A' |
1166 |
1057 |
54.84 |
|
|
|
| 18 |
A' |
910 |
825 |
7.25 |
|
|
|
| 19 |
A' |
576 |
522 |
7.60 |
|
|
|
| 20 |
A' |
481 |
436 |
13.43 |
|
|
|
| 21 |
A' |
276 |
250 |
3.28 |
|
|
|
| 22 |
A' |
179 |
162 |
1.91 |
|
|
|
| 23 |
A' |
56 |
50 |
4.38 |
|
|
|
| 24 |
A" |
3257 |
2952 |
36.15 |
|
|
|
| 25 |
A" |
3233 |
2931 |
58.75 |
|
|
|
| 26 |
A" |
3179 |
2882 |
14.15 |
|
|
|
| 27 |
A" |
3135 |
2842 |
53.43 |
|
|
|
| 28 |
A" |
1634 |
1481 |
1.01 |
|
|
|
| 29 |
A" |
1623 |
1471 |
6.53 |
|
|
|
| 30 |
A" |
1617 |
1466 |
6.99 |
|
|
|
| 31 |
A" |
1601 |
1451 |
1.20 |
|
|
|
| 32 |
A" |
1543 |
1399 |
64.71 |
|
|
|
| 33 |
A" |
1343 |
1217 |
227.90 |
|
|
|
| 34 |
A" |
1285 |
1165 |
2.84 |
|
|
|
| 35 |
A" |
1271 |
1152 |
268.99 |
|
|
|
| 36 |
A" |
1187 |
1076 |
54.93 |
|
|
|
| 37 |
A" |
1006 |
912 |
5.78 |
|
|
|
| 38 |
A" |
497 |
450 |
1.17 |
|
|
|
| 39 |
A" |
377 |
342 |
1.17 |
|
|
|
| 40 |
A" |
251 |
228 |
0.02 |
|
|
|
| 41 |
A" |
204 |
185 |
0.16 |
|
|
|
| 42 |
A" |
34 |
31 |
0.30 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32860.8 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 29785.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.386 |
|
|
|
| 2 |
O |
-0.386 |
|
|
|
| 3 |
C |
0.017 |
|
|
|
| 4 |
C |
0.017 |
|
|
|
| 5 |
C |
0.449 |
|
|
|
| 6 |
C |
-0.191 |
|
|
|
| 7 |
H |
0.046 |
|
|
|
| 8 |
H |
0.046 |
|
|
|
| 9 |
H |
0.078 |
|
|
|
| 10 |
H |
0.078 |
|
|
|
| 11 |
H |
0.047 |
|
|
|
| 12 |
H |
0.047 |
|
|
|
| 13 |
H |
-0.022 |
|
|
|
| 14 |
H |
0.069 |
|
|
|
| 15 |
H |
0.044 |
|
|
|
| 16 |
H |
0.044 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.109 |
2.301 |
0.000 |
2.555 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.750 |
-1.546 |
0.000 |
| y |
-1.546 |
-39.520 |
0.000 |
| z |
0.000 |
0.000 |
-32.629 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.675 |
-1.546 |
0.000 |
| y |
-1.546 |
-3.831 |
0.000 |
| z |
0.000 |
0.000 |
6.506 |
|
| Polar |
|---|
| 3z2-r2 | 13.012 |
|---|
| x2-y2 | 0.770 |
|---|
| xy | -1.546 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.767 |
-0.190 |
0.000 |
| y |
-0.190 |
7.235 |
0.000 |
| z |
0.000 |
0.000 |
9.388 |
<r2> (average value of r
2) Å
2
| <r2> |
197.295 |
| (<r2>)1/2 |
14.046 |