Jump to
S1C2
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -267.774641 |
| Energy at 298.15K | -267.781681 |
| HF Energy | -267.774641 |
| Nuclear repulsion energy | 224.905252 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3433 |
3112 |
1.64 |
|
|
|
| 2 |
A |
3404 |
3085 |
1.50 |
|
|
|
| 3 |
A |
3391 |
3074 |
7.22 |
|
|
|
| 4 |
A |
3255 |
2951 |
18.97 |
|
|
|
| 5 |
A |
3218 |
2917 |
21.49 |
|
|
|
| 6 |
A |
3169 |
2873 |
32.67 |
|
|
|
| 7 |
A |
1785 |
1618 |
20.29 |
|
|
|
| 8 |
A |
1693 |
1534 |
32.49 |
|
|
|
| 9 |
A |
1608 |
1458 |
7.30 |
|
|
|
| 10 |
A |
1592 |
1443 |
6.28 |
|
|
|
| 11 |
A |
1549 |
1404 |
6.56 |
|
|
|
| 12 |
A |
1527 |
1384 |
1.89 |
|
|
|
| 13 |
A |
1368 |
1240 |
8.54 |
|
|
|
| 14 |
A |
1325 |
1201 |
14.44 |
|
|
|
| 15 |
A |
1264 |
1146 |
21.70 |
|
|
|
| 16 |
A |
1204 |
1091 |
17.41 |
|
|
|
| 17 |
A |
1164 |
1055 |
0.83 |
|
|
|
| 18 |
A |
1096 |
993 |
8.80 |
|
|
|
| 19 |
A |
1072 |
971 |
8.69 |
|
|
|
| 20 |
A |
1019 |
923 |
28.46 |
|
|
|
| 21 |
A |
1004 |
910 |
0.20 |
|
|
|
| 22 |
A |
971 |
880 |
4.33 |
|
|
|
| 23 |
A |
917 |
831 |
15.25 |
|
|
|
| 24 |
A |
850 |
770 |
76.24 |
|
|
|
| 25 |
A |
701 |
636 |
2.11 |
|
|
|
| 26 |
A |
696 |
631 |
1.13 |
|
|
|
| 27 |
A |
659 |
598 |
9.44 |
|
|
|
| 28 |
A |
361 |
327 |
2.41 |
|
|
|
| 29 |
A |
268 |
243 |
7.06 |
|
|
|
| 30 |
A |
165 |
149 |
0.21 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22864.6 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 20724.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-0.084 |
-1.033 |
-0.003 |
| C2 |
0.629 |
0.106 |
-0.005 |
| C3 |
-1.391 |
-0.717 |
0.002 |
| C4 |
-0.207 |
1.146 |
-0.002 |
| C5 |
-1.535 |
0.604 |
0.003 |
| C6 |
2.113 |
-0.026 |
0.003 |
| H7 |
-2.078 |
-1.530 |
0.002 |
| H8 |
0.074 |
2.176 |
-0.003 |
| H9 |
-2.459 |
1.137 |
0.005 |
| H10 |
2.388 |
-1.059 |
-0.155 |
| H11 |
2.561 |
0.571 |
-0.781 |
| H12 |
2.532 |
0.295 |
0.949 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| O1 | | 1.3446 | 1.3440 | 2.1831 | 2.1872 | 2.4178 | 2.0549 | 3.2132 | 3.2170 | 2.4772 | 3.1899 | 3.0848 |
C2 | 1.3446 | | 2.1811 | 1.3341 | 2.2201 | 1.4905 | 3.1634 | 2.1425 | 3.2550 | 2.1153 | 2.1331 | 2.1369 | C3 | 1.3440 | 2.1811 | | 2.2077 | 1.3288 | 3.5717 | 1.0647 | 3.2427 | 2.1401 | 3.7973 | 4.2291 | 4.1605 | C4 | 2.1831 | 1.3341 | 2.2077 | | 1.4345 | 2.5997 | 3.2658 | 1.0672 | 2.2517 | 3.4086 | 2.9320 | 3.0217 | C5 | 2.1872 | 2.2201 | 1.3288 | 1.4345 | | 3.7022 | 2.2019 | 2.2494 | 1.0668 | 4.2634 | 4.1699 | 4.1869 | C6 | 2.4178 | 1.4905 | 3.5717 | 2.5997 | 3.7022 | | 4.4535 | 3.0013 | 4.7177 | 1.0802 | 1.0820 | 1.0829 | H7 | 2.0549 | 3.1634 | 1.0647 | 3.2658 | 2.2019 | 4.4535 | | 4.2856 | 2.6944 | 4.4939 | 5.1525 | 5.0481 | H8 | 3.2132 | 2.1425 | 3.2427 | 1.0672 | 2.2494 | 3.0013 | 4.2856 | | 2.7371 | 3.9799 | 3.0600 | 3.2383 | H9 | 3.2170 | 3.2550 | 2.1401 | 2.2517 | 1.0668 | 4.7177 | 2.6944 | 2.7371 | | 5.3232 | 5.1117 | 5.1486 | H10 | 2.4772 | 2.1153 | 3.7973 | 3.4086 | 4.2634 | 1.0802 | 4.4939 | 3.9799 | 5.3232 | | 1.7541 | 1.7527 | H11 | 3.1899 | 2.1331 | 4.2291 | 2.9320 | 4.1699 | 1.0820 | 5.1525 | 3.0600 | 5.1117 | 1.7541 | | 1.7516 | H12 | 3.0848 | 2.1369 | 4.1605 | 3.0217 | 4.1869 | 1.0829 | 5.0481 | 3.2383 | 5.1486 | 1.7527 | 1.7516 | |
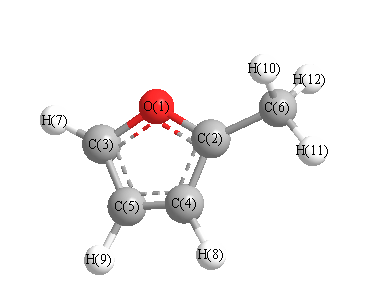 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C4 |
109.172 |
|
O1 |
C2 |
C6 |
116.939 |
| O1 |
C3 |
C5 |
109.832 |
|
O1 |
C3 |
H7 |
116.626 |
| C2 |
O1 |
C3 |
108.431 |
|
C2 |
C4 |
C5 |
106.567 |
| C2 |
C4 |
H8 |
125.953 |
|
C2 |
C6 |
H10 |
109.712 |
| C2 |
C6 |
H11 |
111.036 |
|
C2 |
C6 |
H12 |
111.295 |
| C3 |
C5 |
C6 |
73.981 |
|
C3 |
C5 |
H9 |
126.246 |
| C4 |
C2 |
C6 |
133.886 |
|
C4 |
C5 |
H9 |
127.757 |
| C5 |
C3 |
H7 |
133.541 |
|
C5 |
C4 |
H8 |
127.481 |
| H10 |
C6 |
H11 |
108.439 |
|
H10 |
C6 |
H12 |
108.248 |
| H11 |
C6 |
H12 |
108.017 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.376 |
|
|
|
| 2 |
C |
0.253 |
|
|
|
| 3 |
C |
0.072 |
|
|
|
| 4 |
C |
-0.185 |
|
|
|
| 5 |
C |
-0.117 |
|
|
|
| 6 |
C |
-0.181 |
|
|
|
| 7 |
H |
0.107 |
|
|
|
| 8 |
H |
0.097 |
|
|
|
| 9 |
H |
0.102 |
|
|
|
| 10 |
H |
0.077 |
|
|
|
| 11 |
H |
0.074 |
|
|
|
| 12 |
H |
0.077 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.272 |
0.547 |
-0.000 |
0.611 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-30.662 |
1.058 |
0.000 |
| y |
1.058 |
-34.270 |
-0.000 |
| z |
0.000 |
-0.000 |
-38.406 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.677 |
1.058 |
0.000 |
| y |
1.058 |
0.264 |
-0.000 |
| z |
0.000 |
-0.000 |
-5.940 |
|
| Polar |
|---|
| 3z2-r2 | -11.881 |
|---|
| x2-y2 | 3.608 |
|---|
| xy | 1.058 |
|---|
| xz | 0.000 |
|---|
| yz | -0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.034 |
-0.176 |
0.000 |
| y |
-0.176 |
8.705 |
0.000 |
| z |
0.000 |
0.000 |
5.796 |
<r2> (average value of r
2) Å
2
| <r2> |
138.897 |
| (<r2>)1/2 |
11.785 |
Jump to
S1C1
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -267.774641 |
| Energy at 298.15K | -267.781683 |
| HF Energy | -267.774641 |
| Nuclear repulsion energy | 224.911405 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3433 |
3112 |
1.65 |
|
|
|
| 2 |
A' |
3404 |
3085 |
1.51 |
|
|
|
| 3 |
A' |
3391 |
3074 |
7.21 |
|
|
|
| 4 |
A' |
3255 |
2951 |
18.97 |
|
|
|
| 5 |
A' |
3169 |
2873 |
32.66 |
|
|
|
| 6 |
A' |
1786 |
1619 |
20.30 |
|
|
|
| 7 |
A' |
1693 |
1534 |
32.50 |
|
|
|
| 8 |
A' |
1608 |
1458 |
7.30 |
|
|
|
| 9 |
A' |
1549 |
1404 |
6.53 |
|
|
|
| 10 |
A' |
1527 |
1384 |
1.88 |
|
|
|
| 11 |
A' |
1368 |
1240 |
8.51 |
|
|
|
| 12 |
A' |
1325 |
1201 |
14.46 |
|
|
|
| 13 |
A' |
1264 |
1146 |
21.81 |
|
|
|
| 14 |
A' |
1204 |
1091 |
17.36 |
|
|
|
| 15 |
A' |
1096 |
993 |
8.74 |
|
|
|
| 16 |
A' |
1072 |
971 |
8.72 |
|
|
|
| 17 |
A' |
1019 |
923 |
28.48 |
|
|
|
| 18 |
A' |
971 |
881 |
4.32 |
|
|
|
| 19 |
A' |
696 |
631 |
1.13 |
|
|
|
| 20 |
A' |
361 |
327 |
2.41 |
|
|
|
| 21 |
A" |
3218 |
2917 |
21.48 |
|
|
|
| 22 |
A" |
1592 |
1443 |
6.28 |
|
|
|
| 23 |
A" |
1165 |
1056 |
0.83 |
|
|
|
| 24 |
A" |
1004 |
910 |
0.20 |
|
|
|
| 25 |
A" |
917 |
832 |
15.23 |
|
|
|
| 26 |
A" |
850 |
770 |
76.27 |
|
|
|
| 27 |
A" |
701 |
636 |
2.10 |
|
|
|
| 28 |
A" |
659 |
598 |
9.44 |
|
|
|
| 29 |
A" |
268 |
243 |
7.05 |
|
|
|
| 30 |
A" |
165 |
150 |
0.21 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22865.2 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 20725.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
1.009 |
-0.253 |
0.000 |
| C2 |
0.000 |
0.639 |
0.000 |
| C3 |
0.477 |
-1.489 |
0.000 |
| C4 |
-1.164 |
-0.012 |
0.000 |
| C5 |
-0.849 |
-1.412 |
0.000 |
| C6 |
0.373 |
2.074 |
0.000 |
| H7 |
1.165 |
-2.301 |
0.000 |
| H8 |
-2.131 |
0.438 |
0.000 |
| H9 |
-1.528 |
-2.235 |
0.000 |
| H10 |
-0.518 |
2.687 |
0.000 |
| H11 |
0.961 |
2.320 |
0.876 |
| H12 |
0.961 |
2.320 |
-0.876 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| O1 | | 1.3464 | 1.3454 | 2.1857 | 2.1899 | 2.4128 | 2.0542 | 3.2151 | 3.2194 | 3.3129 | 2.7187 | 2.7187 |
C2 | 1.3464 | | 2.1803 | 1.3333 | 2.2196 | 1.4833 | 3.1624 | 2.1406 | 3.2548 | 2.1131 | 2.1255 | 2.1255 | C3 | 1.3454 | 2.1803 | | 2.2073 | 1.3286 | 3.5648 | 1.0647 | 3.2429 | 2.1396 | 4.2930 | 3.9383 | 3.9383 | C4 | 2.1857 | 1.3333 | 2.2073 | | 1.4347 | 2.5912 | 3.2655 | 1.0672 | 2.2525 | 2.7758 | 3.2742 | 3.2742 | C5 | 2.1899 | 2.2196 | 1.3286 | 1.4347 | | 3.6944 | 2.2021 | 2.2508 | 1.0669 | 4.1128 | 4.2395 | 4.2395 | C6 | 2.4128 | 1.4833 | 3.5648 | 2.5912 | 3.6944 | | 4.4469 | 2.9911 | 4.7101 | 1.0809 | 1.0832 | 1.0832 | H7 | 2.0542 | 3.1624 | 1.0647 | 3.2655 | 2.2021 | 4.4469 | | 4.2861 | 2.6943 | 5.2648 | 4.7082 | 4.7082 | H8 | 3.2151 | 2.1406 | 3.2429 | 1.0672 | 2.2508 | 2.9911 | 4.2861 | | 2.7404 | 2.7682 | 3.7241 | 3.7241 | H9 | 3.2194 | 3.2548 | 2.1396 | 2.2525 | 1.0669 | 4.7101 | 2.6943 | 2.7404 | | 5.0251 | 5.2643 | 5.2643 | H10 | 3.3129 | 2.1131 | 4.2930 | 2.7758 | 4.1128 | 1.0809 | 5.2648 | 2.7682 | 5.0251 | | 1.7571 | 1.7571 | H11 | 2.7187 | 2.1255 | 3.9383 | 3.2742 | 4.2395 | 1.0832 | 4.7082 | 3.7241 | 5.2643 | 1.7571 | | 1.7517 | H12 | 2.7187 | 2.1255 | 3.9383 | 3.2742 | 4.2395 | 1.0832 | 4.7082 | 3.7241 | 5.2643 | 1.7571 | 1.7517 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C4 |
109.307 |
|
O1 |
C2 |
C6 |
116.924 |
| O1 |
C3 |
C5 |
109.965 |
|
O1 |
C3 |
H7 |
116.449 |
| C2 |
O1 |
C3 |
108.194 |
|
C2 |
C4 |
C5 |
106.563 |
| C2 |
C4 |
H8 |
125.821 |
|
C2 |
C6 |
H10 |
109.999 |
| C2 |
C6 |
H11 |
110.858 |
|
C2 |
C6 |
H12 |
110.858 |
| C3 |
C5 |
C6 |
74.000 |
|
C3 |
C5 |
H9 |
126.204 |
| C4 |
C2 |
C6 |
133.769 |
|
C4 |
C5 |
H9 |
127.825 |
| C5 |
C3 |
H7 |
133.586 |
|
C5 |
C4 |
H8 |
127.616 |
| H10 |
C6 |
H11 |
108.568 |
|
H10 |
C6 |
H12 |
108.568 |
| H11 |
C6 |
H12 |
107.911 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.376 |
|
|
|
| 2 |
C |
0.253 |
|
|
|
| 3 |
C |
0.072 |
|
|
|
| 4 |
C |
-0.185 |
|
|
|
| 5 |
C |
-0.117 |
|
|
|
| 6 |
C |
-0.181 |
|
|
|
| 7 |
H |
0.107 |
|
|
|
| 8 |
H |
0.097 |
|
|
|
| 9 |
H |
0.102 |
|
|
|
| 10 |
H |
0.074 |
|
|
|
| 11 |
H |
0.077 |
|
|
|
| 12 |
H |
0.077 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.495 |
0.357 |
0.000 |
0.611 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-34.514 |
-0.425 |
0.000 |
| y |
-0.425 |
-30.419 |
0.000 |
| z |
0.000 |
0.000 |
-38.405 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.101 |
-0.425 |
0.000 |
| y |
-0.425 |
6.041 |
0.000 |
| z |
0.000 |
0.000 |
-5.939 |
|
| Polar |
|---|
| 3z2-r2 | -11.879 |
|---|
| x2-y2 | -4.095 |
|---|
| xy | -0.425 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.797 |
0.380 |
0.000 |
| y |
0.380 |
9.942 |
0.000 |
| z |
0.000 |
0.000 |
5.796 |
<r2> (average value of r
2) Å
2
| <r2> |
138.889 |
| (<r2>)1/2 |
11.785 |
