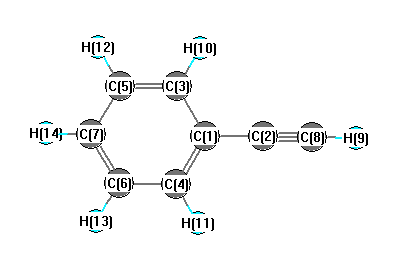
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3617 |
3279 |
94.86 |
|
|
|
| 2 |
A1 |
3356 |
3042 |
5.05 |
|
|
|
| 3 |
A1 |
3342 |
3029 |
20.35 |
|
|
|
| 4 |
A1 |
3320 |
3009 |
0.20 |
|
|
|
| 5 |
A1 |
2379 |
2156 |
10.47 |
|
|
|
| 6 |
A1 |
1784 |
1617 |
2.14 |
|
|
|
| 7 |
A1 |
1650 |
1496 |
20.85 |
|
|
|
| 8 |
A1 |
1294 |
1173 |
1.67 |
|
|
|
| 9 |
A1 |
1287 |
1167 |
0.00 |
|
|
|
| 10 |
A1 |
1119 |
1014 |
2.79 |
|
|
|
| 11 |
A1 |
1084 |
983 |
0.06 |
|
|
|
| 12 |
A1 |
819 |
743 |
3.69 |
|
|
|
| 13 |
A1 |
500 |
453 |
0.21 |
|
|
|
| 14 |
A2 |
1107 |
1003 |
0.00 |
|
|
|
| 15 |
A2 |
953 |
864 |
0.00 |
|
|
|
| 16 |
A2 |
451 |
409 |
0.00 |
|
|
|
| 17 |
B1 |
1120 |
1015 |
0.05 |
|
|
|
| 18 |
B1 |
1050 |
952 |
4.63 |
|
|
|
| 19 |
B1 |
857 |
777 |
55.70 |
|
|
|
| 20 |
B1 |
792 |
718 |
56.37 |
|
|
|
| 21 |
B1 |
767 |
695 |
39.37 |
|
|
|
| 22 |
B1 |
609 |
552 |
2.22 |
|
|
|
| 23 |
B1 |
410 |
372 |
6.20 |
|
|
|
| 24 |
B1 |
157 |
142 |
2.46 |
|
|
|
| 25 |
B2 |
3351 |
3038 |
20.65 |
|
|
|
| 26 |
B2 |
3331 |
3019 |
5.57 |
|
|
|
| 27 |
B2 |
1753 |
1589 |
1.39 |
|
|
|
| 28 |
B2 |
1594 |
1445 |
6.91 |
|
|
|
| 29 |
B2 |
1461 |
1324 |
0.25 |
|
|
|
| 30 |
B2 |
1319 |
1196 |
1.02 |
|
|
|
| 31 |
B2 |
1194 |
1083 |
5.12 |
|
|
|
| 32 |
B2 |
1166 |
1057 |
0.47 |
|
|
|
| 33 |
B2 |
829 |
752 |
42.82 |
|
|
|
| 34 |
B2 |
681 |
617 |
0.73 |
|
|
|
| 35 |
B2 |
587 |
532 |
8.88 |
|
|
|
| 36 |
B2 |
180 |
163 |
1.69 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25635.2 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 23235.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.207 |
|
|
|
| 2 |
C |
-0.010 |
|
|
|
| 3 |
C |
-0.106 |
|
|
|
| 4 |
C |
-0.106 |
|
|
|
| 5 |
C |
-0.160 |
|
|
|
| 6 |
C |
-0.160 |
|
|
|
| 7 |
C |
-0.023 |
|
|
|
| 8 |
C |
0.096 |
|
|
|
| 9 |
H |
0.192 |
|
|
|
| 10 |
H |
0.100 |
|
|
|
| 11 |
H |
0.100 |
|
|
|
| 12 |
H |
0.094 |
|
|
|
| 13 |
H |
0.094 |
|
|
|
| 14 |
H |
0.097 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.774 |
0.774 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-51.863 |
0.000 |
0.000 |
| y |
0.000 |
-42.590 |
0.000 |
| z |
0.000 |
0.000 |
-38.584 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-11.276 |
0.000 |
0.000 |
| y |
0.000 |
2.634 |
0.000 |
| z |
0.000 |
0.000 |
8.642 |
|
| Polar |
|---|
| 3z2-r2 | 17.284 |
|---|
| x2-y2 | -9.273 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.197 |
0.000 |
0.000 |
| y |
0.000 |
12.272 |
0.000 |
| z |
0.000 |
0.000 |
18.926 |
<r2> (average value of r
2) Å
2
| <r2> |
261.710 |
| (<r2>)1/2 |
16.177 |