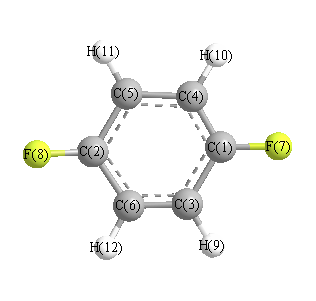
Vibrational Frequencies calculated at B2PLYP/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3237 |
3237 |
0.00 |
288.57 |
0.14 |
0.25 |
| 2 |
Ag |
1655 |
1655 |
0.00 |
4.67 |
0.71 |
0.83 |
| 3 |
Ag |
1278 |
1278 |
0.00 |
25.44 |
0.07 |
0.13 |
| 4 |
Ag |
1166 |
1166 |
0.00 |
7.11 |
0.62 |
0.77 |
| 5 |
Ag |
868 |
868 |
0.00 |
36.80 |
0.06 |
0.11 |
| 6 |
Ag |
457 |
457 |
0.00 |
5.39 |
0.38 |
0.56 |
| 7 |
Au |
963 |
963 |
0.00 |
0.00 |
0.00 |
0.00 |
| 8 |
Au |
437 |
437 |
0.00 |
0.00 |
0.00 |
0.00 |
| 9 |
B1g |
818 |
818 |
0.00 |
0.02 |
0.75 |
0.86 |
| 10 |
B1u |
3223 |
3223 |
3.56 |
0.00 |
0.45 |
0.62 |
| 11 |
B1u |
1548 |
1548 |
241.23 |
0.00 |
0.00 |
0.00 |
| 12 |
B1u |
1236 |
1236 |
194.42 |
0.00 |
0.00 |
0.00 |
| 13 |
B1u |
1031 |
1031 |
1.68 |
0.00 |
0.00 |
0.00 |
| 14 |
B1u |
751 |
751 |
61.71 |
0.00 |
0.00 |
0.00 |
| 15 |
B2g |
931 |
931 |
0.00 |
0.06 |
0.75 |
0.86 |
| 16 |
B2g |
696 |
696 |
0.00 |
0.61 |
0.75 |
0.86 |
| 17 |
B2g |
377 |
377 |
0.00 |
2.99 |
0.75 |
0.86 |
| 18 |
B2u |
3236 |
3236 |
0.17 |
0.00 |
0.34 |
0.51 |
| 19 |
B2u |
1448 |
1448 |
0.47 |
0.00 |
0.72 |
0.84 |
| 20 |
B2u |
1355 |
1355 |
0.13 |
0.00 |
0.54 |
0.70 |
| 21 |
B2u |
1110 |
1110 |
12.01 |
0.00 |
0.66 |
0.79 |
| 22 |
B2u |
349 |
349 |
4.29 |
0.00 |
0.00 |
0.00 |
| 23 |
B3g |
3225 |
3225 |
0.00 |
114.41 |
0.75 |
0.86 |
| 24 |
B3g |
1658 |
1658 |
0.00 |
8.23 |
0.75 |
0.86 |
| 25 |
B3g |
1310 |
1310 |
0.00 |
1.39 |
0.75 |
0.86 |
| 26 |
B3g |
648 |
648 |
0.00 |
6.57 |
0.75 |
0.86 |
| 27 |
B3g |
448 |
448 |
0.00 |
0.07 |
0.75 |
0.86 |
| 28 |
B3u |
857 |
857 |
76.59 |
0.00 |
0.00 |
0.00 |
| 29 |
B3u |
520 |
520 |
18.72 |
0.00 |
0.00 |
0.00 |
| 30 |
B3u |
160 |
160 |
1.53 |
0.00 |
0.00 |
0.00 |
Unscaled Zero Point Vibrational Energy (zpe) 18496.4 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 18496.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.239 |
|
|
|
| 2 |
C |
0.239 |
|
|
|
| 3 |
C |
-0.137 |
|
|
|
| 4 |
C |
-0.137 |
|
|
|
| 5 |
C |
-0.137 |
|
|
|
| 6 |
C |
-0.137 |
|
|
|
| 7 |
F |
-0.213 |
|
|
|
| 8 |
F |
-0.213 |
|
|
|
| 9 |
H |
0.124 |
|
|
|
| 10 |
H |
0.124 |
|
|
|
| 11 |
H |
0.124 |
|
|
|
| 12 |
H |
0.124 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.706 |
0.000 |
0.000 |
| y |
0.000 |
-37.424 |
0.000 |
| z |
0.000 |
0.000 |
-51.954 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.017 |
0.000 |
0.000 |
| y |
0.000 |
11.906 |
0.000 |
| z |
0.000 |
0.000 |
-9.889 |
|
| Polar |
|---|
| 3z2-r2 | -19.778 |
|---|
| x2-y2 | -9.282 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.713 |
0.000 |
0.000 |
| y |
0.000 |
11.384 |
0.000 |
| z |
0.000 |
0.000 |
11.934 |
<r2> (average value of r
2) Å
2
| <r2> |
254.610 |
| (<r2>)1/2 |
15.957 |