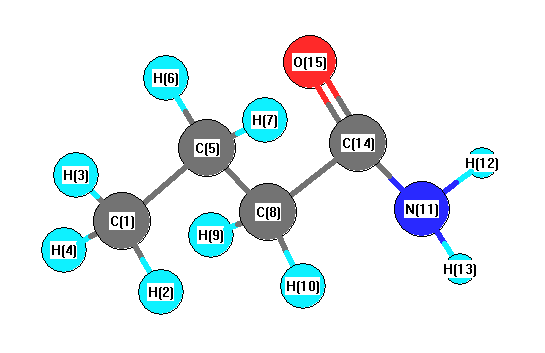
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3965 |
3594 |
51.13 |
|
|
|
| 2 |
A |
3828 |
3470 |
54.53 |
|
|
|
| 3 |
A |
3235 |
2932 |
68.46 |
|
|
|
| 4 |
A |
3219 |
2918 |
59.86 |
|
|
|
| 5 |
A |
3195 |
2896 |
20.98 |
|
|
|
| 6 |
A |
3187 |
2888 |
9.54 |
|
|
|
| 7 |
A |
3180 |
2882 |
18.19 |
|
|
|
| 8 |
A |
3155 |
2860 |
33.91 |
|
|
|
| 9 |
A |
3149 |
2855 |
22.55 |
|
|
|
| 10 |
A |
1933 |
1752 |
408.78 |
|
|
|
| 11 |
A |
1769 |
1604 |
125.23 |
|
|
|
| 12 |
A |
1628 |
1476 |
6.26 |
|
|
|
| 13 |
A |
1616 |
1464 |
6.24 |
|
|
|
| 14 |
A |
1609 |
1458 |
1.72 |
|
|
|
| 15 |
A |
1589 |
1440 |
6.74 |
|
|
|
| 16 |
A |
1556 |
1411 |
62.09 |
|
|
|
| 17 |
A |
1538 |
1394 |
5.53 |
|
|
|
| 18 |
A |
1469 |
1332 |
53.26 |
|
|
|
| 19 |
A |
1437 |
1302 |
4.16 |
|
|
|
| 20 |
A |
1364 |
1236 |
122.10 |
|
|
|
| 21 |
A |
1361 |
1234 |
2.99 |
|
|
|
| 22 |
A |
1228 |
1113 |
1.10 |
|
|
|
| 23 |
A |
1220 |
1106 |
1.33 |
|
|
|
| 24 |
A |
1164 |
1055 |
7.12 |
|
|
|
| 25 |
A |
1111 |
1007 |
2.17 |
|
|
|
| 26 |
A |
990 |
897 |
2.89 |
|
|
|
| 27 |
A |
961 |
871 |
1.76 |
|
|
|
| 28 |
A |
907 |
822 |
1.07 |
|
|
|
| 29 |
A |
810 |
734 |
6.32 |
|
|
|
| 30 |
A |
682 |
618 |
9.24 |
|
|
|
| 31 |
A |
664 |
602 |
18.14 |
|
|
|
| 32 |
A |
542 |
492 |
3.26 |
|
|
|
| 33 |
A |
464 |
420 |
2.48 |
|
|
|
| 34 |
A |
366 |
332 |
3.74 |
|
|
|
| 35 |
A |
262 |
237 |
0.04 |
|
|
|
| 36 |
A |
194 |
176 |
9.77 |
|
|
|
| 37 |
A |
165 |
150 |
224.97 |
|
|
|
| 38 |
A |
106 |
96 |
1.20 |
|
|
|
| 39 |
A |
20 |
19 |
1.77 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30417.7 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 27570.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.178 |
|
|
|
| 2 |
H |
0.046 |
|
|
|
| 3 |
H |
0.066 |
|
|
|
| 4 |
H |
0.044 |
|
|
|
| 5 |
C |
-0.001 |
|
|
|
| 6 |
H |
0.037 |
|
|
|
| 7 |
H |
0.049 |
|
|
|
| 8 |
C |
-0.110 |
|
|
|
| 9 |
H |
0.055 |
|
|
|
| 10 |
H |
0.064 |
|
|
|
| 11 |
N |
-0.421 |
|
|
|
| 12 |
H |
0.207 |
|
|
|
| 13 |
H |
0.188 |
|
|
|
| 14 |
C |
0.442 |
|
|
|
| 15 |
O |
-0.487 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.007 |
-3.773 |
0.132 |
3.775 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.790 |
-6.511 |
0.302 |
| y |
-6.511 |
-39.738 |
-0.284 |
| z |
0.302 |
-0.284 |
-37.994 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.076 |
-6.511 |
0.302 |
| y |
-6.511 |
-3.846 |
-0.284 |
| z |
0.302 |
-0.284 |
-1.230 |
|
| Polar |
|---|
| 3z2-r2 | -2.459 |
|---|
| x2-y2 | 5.948 |
|---|
| xy | -6.511 |
|---|
| xz | 0.302 |
|---|
| yz | -0.284 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.416 |
-0.196 |
-0.045 |
| y |
-0.196 |
8.367 |
0.005 |
| z |
-0.045 |
0.005 |
6.619 |
<r2> (average value of r
2) Å
2
| <r2> |
216.426 |
| (<r2>)1/2 |
14.711 |