Jump to
S1C2
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -321.878450 |
| Energy at 298.15K | -321.887603 |
| HF Energy | -321.878450 |
| Nuclear repulsion energy | 238.494517 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3230 |
2928 |
46.33 |
|
|
|
| 2 |
A' |
3187 |
2888 |
56.11 |
|
|
|
| 3 |
A' |
3177 |
2879 |
7.46 |
|
|
|
| 4 |
A' |
3159 |
2864 |
25.50 |
|
|
|
| 5 |
A' |
1983 |
1798 |
265.75 |
|
|
|
| 6 |
A' |
1650 |
1495 |
11.30 |
|
|
|
| 7 |
A' |
1629 |
1477 |
3.19 |
|
|
|
| 8 |
A' |
1611 |
1461 |
1.56 |
|
|
|
| 9 |
A' |
1562 |
1416 |
21.49 |
|
|
|
| 10 |
A' |
1542 |
1397 |
4.14 |
|
|
|
| 11 |
A' |
1457 |
1321 |
12.59 |
|
|
|
| 12 |
A' |
1267 |
1149 |
16.57 |
|
|
|
| 13 |
A' |
1192 |
1080 |
131.21 |
|
|
|
| 14 |
A' |
1111 |
1007 |
171.28 |
|
|
|
| 15 |
A' |
1103 |
999 |
426.88 |
|
|
|
| 16 |
A' |
976 |
885 |
6.05 |
|
|
|
| 17 |
A' |
779 |
706 |
3.90 |
|
|
|
| 18 |
A' |
443 |
401 |
1.74 |
|
|
|
| 19 |
A' |
383 |
347 |
0.12 |
|
|
|
| 20 |
A' |
173 |
157 |
0.84 |
|
|
|
| 21 |
A" |
3241 |
2937 |
93.85 |
|
|
|
| 22 |
A" |
3216 |
2915 |
11.19 |
|
|
|
| 23 |
A" |
3197 |
2897 |
1.80 |
|
|
|
| 24 |
A" |
1617 |
1465 |
6.49 |
|
|
|
| 25 |
A" |
1432 |
1298 |
0.24 |
|
|
|
| 26 |
A" |
1383 |
1253 |
0.89 |
|
|
|
| 27 |
A" |
1294 |
1173 |
1.68 |
|
|
|
| 28 |
A" |
967 |
877 |
0.96 |
|
|
|
| 29 |
A" |
817 |
740 |
1.12 |
|
|
|
| 30 |
A" |
250 |
227 |
0.02 |
|
|
|
| 31 |
A" |
212 |
192 |
0.57 |
|
|
|
| 32 |
A" |
110 |
99 |
2.39 |
|
|
|
| 33 |
A" |
49 |
45 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24697.3 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 22385.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
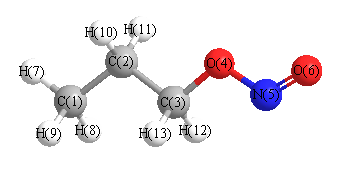
Geometric Data calculated at HF/Def2TZVPP
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.994 |
2.108 |
0.000 |
| C2 |
-1.511 |
0.662 |
0.000 |
| C3 |
0.000 |
0.574 |
0.000 |
| O4 |
0.355 |
-0.802 |
0.000 |
| N5 |
1.675 |
-0.973 |
0.000 |
| O6 |
1.972 |
-2.081 |
0.000 |
| H7 |
-3.076 |
2.149 |
0.000 |
| H8 |
-1.643 |
2.641 |
0.877 |
| H9 |
-1.643 |
2.641 |
-0.877 |
| H10 |
-1.891 |
0.141 |
-0.871 |
| H11 |
-1.891 |
0.141 |
0.871 |
| H12 |
0.418 |
1.048 |
0.879 |
| H13 |
0.418 |
1.048 |
-0.879 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
N5 |
O6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
| C1 | | 1.5243 | 2.5156 | 3.7394 | 4.7908 | 5.7689 | 1.0824 | 1.0843 | 1.0843 | 2.1534 | 2.1534 | 2.7778 | 2.7778 |
C2 | 1.5243 | | 1.5136 | 2.3716 | 3.5809 | 4.4340 | 2.1582 | 2.1686 | 2.1686 | 1.0836 | 1.0836 | 2.1551 | 2.1551 | C3 | 2.5156 | 1.5136 | | 1.4212 | 2.2802 | 3.3078 | 3.4552 | 2.7823 | 2.7823 | 2.1267 | 2.1267 | 1.0826 | 1.0826 | O4 | 3.7394 | 2.3716 | 1.4212 | | 1.3310 | 2.0624 | 4.5247 | 4.0762 | 4.0762 | 2.5868 | 2.5868 | 2.0490 | 2.0490 | N5 | 4.7908 | 3.5809 | 2.2802 | 1.3310 | | 1.1472 | 5.6843 | 4.9841 | 4.9841 | 3.8361 | 3.8361 | 2.5368 | 2.5368 | O6 | 5.7689 | 4.4340 | 3.3078 | 2.0624 | 1.1472 | | 6.5859 | 6.0119 | 6.0119 | 4.5414 | 4.5414 | 3.6025 | 3.6025 | H7 | 1.0824 | 2.1582 | 3.4552 | 4.5247 | 5.6843 | 6.5859 | | 1.7499 | 1.7499 | 2.4883 | 2.4883 | 3.7673 | 3.7673 | H8 | 1.0843 | 2.1686 | 2.7823 | 4.0762 | 4.9841 | 6.0119 | 1.7499 | | 1.7532 | 3.0603 | 2.5124 | 2.6057 | 3.1419 | H9 | 1.0843 | 2.1686 | 2.7823 | 4.0762 | 4.9841 | 6.0119 | 1.7499 | 1.7532 | | 2.5124 | 3.0603 | 3.1419 | 2.6057 | H10 | 2.1534 | 1.0836 | 2.1267 | 2.5868 | 3.8361 | 4.5414 | 2.4883 | 3.0603 | 2.5124 | | 1.7415 | 3.0362 | 2.4813 | H11 | 2.1534 | 1.0836 | 2.1267 | 2.5868 | 3.8361 | 4.5414 | 2.4883 | 2.5124 | 3.0603 | 1.7415 | | 2.4813 | 3.0362 | H12 | 2.7778 | 2.1551 | 1.0826 | 2.0490 | 2.5368 | 3.6025 | 3.7673 | 2.6057 | 3.1419 | 3.0362 | 2.4813 | | 1.7580 | H13 | 2.7778 | 2.1551 | 1.0826 | 2.0490 | 2.5368 | 3.6025 | 3.7673 | 3.1419 | 2.6057 | 2.4813 | 3.0362 | 1.7580 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
111.798 |
|
C1 |
C2 |
H10 |
110.178 |
| C1 |
C2 |
H11 |
110.178 |
|
C2 |
C1 |
H7 |
110.635 |
| C2 |
C1 |
H8 |
111.349 |
|
C2 |
C1 |
H9 |
111.349 |
| C2 |
C3 |
O4 |
107.777 |
|
C2 |
C3 |
H12 |
111.129 |
| C2 |
C3 |
H13 |
111.129 |
|
C3 |
C2 |
H10 |
108.805 |
| C3 |
C2 |
H11 |
108.805 |
|
C3 |
O4 |
N5 |
111.844 |
| O4 |
C3 |
H12 |
109.097 |
|
O4 |
C3 |
H13 |
109.097 |
| O4 |
N5 |
O6 |
112.443 |
|
H7 |
C1 |
H8 |
107.731 |
| H7 |
C1 |
H9 |
107.731 |
|
H8 |
C1 |
H9 |
107.885 |
| H10 |
C2 |
H11 |
106.939 |
|
H12 |
C3 |
H13 |
108.572 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.195 |
|
|
|
| 2 |
C |
-0.019 |
|
|
|
| 3 |
C |
0.096 |
|
|
|
| 4 |
O |
-0.221 |
|
|
|
| 5 |
N |
0.273 |
|
|
|
| 6 |
O |
-0.305 |
|
|
|
| 7 |
H |
0.069 |
|
|
|
| 8 |
H |
0.053 |
|
|
|
| 9 |
H |
0.053 |
|
|
|
| 10 |
H |
0.043 |
|
|
|
| 11 |
H |
0.043 |
|
|
|
| 12 |
H |
0.055 |
|
|
|
| 13 |
H |
0.055 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.670 |
2.738 |
0.000 |
3.207 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.615 |
2.271 |
0.000 |
| y |
2.271 |
-40.497 |
0.000 |
| z |
0.000 |
0.000 |
-34.735 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.998 |
2.271 |
0.000 |
| y |
2.271 |
-3.822 |
0.000 |
| z |
0.000 |
0.000 |
4.820 |
|
| Polar |
|---|
| 3z2-r2 | 9.641 |
|---|
| x2-y2 | 1.882 |
|---|
| xy | 2.271 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.078 |
-1.779 |
0.000 |
| y |
-1.779 |
8.379 |
0.000 |
| z |
0.000 |
0.000 |
5.985 |
<r2> (average value of r
2) Å
2
| <r2> |
238.342 |
| (<r2>)1/2 |
15.438 |
Jump to
S1C1
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -321.878235 |
| Energy at 298.15K | -321.887499 |
| HF Energy | -321.878235 |
| Nuclear repulsion energy | 242.481855 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3242 |
2938 |
47.87 |
|
|
|
| 2 |
A |
3236 |
2933 |
58.21 |
|
|
|
| 3 |
A |
3220 |
2918 |
49.76 |
|
|
|
| 4 |
A |
3200 |
2900 |
27.86 |
|
|
|
| 5 |
A |
3186 |
2888 |
19.61 |
|
|
|
| 6 |
A |
3167 |
2871 |
12.97 |
|
|
|
| 7 |
A |
3162 |
2866 |
35.05 |
|
|
|
| 8 |
A |
1982 |
1797 |
258.50 |
|
|
|
| 9 |
A |
1647 |
1493 |
8.23 |
|
|
|
| 10 |
A |
1625 |
1472 |
8.82 |
|
|
|
| 11 |
A |
1613 |
1462 |
7.63 |
|
|
|
| 12 |
A |
1602 |
1452 |
1.18 |
|
|
|
| 13 |
A |
1552 |
1406 |
23.78 |
|
|
|
| 14 |
A |
1542 |
1398 |
4.86 |
|
|
|
| 15 |
A |
1502 |
1361 |
3.55 |
|
|
|
| 16 |
A |
1419 |
1286 |
2.77 |
|
|
|
| 17 |
A |
1385 |
1255 |
2.83 |
|
|
|
| 18 |
A |
1289 |
1168 |
16.16 |
|
|
|
| 19 |
A |
1254 |
1136 |
14.38 |
|
|
|
| 20 |
A |
1176 |
1066 |
122.43 |
|
|
|
| 21 |
A |
1104 |
1000 |
498.62 |
|
|
|
| 22 |
A |
1099 |
996 |
77.53 |
|
|
|
| 23 |
A |
998 |
904 |
1.21 |
|
|
|
| 24 |
A |
939 |
851 |
0.22 |
|
|
|
| 25 |
A |
855 |
775 |
6.45 |
|
|
|
| 26 |
A |
717 |
650 |
1.90 |
|
|
|
| 27 |
A |
517 |
469 |
0.64 |
|
|
|
| 28 |
A |
355 |
322 |
0.60 |
|
|
|
| 29 |
A |
293 |
265 |
0.38 |
|
|
|
| 30 |
A |
207 |
188 |
0.37 |
|
|
|
| 31 |
A |
194 |
176 |
0.25 |
|
|
|
| 32 |
A |
123 |
111 |
1.72 |
|
|
|
| 33 |
A |
48 |
43 |
0.33 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24722.6 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 22408.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.837 |
1.124 |
0.070 |
| C2 |
1.746 |
-0.370 |
-0.212 |
| C3 |
0.468 |
-1.012 |
0.289 |
| O4 |
-0.661 |
-0.503 |
-0.414 |
| N5 |
-1.514 |
0.146 |
0.418 |
| O6 |
-2.402 |
0.548 |
-0.171 |
| H7 |
2.772 |
1.527 |
-0.298 |
| H8 |
1.784 |
1.327 |
1.134 |
| H9 |
1.033 |
1.664 |
-0.414 |
| H10 |
1.835 |
-0.554 |
-1.276 |
| H11 |
2.574 |
-0.887 |
0.264 |
| H12 |
0.479 |
-2.078 |
0.115 |
| H13 |
0.325 |
-0.835 |
1.346 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
N5 |
O6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
| C1 | | 1.5229 | 2.5465 | 3.0198 | 3.5082 | 4.2849 | 1.0827 | 1.0848 | 1.0824 | 2.1516 | 2.1512 | 3.4786 | 2.7848 |
C2 | 1.5229 | | 1.5157 | 2.4192 | 3.3605 | 4.2493 | 2.1581 | 2.1659 | 2.1646 | 1.0843 | 1.0859 | 2.1521 | 2.1595 | C3 | 2.5465 | 1.5157 | | 1.4231 | 2.2986 | 3.2987 | 3.4784 | 2.8139 | 2.8232 | 2.1277 | 2.1102 | 1.0805 | 1.0813 | O4 | 3.0198 | 2.4192 | 1.4231 | | 1.3569 | 2.0487 | 3.9897 | 3.4239 | 2.7500 | 2.6408 | 3.3273 | 2.0149 | 2.0440 | N5 | 3.5082 | 3.3605 | 2.2986 | 1.3569 | | 1.1395 | 4.5598 | 3.5760 | 3.0795 | 3.8179 | 4.2194 | 3.0016 | 2.2812 | O6 | 4.2849 | 4.2493 | 3.2987 | 2.0487 | 1.1395 | | 5.2676 | 4.4540 | 3.6198 | 4.5156 | 5.1976 | 3.9092 | 3.4135 | H7 | 1.0827 | 2.1581 | 3.4784 | 3.9897 | 4.5598 | 5.2676 | | 1.7514 | 1.7485 | 2.4834 | 2.4869 | 4.2925 | 3.7781 | H8 | 1.0848 | 2.1659 | 2.8139 | 3.4239 | 3.5760 | 4.4540 | 1.7514 | | 1.7535 | 3.0581 | 2.5067 | 3.7865 | 2.6175 | H9 | 1.0824 | 2.1646 | 2.8232 | 2.7500 | 3.0795 | 3.6198 | 1.7485 | 1.7535 | | 2.5114 | 3.0567 | 3.8194 | 3.1375 | H10 | 2.1516 | 1.0843 | 2.1277 | 2.6408 | 3.8179 | 4.5156 | 2.4834 | 3.0581 | 2.5114 | | 1.7409 | 2.4690 | 3.0389 | H11 | 2.1512 | 1.0859 | 2.1102 | 3.3273 | 4.2194 | 5.1976 | 2.4869 | 2.5067 | 3.0567 | 1.7409 | | 2.4144 | 2.4964 | H12 | 3.4786 | 2.1521 | 1.0805 | 2.0149 | 3.0016 | 3.9092 | 4.2925 | 3.7865 | 3.8194 | 2.4690 | 2.4144 | | 1.7558 | H13 | 2.7848 | 2.1595 | 1.0813 | 2.0440 | 2.2812 | 3.4135 | 3.7781 | 2.6175 | 3.1375 | 3.0389 | 2.4964 | 1.7558 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
113.868 |
|
C1 |
C2 |
H10 |
110.094 |
| C1 |
C2 |
H11 |
109.959 |
|
C2 |
C1 |
H7 |
110.702 |
| C2 |
C1 |
H8 |
111.207 |
|
C2 |
C1 |
H9 |
111.245 |
| C2 |
C3 |
O4 |
110.774 |
|
C2 |
C3 |
H12 |
110.864 |
| C2 |
C3 |
H13 |
111.416 |
|
C3 |
C2 |
H10 |
108.704 |
| C3 |
C2 |
H11 |
107.252 |
|
C3 |
O4 |
N5 |
111.522 |
| O4 |
C3 |
H12 |
106.371 |
|
O4 |
C3 |
H13 |
108.631 |
| O4 |
N5 |
O6 |
110.000 |
|
H7 |
C1 |
H8 |
107.803 |
| H7 |
C1 |
H9 |
107.718 |
|
H8 |
C1 |
H9 |
108.016 |
| H10 |
C2 |
H11 |
106.680 |
|
H12 |
C3 |
H13 |
108.618 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.181 |
|
|
|
| 2 |
C |
-0.029 |
|
|
|
| 3 |
C |
0.088 |
|
|
|
| 4 |
O |
-0.227 |
|
|
|
| 5 |
N |
0.270 |
|
|
|
| 6 |
O |
-0.306 |
|
|
|
| 7 |
H |
0.063 |
|
|
|
| 8 |
H |
0.044 |
|
|
|
| 9 |
H |
0.076 |
|
|
|
| 10 |
H |
0.042 |
|
|
|
| 11 |
H |
0.033 |
|
|
|
| 12 |
H |
0.067 |
|
|
|
| 13 |
H |
0.059 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.669 |
-1.463 |
0.544 |
3.092 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.480 |
0.771 |
-0.211 |
| y |
0.771 |
-35.823 |
0.801 |
| z |
-0.211 |
0.801 |
-35.938 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.600 |
0.771 |
-0.211 |
| y |
0.771 |
2.386 |
0.801 |
| z |
-0.211 |
0.801 |
2.213 |
|
| Polar |
|---|
| 3z2-r2 | 4.427 |
|---|
| x2-y2 | -4.658 |
|---|
| xy | 0.771 |
|---|
| xz | -0.211 |
|---|
| yz | 0.801 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.237 |
-0.532 |
0.071 |
| y |
-0.532 |
6.895 |
-0.117 |
| z |
0.071 |
-0.117 |
6.179 |
<r2> (average value of r
2) Å
2
| <r2> |
209.192 |
| (<r2>)1/2 |
14.463 |
