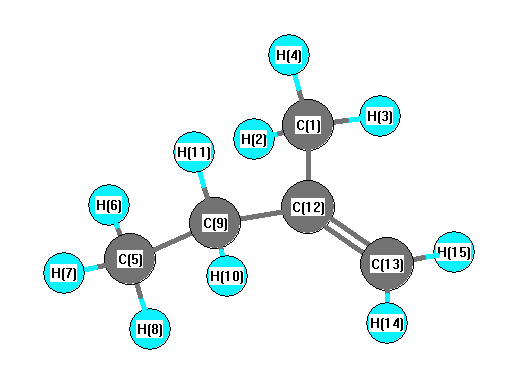
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3353 |
3039 |
23.39 |
|
|
|
| 2 |
A |
3278 |
2971 |
12.07 |
|
|
|
| 3 |
A |
3241 |
2938 |
29.76 |
|
|
|
| 4 |
A |
3230 |
2928 |
66.57 |
|
|
|
| 5 |
A |
3219 |
2918 |
57.82 |
|
|
|
| 6 |
A |
3204 |
2904 |
23.17 |
|
|
|
| 7 |
A |
3197 |
2898 |
15.67 |
|
|
|
| 8 |
A |
3163 |
2867 |
35.63 |
|
|
|
| 9 |
A |
3157 |
2861 |
33.83 |
|
|
|
| 10 |
A |
3145 |
2851 |
27.76 |
|
|
|
| 11 |
A |
1845 |
1673 |
30.49 |
|
|
|
| 12 |
A |
1628 |
1475 |
3.90 |
|
|
|
| 13 |
A |
1618 |
1467 |
5.37 |
|
|
|
| 14 |
A |
1615 |
1463 |
11.14 |
|
|
|
| 15 |
A |
1607 |
1456 |
7.64 |
|
|
|
| 16 |
A |
1596 |
1446 |
3.60 |
|
|
|
| 17 |
A |
1569 |
1422 |
0.31 |
|
|
|
| 18 |
A |
1537 |
1393 |
2.76 |
|
|
|
| 19 |
A |
1532 |
1389 |
2.46 |
|
|
|
| 20 |
A |
1477 |
1338 |
3.83 |
|
|
|
| 21 |
A |
1417 |
1284 |
1.43 |
|
|
|
| 22 |
A |
1374 |
1246 |
1.58 |
|
|
|
| 23 |
A |
1214 |
1100 |
0.76 |
|
|
|
| 24 |
A |
1190 |
1078 |
3.55 |
|
|
|
| 25 |
A |
1138 |
1031 |
0.83 |
|
|
|
| 26 |
A |
1088 |
986 |
0.13 |
|
|
|
| 27 |
A |
1083 |
982 |
0.75 |
|
|
|
| 28 |
A |
1045 |
947 |
53.99 |
|
|
|
| 29 |
A |
1002 |
908 |
1.03 |
|
|
|
| 30 |
A |
857 |
777 |
1.19 |
|
|
|
| 31 |
A |
820 |
743 |
0.68 |
|
|
|
| 32 |
A |
781 |
708 |
0.16 |
|
|
|
| 33 |
A |
583 |
529 |
8.05 |
|
|
|
| 34 |
A |
467 |
424 |
1.94 |
|
|
|
| 35 |
A |
425 |
385 |
0.93 |
|
|
|
| 36 |
A |
280 |
254 |
0.38 |
|
|
|
| 37 |
A |
241 |
218 |
0.40 |
|
|
|
| 38 |
A |
196 |
178 |
0.12 |
|
|
|
| 39 |
A |
71 |
64 |
0.05 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31739.7 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 28768.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.167 |
|
|
|
| 2 |
H |
0.064 |
|
|
|
| 3 |
H |
0.067 |
|
|
|
| 4 |
H |
0.060 |
|
|
|
| 5 |
C |
-0.184 |
|
|
|
| 6 |
H |
0.048 |
|
|
|
| 7 |
H |
0.061 |
|
|
|
| 8 |
H |
0.059 |
|
|
|
| 9 |
C |
-0.009 |
|
|
|
| 10 |
H |
0.050 |
|
|
|
| 11 |
H |
0.036 |
|
|
|
| 12 |
C |
-0.034 |
|
|
|
| 13 |
C |
-0.156 |
|
|
|
| 14 |
H |
0.052 |
|
|
|
| 15 |
H |
0.053 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.387 |
0.385 |
0.138 |
0.563 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.450 |
0.636 |
-0.922 |
| y |
0.636 |
-32.826 |
0.182 |
| z |
-0.922 |
0.182 |
-34.984 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.455 |
0.636 |
-0.922 |
| y |
0.636 |
1.391 |
0.182 |
| z |
-0.922 |
0.182 |
-1.845 |
|
| Polar |
|---|
| 3z2-r2 | -3.691 |
|---|
| x2-y2 | -0.624 |
|---|
| xy | 0.636 |
|---|
| xz | -0.922 |
|---|
| yz | 0.182 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.793 |
-0.995 |
-0.454 |
| y |
-0.995 |
9.381 |
0.341 |
| z |
-0.454 |
0.341 |
7.254 |
<r2> (average value of r
2) Å
2
| <r2> |
142.980 |
| (<r2>)1/2 |
11.957 |