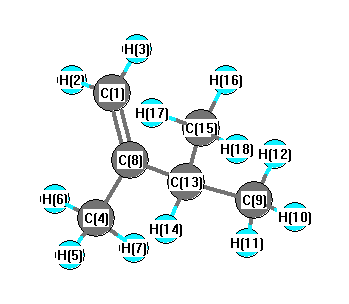
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3351 |
3038 |
24.24 |
|
|
|
| 2 |
A' |
3278 |
2971 |
13.50 |
|
|
|
| 3 |
A' |
3243 |
2939 |
31.04 |
|
|
|
| 4 |
A' |
3227 |
2925 |
78.72 |
|
|
|
| 5 |
A' |
3220 |
2918 |
89.28 |
|
|
|
| 6 |
A' |
3173 |
2876 |
4.77 |
|
|
|
| 7 |
A' |
3167 |
2870 |
36.02 |
|
|
|
| 8 |
A' |
3161 |
2865 |
17.58 |
|
|
|
| 9 |
A' |
1841 |
1668 |
30.56 |
|
|
|
| 10 |
A' |
1630 |
1477 |
4.36 |
|
|
|
| 11 |
A' |
1624 |
1472 |
8.18 |
|
|
|
| 12 |
A' |
1614 |
1463 |
12.78 |
|
|
|
| 13 |
A' |
1576 |
1428 |
0.93 |
|
|
|
| 14 |
A' |
1544 |
1399 |
1.89 |
|
|
|
| 15 |
A' |
1536 |
1392 |
5.44 |
|
|
|
| 16 |
A' |
1458 |
1321 |
6.83 |
|
|
|
| 17 |
A' |
1412 |
1280 |
0.29 |
|
|
|
| 18 |
A' |
1288 |
1167 |
2.81 |
|
|
|
| 19 |
A' |
1205 |
1092 |
7.42 |
|
|
|
| 20 |
A' |
1094 |
992 |
0.87 |
|
|
|
| 21 |
A' |
1036 |
939 |
0.45 |
|
|
|
| 22 |
A' |
954 |
865 |
0.73 |
|
|
|
| 23 |
A' |
769 |
697 |
0.62 |
|
|
|
| 24 |
A' |
556 |
504 |
0.67 |
|
|
|
| 25 |
A' |
480 |
435 |
1.20 |
|
|
|
| 26 |
A' |
355 |
322 |
0.13 |
|
|
|
| 27 |
A' |
297 |
270 |
0.56 |
|
|
|
| 28 |
A' |
270 |
245 |
0.08 |
|
|
|
| 29 |
A" |
3222 |
2921 |
5.09 |
|
|
|
| 30 |
A" |
3216 |
2915 |
39.11 |
|
|
|
| 31 |
A" |
3207 |
2906 |
16.56 |
|
|
|
| 32 |
A" |
3157 |
2861 |
31.42 |
|
|
|
| 33 |
A" |
1617 |
1466 |
0.28 |
|
|
|
| 34 |
A" |
1606 |
1456 |
0.87 |
|
|
|
| 35 |
A" |
1598 |
1449 |
7.56 |
|
|
|
| 36 |
A" |
1522 |
1380 |
4.71 |
|
|
|
| 37 |
A" |
1463 |
1326 |
2.00 |
|
|
|
| 38 |
A" |
1213 |
1100 |
3.64 |
|
|
|
| 39 |
A" |
1154 |
1046 |
0.59 |
|
|
|
| 40 |
A" |
1046 |
948 |
51.55 |
|
|
|
| 41 |
A" |
1032 |
936 |
0.33 |
|
|
|
| 42 |
A" |
1001 |
907 |
0.16 |
|
|
|
| 43 |
A" |
804 |
728 |
0.97 |
|
|
|
| 44 |
A" |
604 |
548 |
9.69 |
|
|
|
| 45 |
A" |
279 |
253 |
0.18 |
|
|
|
| 46 |
A" |
229 |
208 |
0.46 |
|
|
|
| 47 |
A" |
184 |
167 |
0.07 |
|
|
|
| 48 |
A" |
45 |
40 |
0.11 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 38277.5 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 34694.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.155 |
|
|
|
| 2 |
C |
-0.051 |
|
|
|
| 3 |
C |
0.176 |
|
|
|
| 4 |
C |
-0.199 |
|
|
|
| 5 |
C |
-0.231 |
|
|
|
| 6 |
C |
-0.231 |
|
|
|
| 7 |
H |
0.054 |
|
|
|
| 8 |
H |
0.051 |
|
|
|
| 9 |
H |
0.019 |
|
|
|
| 10 |
H |
0.069 |
|
|
|
| 11 |
H |
0.066 |
|
|
|
| 12 |
H |
0.066 |
|
|
|
| 13 |
H |
0.061 |
|
|
|
| 14 |
H |
0.072 |
|
|
|
| 15 |
H |
0.050 |
|
|
|
| 16 |
H |
0.061 |
|
|
|
| 17 |
H |
0.072 |
|
|
|
| 18 |
H |
0.050 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.281 |
0.427 |
0.000 |
0.511 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.345 |
-0.775 |
0.000 |
| y |
-0.775 |
-40.236 |
0.000 |
| z |
0.000 |
0.000 |
-42.243 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.894 |
-0.775 |
0.000 |
| y |
-0.775 |
0.559 |
0.000 |
| z |
0.000 |
0.000 |
-2.453 |
|
| Polar |
|---|
| 3z2-r2 | -4.906 |
|---|
| x2-y2 | 0.890 |
|---|
| xy | -0.775 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.434 |
0.958 |
0.000 |
| y |
0.958 |
11.738 |
0.000 |
| z |
0.000 |
0.000 |
9.106 |
<r2> (average value of r
2) Å
2
| <r2> |
190.503 |
| (<r2>)1/2 |
13.802 |