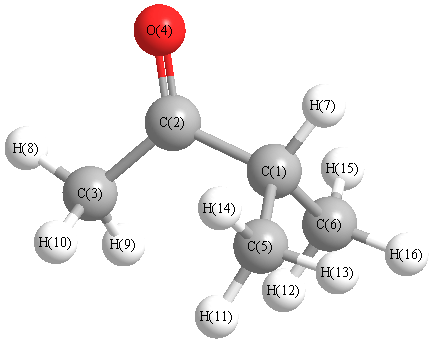
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3278 |
2971 |
16.93 |
|
|
|
| 2 |
A |
3237 |
2934 |
54.12 |
|
|
|
| 3 |
A |
3232 |
2930 |
41.17 |
|
|
|
| 4 |
A |
3231 |
2928 |
50.04 |
|
|
|
| 5 |
A |
3221 |
2919 |
27.03 |
|
|
|
| 6 |
A |
3218 |
2917 |
10.51 |
|
|
|
| 7 |
A |
3198 |
2899 |
0.88 |
|
|
|
| 8 |
A |
3177 |
2880 |
7.33 |
|
|
|
| 9 |
A |
3164 |
2868 |
14.22 |
|
|
|
| 10 |
A |
3160 |
2864 |
31.45 |
|
|
|
| 11 |
A |
1969 |
1785 |
256.69 |
|
|
|
| 12 |
A |
1629 |
1477 |
16.69 |
|
|
|
| 13 |
A |
1623 |
1471 |
5.85 |
|
|
|
| 14 |
A |
1613 |
1462 |
2.17 |
|
|
|
| 15 |
A |
1608 |
1457 |
1.02 |
|
|
|
| 16 |
A |
1598 |
1449 |
7.65 |
|
|
|
| 17 |
A |
1584 |
1435 |
11.50 |
|
|
|
| 18 |
A |
1547 |
1402 |
3.68 |
|
|
|
| 19 |
A |
1526 |
1383 |
3.04 |
|
|
|
| 20 |
A |
1522 |
1379 |
32.04 |
|
|
|
| 21 |
A |
1471 |
1333 |
2.34 |
|
|
|
| 22 |
A |
1458 |
1321 |
2.18 |
|
|
|
| 23 |
A |
1366 |
1238 |
45.45 |
|
|
|
| 24 |
A |
1287 |
1166 |
4.23 |
|
|
|
| 25 |
A |
1224 |
1110 |
1.23 |
|
|
|
| 26 |
A |
1190 |
1078 |
14.42 |
|
|
|
| 27 |
A |
1097 |
994 |
2.37 |
|
|
|
| 28 |
A |
1041 |
944 |
0.48 |
|
|
|
| 29 |
A |
1025 |
929 |
16.09 |
|
|
|
| 30 |
A |
1006 |
912 |
0.53 |
|
|
|
| 31 |
A |
955 |
866 |
2.91 |
|
|
|
| 32 |
A |
769 |
697 |
3.74 |
|
|
|
| 33 |
A |
655 |
593 |
1.21 |
|
|
|
| 34 |
A |
564 |
512 |
12.34 |
|
|
|
| 35 |
A |
534 |
484 |
5.98 |
|
|
|
| 36 |
A |
356 |
323 |
1.62 |
|
|
|
| 37 |
A |
291 |
264 |
1.76 |
|
|
|
| 38 |
A |
274 |
248 |
0.40 |
|
|
|
| 39 |
A |
253 |
229 |
0.45 |
|
|
|
| 40 |
A |
225 |
204 |
0.68 |
|
|
|
| 41 |
A |
139 |
126 |
0.04 |
|
|
|
| 42 |
A |
56 |
51 |
4.75 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32784.0 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 29715.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.417 |
|
|
|
| 2 |
C |
0.298 |
|
|
|
| 3 |
H |
0.072 |
|
|
|
| 4 |
H |
0.110 |
|
|
|
| 5 |
H |
0.066 |
|
|
|
| 6 |
C |
-0.233 |
|
|
|
| 7 |
H |
0.044 |
|
|
|
| 8 |
C |
0.131 |
|
|
|
| 9 |
H |
0.073 |
|
|
|
| 10 |
H |
0.086 |
|
|
|
| 11 |
H |
0.055 |
|
|
|
| 12 |
H |
0.053 |
|
|
|
| 13 |
H |
0.065 |
|
|
|
| 14 |
H |
0.070 |
|
|
|
| 15 |
C |
-0.240 |
|
|
|
| 16 |
C |
-0.233 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.148 |
1.797 |
1.556 |
3.203 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.475 |
3.735 |
3.398 |
| y |
3.735 |
-39.302 |
-1.596 |
| z |
3.398 |
-1.596 |
-39.049 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.299 |
3.735 |
3.398 |
| y |
3.735 |
0.960 |
-1.596 |
| z |
3.398 |
-1.596 |
1.339 |
|
| Polar |
|---|
| 3z2-r2 | 2.678 |
|---|
| x2-y2 | -2.173 |
|---|
| xy | 3.735 |
|---|
| xz | 3.398 |
|---|
| yz | -1.596 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.322 |
-0.136 |
-0.104 |
| y |
-0.136 |
8.624 |
0.274 |
| z |
-0.104 |
0.274 |
8.137 |
<r2> (average value of r
2) Å
2
| <r2> |
179.085 |
| (<r2>)1/2 |
13.382 |