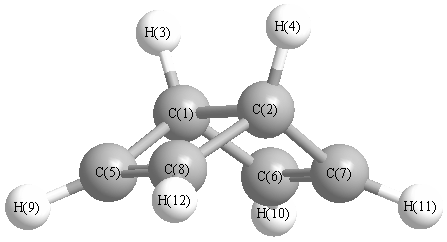
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3360 |
3046 |
7.90 |
254.95 |
0.03 |
0.07 |
| 2 |
A1 |
3247 |
2943 |
45.40 |
146.64 |
0.12 |
0.21 |
| 3 |
A1 |
1762 |
1597 |
0.14 |
62.59 |
0.05 |
0.09 |
| 4 |
A1 |
1299 |
1178 |
4.48 |
29.01 |
0.12 |
0.22 |
| 5 |
A1 |
1143 |
1036 |
1.06 |
11.77 |
0.40 |
0.57 |
| 6 |
A1 |
1024 |
928 |
28.25 |
11.87 |
0.22 |
0.36 |
| 7 |
A1 |
941 |
853 |
0.00 |
4.67 |
0.62 |
0.76 |
| 8 |
A1 |
894 |
810 |
77.10 |
9.43 |
0.05 |
0.09 |
| 9 |
A1 |
421 |
382 |
5.46 |
4.89 |
0.51 |
0.67 |
| 10 |
A2 |
3325 |
3014 |
0.00 |
147.16 |
0.75 |
0.86 |
| 11 |
A2 |
1427 |
1293 |
0.00 |
5.90 |
0.75 |
0.86 |
| 12 |
A2 |
1325 |
1201 |
0.00 |
8.76 |
0.75 |
0.86 |
| 13 |
A2 |
1069 |
969 |
0.00 |
1.35 |
0.75 |
0.86 |
| 14 |
A2 |
1022 |
926 |
0.00 |
1.17 |
0.75 |
0.86 |
| 15 |
A2 |
857 |
776 |
0.00 |
3.81 |
0.75 |
0.86 |
| 16 |
A2 |
366 |
332 |
0.00 |
1.79 |
0.75 |
0.86 |
| 17 |
B1 |
3358 |
3043 |
37.09 |
56.61 |
0.75 |
0.86 |
| 18 |
B1 |
1734 |
1571 |
15.08 |
0.10 |
0.75 |
0.86 |
| 19 |
B1 |
1346 |
1220 |
9.42 |
0.23 |
0.75 |
0.86 |
| 20 |
B1 |
1191 |
1080 |
0.28 |
0.01 |
0.75 |
0.86 |
| 21 |
B1 |
1071 |
971 |
2.08 |
8.49 |
0.75 |
0.86 |
| 22 |
B1 |
804 |
729 |
48.29 |
1.04 |
0.75 |
0.86 |
| 23 |
B2 |
3327 |
3016 |
30.89 |
40.22 |
0.75 |
0.86 |
| 24 |
B2 |
3238 |
2935 |
27.06 |
116.33 |
0.75 |
0.86 |
| 25 |
B2 |
1424 |
1291 |
43.78 |
0.01 |
0.75 |
0.86 |
| 26 |
B2 |
1287 |
1166 |
6.71 |
0.36 |
0.75 |
0.86 |
| 27 |
B2 |
1091 |
989 |
3.04 |
3.59 |
0.75 |
0.86 |
| 28 |
B2 |
1021 |
925 |
5.73 |
0.00 |
0.75 |
0.86 |
| 29 |
B2 |
912 |
827 |
10.71 |
0.78 |
0.75 |
0.86 |
| 30 |
B2 |
542 |
492 |
7.46 |
2.81 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 22912.8 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 20768.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.077 |
|
|
|
| 2 |
C |
0.077 |
|
|
|
| 3 |
H |
0.050 |
|
|
|
| 4 |
H |
0.050 |
|
|
|
| 5 |
C |
-0.156 |
|
|
|
| 6 |
C |
-0.156 |
|
|
|
| 7 |
C |
-0.156 |
|
|
|
| 8 |
C |
-0.156 |
|
|
|
| 9 |
H |
0.092 |
|
|
|
| 10 |
H |
0.092 |
|
|
|
| 11 |
H |
0.092 |
|
|
|
| 12 |
H |
0.092 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.096 |
0.096 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.116 |
0.000 |
0.000 |
| y |
0.000 |
-32.795 |
0.000 |
| z |
0.000 |
0.000 |
-36.737 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.350 |
0.000 |
0.000 |
| y |
0.000 |
3.631 |
0.000 |
| z |
0.000 |
0.000 |
-2.282 |
|
| Polar |
|---|
| 3z2-r2 | -4.563 |
|---|
| x2-y2 | -3.321 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.234 |
0.000 |
0.000 |
| y |
0.000 |
9.940 |
0.000 |
| z |
0.000 |
0.000 |
7.009 |
<r2> (average value of r
2) Å
2
| <r2> |
117.823 |
| (<r2>)1/2 |
10.855 |