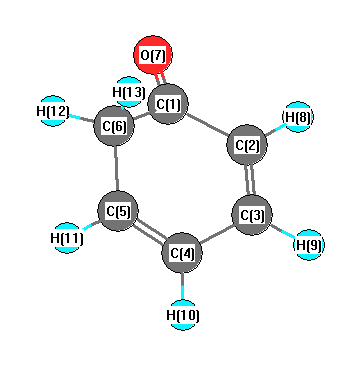
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3349 |
3035 |
0.20 |
|
|
|
| 2 |
A1 |
3309 |
2999 |
10.45 |
|
|
|
| 3 |
A1 |
3146 |
2852 |
19.40 |
|
|
|
| 4 |
A1 |
1946 |
1764 |
516.37 |
|
|
|
| 5 |
A1 |
1846 |
1674 |
0.61 |
|
|
|
| 6 |
A1 |
1568 |
1422 |
8.46 |
|
|
|
| 7 |
A1 |
1533 |
1389 |
4.46 |
|
|
|
| 8 |
A1 |
1285 |
1165 |
10.02 |
|
|
|
| 9 |
A1 |
1043 |
945 |
3.76 |
|
|
|
| 10 |
A1 |
929 |
842 |
5.36 |
|
|
|
| 11 |
A1 |
814 |
738 |
2.84 |
|
|
|
| 12 |
A1 |
538 |
487 |
3.54 |
|
|
|
| 13 |
A2 |
1324 |
1200 |
0.00 |
|
|
|
| 14 |
A2 |
1128 |
1022 |
0.00 |
|
|
|
| 15 |
A2 |
826 |
749 |
0.00 |
|
|
|
| 16 |
A2 |
399 |
362 |
0.00 |
|
|
|
| 17 |
B1 |
3161 |
2865 |
11.91 |
|
|
|
| 18 |
B1 |
1150 |
1043 |
1.31 |
|
|
|
| 19 |
B1 |
1049 |
951 |
39.61 |
|
|
|
| 20 |
B1 |
945 |
857 |
44.86 |
|
|
|
| 21 |
B1 |
637 |
577 |
21.58 |
|
|
|
| 22 |
B1 |
348 |
316 |
2.31 |
|
|
|
| 23 |
B1 |
134 |
122 |
1.74 |
|
|
|
| 24 |
B2 |
3347 |
3034 |
16.04 |
|
|
|
| 25 |
B2 |
3308 |
2999 |
13.70 |
|
|
|
| 26 |
B2 |
1809 |
1640 |
0.80 |
|
|
|
| 27 |
B2 |
1542 |
1398 |
29.44 |
|
|
|
| 28 |
B2 |
1506 |
1365 |
4.92 |
|
|
|
| 29 |
B2 |
1377 |
1248 |
42.84 |
|
|
|
| 30 |
B2 |
1231 |
1115 |
9.63 |
|
|
|
| 31 |
B2 |
1052 |
953 |
5.35 |
|
|
|
| 32 |
B2 |
616 |
558 |
0.36 |
|
|
|
| 33 |
B2 |
493 |
447 |
18.20 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24343.6 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 22065.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.358 |
|
|
|
| 2 |
C |
0.115 |
|
|
|
| 3 |
C |
-0.134 |
|
|
|
| 4 |
C |
-0.134 |
|
|
|
| 5 |
C |
-0.136 |
|
|
|
| 6 |
C |
-0.136 |
|
|
|
| 7 |
O |
-0.443 |
|
|
|
| 8 |
H |
0.109 |
|
|
|
| 9 |
H |
0.109 |
|
|
|
| 10 |
H |
0.087 |
|
|
|
| 11 |
H |
0.087 |
|
|
|
| 12 |
H |
0.059 |
|
|
|
| 13 |
H |
0.059 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-4.833 |
4.833 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.729 |
0.000 |
0.000 |
| y |
0.000 |
-36.528 |
0.000 |
| z |
0.000 |
0.000 |
-46.374 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.278 |
0.000 |
0.000 |
| y |
0.000 |
8.023 |
0.000 |
| z |
0.000 |
0.000 |
-6.745 |
|
| Polar |
|---|
| 3z2-r2 | -13.491 |
|---|
| x2-y2 | -6.201 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.229 |
0.000 |
0.000 |
| y |
0.000 |
9.111 |
0.000 |
| z |
0.000 |
0.000 |
13.975 |
<r2> (average value of r
2) Å
2
| <r2> |
185.231 |
| (<r2>)1/2 |
13.610 |