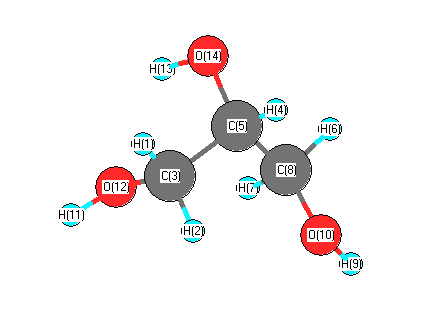
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
4194 |
3801 |
73.86 |
|
|
|
| 2 |
A |
4191 |
3799 |
71.63 |
|
|
|
| 3 |
A |
4149 |
3761 |
69.51 |
|
|
|
| 4 |
A |
3222 |
2921 |
43.31 |
|
|
|
| 5 |
A |
3217 |
2916 |
71.04 |
|
|
|
| 6 |
A |
3193 |
2894 |
27.64 |
|
|
|
| 7 |
A |
3165 |
2869 |
18.59 |
|
|
|
| 8 |
A |
3162 |
2866 |
85.15 |
|
|
|
| 9 |
A |
1649 |
1495 |
6.94 |
|
|
|
| 10 |
A |
1641 |
1487 |
8.57 |
|
|
|
| 11 |
A |
1596 |
1447 |
3.47 |
|
|
|
| 12 |
A |
1584 |
1436 |
3.69 |
|
|
|
| 13 |
A |
1544 |
1399 |
45.81 |
|
|
|
| 14 |
A |
1496 |
1356 |
10.29 |
|
|
|
| 15 |
A |
1423 |
1289 |
66.82 |
|
|
|
| 16 |
A |
1395 |
1265 |
19.32 |
|
|
|
| 17 |
A |
1355 |
1228 |
21.58 |
|
|
|
| 18 |
A |
1325 |
1201 |
13.10 |
|
|
|
| 19 |
A |
1284 |
1164 |
52.42 |
|
|
|
| 20 |
A |
1249 |
1132 |
31.19 |
|
|
|
| 21 |
A |
1214 |
1100 |
142.92 |
|
|
|
| 22 |
A |
1174 |
1064 |
102.77 |
|
|
|
| 23 |
A |
1155 |
1047 |
55.32 |
|
|
|
| 24 |
A |
1080 |
978 |
17.58 |
|
|
|
| 25 |
A |
1030 |
933 |
7.22 |
|
|
|
| 26 |
A |
873 |
792 |
15.20 |
|
|
|
| 27 |
A |
695 |
630 |
31.84 |
|
|
|
| 28 |
A |
521 |
472 |
6.37 |
|
|
|
| 29 |
A |
442 |
401 |
68.93 |
|
|
|
| 30 |
A |
419 |
380 |
77.67 |
|
|
|
| 31 |
A |
295 |
268 |
3.04 |
|
|
|
| 32 |
A |
270 |
245 |
14.00 |
|
|
|
| 33 |
A |
245 |
222 |
140.35 |
|
|
|
| 34 |
A |
237 |
215 |
72.96 |
|
|
|
| 35 |
A |
176 |
159 |
38.07 |
|
|
|
| 36 |
A |
98 |
89 |
2.88 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 27979.0 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 25360.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.060 |
|
|
|
| 2 |
H |
0.065 |
|
|
|
| 3 |
C |
0.055 |
|
|
|
| 4 |
H |
0.030 |
|
|
|
| 5 |
C |
0.336 |
|
|
|
| 6 |
H |
0.061 |
|
|
|
| 7 |
H |
0.049 |
|
|
|
| 8 |
C |
0.054 |
|
|
|
| 9 |
H |
0.198 |
|
|
|
| 10 |
O |
-0.424 |
|
|
|
| 11 |
H |
0.199 |
|
|
|
| 12 |
O |
-0.439 |
|
|
|
| 13 |
H |
0.214 |
|
|
|
| 14 |
O |
-0.460 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.072 |
-1.933 |
-0.643 |
2.038 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-29.970 |
4.023 |
-3.230 |
| y |
4.023 |
-38.426 |
-0.549 |
| z |
-3.230 |
-0.549 |
-35.693 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
7.090 |
4.023 |
-3.230 |
| y |
4.023 |
-5.595 |
-0.549 |
| z |
-3.230 |
-0.549 |
-1.495 |
|
| Polar |
|---|
| 3z2-r2 | -2.990 |
|---|
| x2-y2 | 8.457 |
|---|
| xy | 4.023 |
|---|
| xz | -3.230 |
|---|
| yz | -0.549 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.205 |
0.060 |
-0.010 |
| y |
0.060 |
6.653 |
-0.123 |
| z |
-0.010 |
-0.123 |
6.453 |
<r2> (average value of r
2) Å
2
| <r2> |
173.000 |
| (<r2>)1/2 |
13.153 |