Jump to
S1C2
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -549.193202 |
| Energy at 298.15K | |
| HF Energy | -549.193202 |
| Nuclear repulsion energy | 359.820387 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
1892 |
1715 |
0.00 |
|
|
|
| 2 |
Ag |
1571 |
1424 |
0.00 |
|
|
|
| 3 |
Ag |
712 |
645 |
0.00 |
|
|
|
| 4 |
Ag |
633 |
574 |
0.00 |
|
|
|
| 5 |
B1g |
746 |
676 |
0.00 |
|
|
|
| 6 |
B1u |
1813 |
1643 |
3.63 |
|
|
|
| 7 |
B1u |
1053 |
955 |
421.01 |
|
|
|
| 8 |
B1u |
320 |
290 |
1.92 |
|
|
|
| 9 |
B2g |
740i |
671i |
0.00 |
|
|
|
| 10 |
B2u |
1615 |
1464 |
1108.50 |
|
|
|
| 11 |
B2u |
1067 |
967 |
17.00 |
|
|
|
| 12 |
B2u |
274 |
249 |
7.89 |
|
|
|
| 13 |
B3g |
842 |
763 |
0.00 |
|
|
|
| 14 |
B3g |
244 |
221 |
0.00 |
|
|
|
| 15 |
B3g |
4848i |
4394i |
0.00 |
|
|
|
| 16 |
B3u |
734 |
665 |
131.04 |
|
|
|
| 17 |
B3u |
153 |
138 |
0.06 |
|
|
|
| 18 |
B3u |
246i |
223i |
30.17 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3917.8 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 3551.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is D2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.956 |
0.000 |
| C2 |
0.000 |
0.000 |
1.039 |
| C3 |
0.000 |
-0.956 |
0.000 |
| C4 |
0.000 |
0.000 |
-1.039 |
| F5 |
0.000 |
2.260 |
0.000 |
| F6 |
0.000 |
0.000 |
2.387 |
| F7 |
0.000 |
-2.260 |
0.000 |
| F8 |
0.000 |
0.000 |
-2.387 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
F5 |
F6 |
F7 |
F8 |
| C1 | | 1.4121 | 1.9128 | 1.4121 | 1.3034 | 2.5717 | 3.2163 | 2.5717 |
C2 | 1.4121 | | 1.4121 | 2.0777 | 2.4872 | 1.3484 | 2.4872 | 3.4261 | C3 | 1.9128 | 1.4121 | | 1.4121 | 3.2163 | 2.5717 | 1.3034 | 2.5717 | C4 | 1.4121 | 2.0777 | 1.4121 | | 2.4872 | 3.4261 | 2.4872 | 1.3484 | F5 | 1.3034 | 2.4872 | 3.2163 | 2.4872 | | 3.2872 | 4.5197 | 3.2872 | F6 | 2.5717 | 1.3484 | 2.5717 | 3.4261 | 3.2872 | | 3.2872 | 4.7745 | F7 | 3.2163 | 2.4872 | 1.3034 | 2.4872 | 4.5197 | 3.2872 | | 3.2872 | F8 | 2.5717 | 3.4261 | 2.5717 | 1.3484 | 3.2872 | 4.7745 | 3.2872 | |
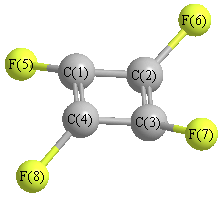 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
85.268 |
|
C1 |
C2 |
F6 |
137.366 |
| C1 |
C4 |
C3 |
85.268 |
|
C1 |
C4 |
F8 |
137.366 |
| C2 |
C1 |
C4 |
94.732 |
|
C2 |
C1 |
F5 |
132.634 |
| C2 |
C3 |
C4 |
94.732 |
|
C2 |
C3 |
F7 |
132.634 |
| C3 |
C2 |
F6 |
137.366 |
|
C3 |
C4 |
F8 |
137.366 |
| C4 |
C1 |
F5 |
132.634 |
|
C4 |
C3 |
F7 |
132.634 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.577 |
|
|
|
| 2 |
C |
-0.123 |
|
|
|
| 3 |
C |
0.577 |
|
|
|
| 4 |
C |
-0.123 |
|
|
|
| 5 |
F |
-0.189 |
|
|
|
| 6 |
F |
-0.265 |
|
|
|
| 7 |
F |
-0.189 |
|
|
|
| 8 |
F |
-0.265 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.589 |
0.000 |
0.000 |
| y |
0.000 |
-38.916 |
0.000 |
| z |
0.000 |
0.000 |
-49.985 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.862 |
0.000 |
0.000 |
| y |
0.000 |
6.371 |
0.000 |
| z |
0.000 |
0.000 |
-10.233 |
|
| Polar |
|---|
| 3z2-r2 | -20.466 |
|---|
| x2-y2 | -1.673 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.341 |
0.000 |
0.000 |
| y |
0.000 |
6.699 |
0.000 |
| z |
0.000 |
0.000 |
7.203 |
<r2> (average value of r
2) Å
2
| <r2> |
239.676 |
| (<r2>)1/2 |
15.481 |
Jump to
S1C1
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -549.252308 |
| Energy at 298.15K | -549.252600 |
| HF Energy | -549.252308 |
| Nuclear repulsion energy | 358.272968 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
2062 |
1869 |
0.00 |
|
|
|
| 2 |
Ag |
1368 |
1240 |
0.00 |
|
|
|
| 3 |
Ag |
709 |
643 |
0.00 |
|
|
|
| 4 |
Ag |
261 |
237 |
0.00 |
|
|
|
| 5 |
Ag |
123 |
111 |
0.00 |
|
|
|
| 6 |
Au |
1467 |
1330 |
570.71 |
|
|
|
| 7 |
Au |
1031 |
935 |
87.89 |
|
|
|
| 8 |
Au |
755 |
684 |
23.81 |
|
|
|
| 9 |
Au |
251 |
228 |
0.34 |
|
|
|
| 10 |
Au |
180 |
163 |
0.04 |
|
|
|
| 11 |
Bg |
1509 |
1368 |
0.00 |
|
|
|
| 12 |
Bg |
843 |
765 |
0.00 |
|
|
|
| 13 |
Bg |
580 |
525 |
0.00 |
|
|
|
| 14 |
Bg |
510 |
462 |
0.00 |
|
|
|
| 15 |
Bu |
2060 |
1867 |
142.18 |
|
|
|
| 16 |
Bu |
1073 |
972 |
227.30 |
|
|
|
| 17 |
Bu |
335 |
304 |
9.65 |
|
|
|
| 18 |
Bu |
227 |
206 |
1.66 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 7672.4 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 6954.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.125 |
0.754 |
0.650 |
| C2 |
-0.125 |
-0.754 |
0.650 |
| C3 |
-0.125 |
-0.754 |
-0.650 |
| C4 |
0.125 |
0.754 |
-0.650 |
| F5 |
-0.125 |
1.649 |
1.555 |
| F6 |
0.125 |
-1.649 |
1.555 |
| F7 |
0.125 |
-1.649 |
-1.555 |
| F8 |
-0.125 |
1.649 |
-1.555 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
F5 |
F6 |
F7 |
F8 |
| C1 | | 1.5293 | 2.0077 | 1.3009 | 1.2966 | 2.5676 | 3.2617 | 2.3932 |
C2 | 1.5293 | | 1.3009 | 2.0077 | 2.5676 | 1.2966 | 2.3932 | 3.2617 | C3 | 2.0077 | 1.3009 | | 1.5293 | 3.2617 | 2.3932 | 1.2966 | 2.5676 | C4 | 1.3009 | 2.0077 | 1.5293 | | 2.3932 | 3.2617 | 2.5676 | 1.2966 | F5 | 1.2966 | 2.5676 | 3.2617 | 2.3932 | | 3.3068 | 4.5397 | 3.1102 | F6 | 2.5676 | 1.2966 | 2.3932 | 3.2617 | 3.3068 | | 3.1102 | 4.5397 | F7 | 3.2617 | 2.3932 | 1.2966 | 2.5676 | 4.5397 | 3.1102 | | 3.3068 | F8 | 2.3932 | 3.2617 | 2.5676 | 1.2966 | 3.1102 | 4.5397 | 3.3068 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
90.000 |
|
C1 |
C2 |
F6 |
130.446 |
| C1 |
C4 |
C3 |
90.000 |
|
C1 |
C4 |
F8 |
134.246 |
| C2 |
C1 |
C4 |
90.000 |
|
C2 |
C1 |
F5 |
130.446 |
| C2 |
C3 |
C4 |
90.000 |
|
C2 |
C3 |
F7 |
134.246 |
| C3 |
C2 |
F6 |
134.246 |
|
C3 |
C4 |
F8 |
130.446 |
| C4 |
C1 |
F5 |
134.246 |
|
C4 |
C3 |
F7 |
130.446 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.240 |
|
|
|
| 2 |
C |
0.240 |
|
|
|
| 3 |
C |
0.240 |
|
|
|
| 4 |
C |
0.240 |
|
|
|
| 5 |
F |
-0.240 |
|
|
|
| 6 |
F |
-0.240 |
|
|
|
| 7 |
F |
-0.240 |
|
|
|
| 8 |
F |
-0.240 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.118 |
-0.097 |
0.000 |
| y |
-0.097 |
-44.625 |
0.000 |
| z |
0.000 |
0.000 |
-44.126 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.258 |
-0.097 |
0.000 |
| y |
-0.097 |
-2.503 |
0.000 |
| z |
0.000 |
0.000 |
-1.755 |
|
| Polar |
|---|
| 3z2-r2 | -3.510 |
|---|
| x2-y2 | 4.507 |
|---|
| xy | -0.097 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.029 |
0.081 |
0.000 |
| y |
0.081 |
5.802 |
0.000 |
| z |
0.000 |
0.000 |
7.411 |
<r2> (average value of r
2) Å
2
| <r2> |
241.318 |
| (<r2>)1/2 |
15.534 |
