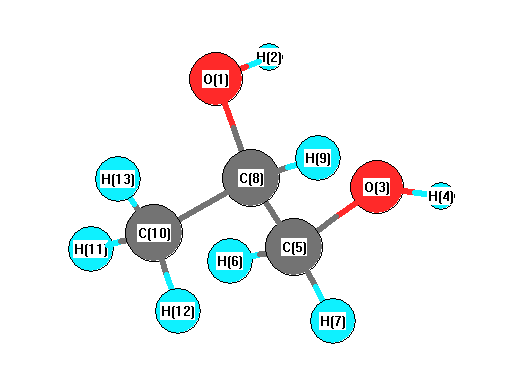
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
4172 |
3782 |
56.28 |
|
|
|
| 2 |
A |
4154 |
3765 |
72.05 |
|
|
|
| 3 |
A |
3242 |
2939 |
39.76 |
|
|
|
| 4 |
A |
3236 |
2933 |
68.63 |
|
|
|
| 5 |
A |
3224 |
2922 |
10.82 |
|
|
|
| 6 |
A |
3170 |
2873 |
57.82 |
|
|
|
| 7 |
A |
3161 |
2865 |
16.57 |
|
|
|
| 8 |
A |
3091 |
2802 |
73.20 |
|
|
|
| 9 |
A |
1625 |
1473 |
3.01 |
|
|
|
| 10 |
A |
1618 |
1466 |
1.58 |
|
|
|
| 11 |
A |
1605 |
1455 |
5.61 |
|
|
|
| 12 |
A |
1575 |
1428 |
11.88 |
|
|
|
| 13 |
A |
1544 |
1399 |
29.39 |
|
|
|
| 14 |
A |
1514 |
1373 |
4.57 |
|
|
|
| 15 |
A |
1502 |
1361 |
29.88 |
|
|
|
| 16 |
A |
1470 |
1332 |
36.88 |
|
|
|
| 17 |
A |
1390 |
1260 |
76.76 |
|
|
|
| 18 |
A |
1327 |
1203 |
21.66 |
|
|
|
| 19 |
A |
1279 |
1159 |
24.67 |
|
|
|
| 20 |
A |
1187 |
1076 |
81.86 |
|
|
|
| 21 |
A |
1172 |
1062 |
4.67 |
|
|
|
| 22 |
A |
1139 |
1032 |
104.82 |
|
|
|
| 23 |
A |
1017 |
922 |
5.64 |
|
|
|
| 24 |
A |
1001 |
907 |
18.75 |
|
|
|
| 25 |
A |
911 |
826 |
12.57 |
|
|
|
| 26 |
A |
562 |
509 |
20.05 |
|
|
|
| 27 |
A |
517 |
469 |
25.11 |
|
|
|
| 28 |
A |
455 |
413 |
141.20 |
|
|
|
| 29 |
A |
388 |
352 |
19.33 |
|
|
|
| 30 |
A |
299 |
271 |
81.54 |
|
|
|
| 31 |
A |
268 |
243 |
4.88 |
|
|
|
| 32 |
A |
239 |
217 |
0.69 |
|
|
|
| 33 |
A |
155 |
140 |
3.72 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26604.0 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 24113.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.438 |
|
|
|
| 2 |
H |
0.212 |
|
|
|
| 3 |
O |
-0.443 |
|
|
|
| 4 |
H |
0.207 |
|
|
|
| 5 |
C |
0.138 |
|
|
|
| 6 |
H |
0.050 |
|
|
|
| 7 |
H |
0.033 |
|
|
|
| 8 |
C |
0.269 |
|
|
|
| 9 |
H |
-0.010 |
|
|
|
| 10 |
C |
-0.192 |
|
|
|
| 11 |
H |
0.057 |
|
|
|
| 12 |
H |
0.042 |
|
|
|
| 13 |
H |
0.074 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.492 |
-1.735 |
1.615 |
2.421 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.624 |
1.831 |
2.845 |
| y |
1.831 |
-32.054 |
0.178 |
| z |
2.845 |
0.178 |
-29.771 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.712 |
1.831 |
2.845 |
| y |
1.831 |
-0.356 |
0.178 |
| z |
2.845 |
0.178 |
3.067 |
|
| Polar |
|---|
| 3z2-r2 | 6.135 |
|---|
| x2-y2 | -1.570 |
|---|
| xy | 1.831 |
|---|
| xz | 2.845 |
|---|
| yz | 0.178 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.914 |
0.223 |
0.136 |
| y |
0.223 |
6.258 |
-0.035 |
| z |
0.136 |
-0.035 |
5.950 |
<r2> (average value of r
2) Å
2
| <r2> |
132.033 |
| (<r2>)1/2 |
11.491 |