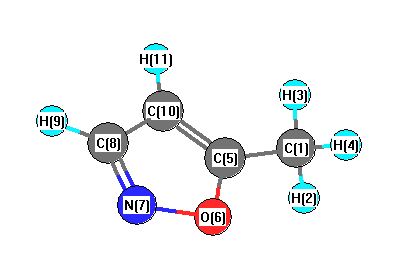
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3425 |
3105 |
0.32 |
|
|
|
| 2 |
A |
3388 |
3071 |
6.75 |
|
|
|
| 3 |
A |
3267 |
2961 |
14.73 |
|
|
|
| 4 |
A |
3238 |
2935 |
11.90 |
|
|
|
| 5 |
A |
3181 |
2883 |
15.83 |
|
|
|
| 6 |
A |
1795 |
1627 |
42.89 |
|
|
|
| 7 |
A |
1695 |
1536 |
94.66 |
|
|
|
| 8 |
A |
1605 |
1455 |
8.10 |
|
|
|
| 9 |
A |
1594 |
1444 |
6.86 |
|
|
|
| 10 |
A |
1538 |
1394 |
4.37 |
|
|
|
| 11 |
A |
1496 |
1356 |
13.00 |
|
|
|
| 12 |
A |
1373 |
1244 |
39.19 |
|
|
|
| 13 |
A |
1310 |
1188 |
13.65 |
|
|
|
| 14 |
A |
1166 |
1057 |
2.37 |
|
|
|
| 15 |
A |
1139 |
1032 |
11.69 |
|
|
|
| 16 |
A |
1108 |
1004 |
11.57 |
|
|
|
| 17 |
A |
1093 |
991 |
0.10 |
|
|
|
| 18 |
A |
1049 |
951 |
3.71 |
|
|
|
| 19 |
A |
1045 |
947 |
15.58 |
|
|
|
| 20 |
A |
1006 |
912 |
23.14 |
|
|
|
| 21 |
A |
885 |
803 |
51.05 |
|
|
|
| 22 |
A |
714 |
647 |
0.91 |
|
|
|
| 23 |
A |
702 |
636 |
0.65 |
|
|
|
| 24 |
A |
700 |
635 |
2.94 |
|
|
|
| 25 |
A |
362 |
328 |
2.55 |
|
|
|
| 26 |
A |
278 |
252 |
2.69 |
|
|
|
| 27 |
A |
140 |
127 |
0.88 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20145.5 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 18259.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.177 |
|
|
|
| 2 |
H |
0.087 |
|
|
|
| 3 |
H |
0.078 |
|
|
|
| 4 |
H |
0.087 |
|
|
|
| 5 |
C |
0.271 |
|
|
|
| 6 |
O |
-0.232 |
|
|
|
| 7 |
N |
-0.190 |
|
|
|
| 8 |
C |
0.102 |
|
|
|
| 9 |
H |
0.100 |
|
|
|
| 10 |
C |
-0.222 |
|
|
|
| 11 |
H |
0.097 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.927 |
2.764 |
-0.001 |
3.370 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.361 |
-4.022 |
0.001 |
| y |
-4.022 |
-35.995 |
0.000 |
| z |
0.001 |
0.000 |
-36.396 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.835 |
-4.022 |
0.001 |
| y |
-4.022 |
-2.116 |
0.000 |
| z |
0.001 |
0.000 |
-2.718 |
|
| Polar |
|---|
| 3z2-r2 | -5.436 |
|---|
| x2-y2 | 4.634 |
|---|
| xy | -4.022 |
|---|
| xz | 0.001 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.331 |
-0.439 |
0.000 |
| y |
-0.439 |
7.730 |
-0.000 |
| z |
0.000 |
-0.000 |
5.286 |
<r2> (average value of r
2) Å
2
| <r2> |
133.250 |
| (<r2>)1/2 |
11.543 |