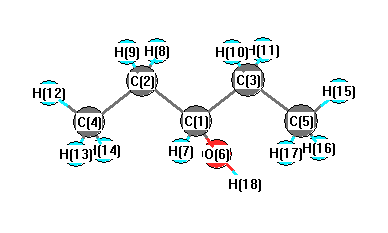
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
4194 |
3801 |
46.97 |
|
|
|
| 2 |
A |
3244 |
2941 |
45.82 |
|
|
|
| 3 |
A |
3218 |
2917 |
60.16 |
|
|
|
| 4 |
A |
3214 |
2913 |
122.33 |
|
|
|
| 5 |
A |
3209 |
2909 |
45.49 |
|
|
|
| 6 |
A |
3192 |
2893 |
12.77 |
|
|
|
| 7 |
A |
3181 |
2883 |
4.48 |
|
|
|
| 8 |
A |
3164 |
2868 |
38.57 |
|
|
|
| 9 |
A |
3157 |
2862 |
35.60 |
|
|
|
| 10 |
A |
3154 |
2858 |
22.03 |
|
|
|
| 11 |
A |
3151 |
2856 |
37.72 |
|
|
|
| 12 |
A |
3104 |
2813 |
40.64 |
|
|
|
| 13 |
A |
1627 |
1475 |
6.23 |
|
|
|
| 14 |
A |
1625 |
1473 |
5.32 |
|
|
|
| 15 |
A |
1616 |
1465 |
6.82 |
|
|
|
| 16 |
A |
1612 |
1462 |
5.21 |
|
|
|
| 17 |
A |
1605 |
1454 |
1.06 |
|
|
|
| 18 |
A |
1596 |
1446 |
0.21 |
|
|
|
| 19 |
A |
1565 |
1419 |
6.15 |
|
|
|
| 20 |
A |
1541 |
1396 |
1.12 |
|
|
|
| 21 |
A |
1532 |
1388 |
3.01 |
|
|
|
| 22 |
A |
1527 |
1384 |
12.63 |
|
|
|
| 23 |
A |
1493 |
1354 |
0.14 |
|
|
|
| 24 |
A |
1437 |
1303 |
20.24 |
|
|
|
| 25 |
A |
1425 |
1292 |
2.75 |
|
|
|
| 26 |
A |
1389 |
1259 |
7.12 |
|
|
|
| 27 |
A |
1357 |
1230 |
48.20 |
|
|
|
| 28 |
A |
1272 |
1153 |
29.48 |
|
|
|
| 29 |
A |
1252 |
1135 |
5.16 |
|
|
|
| 30 |
A |
1174 |
1064 |
15.49 |
|
|
|
| 31 |
A |
1136 |
1030 |
31.61 |
|
|
|
| 32 |
A |
1121 |
1016 |
7.31 |
|
|
|
| 33 |
A |
1092 |
990 |
0.02 |
|
|
|
| 34 |
A |
1056 |
957 |
33.82 |
|
|
|
| 35 |
A |
983 |
891 |
6.90 |
|
|
|
| 36 |
A |
922 |
836 |
3.22 |
|
|
|
| 37 |
A |
838 |
760 |
1.48 |
|
|
|
| 38 |
A |
829 |
752 |
0.13 |
|
|
|
| 39 |
A |
533 |
483 |
4.22 |
|
|
|
| 40 |
A |
512 |
464 |
8.37 |
|
|
|
| 41 |
A |
420 |
381 |
1.94 |
|
|
|
| 42 |
A |
329 |
298 |
2.23 |
|
|
|
| 43 |
A |
294 |
267 |
79.24 |
|
|
|
| 44 |
A |
252 |
229 |
47.96 |
|
|
|
| 45 |
A |
233 |
211 |
2.62 |
|
|
|
| 46 |
A |
204 |
185 |
0.25 |
|
|
|
| 47 |
A |
113 |
103 |
1.60 |
|
|
|
| 48 |
A |
101 |
91 |
0.12 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 38397.0 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 34803.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.256 |
|
|
|
| 2 |
C |
-0.044 |
|
|
|
| 3 |
C |
-0.051 |
|
|
|
| 4 |
C |
-0.195 |
|
|
|
| 5 |
C |
-0.211 |
|
|
|
| 6 |
O |
-0.434 |
|
|
|
| 7 |
H |
0.019 |
|
|
|
| 8 |
H |
0.028 |
|
|
|
| 9 |
H |
0.037 |
|
|
|
| 10 |
H |
0.039 |
|
|
|
| 11 |
H |
0.028 |
|
|
|
| 12 |
H |
0.057 |
|
|
|
| 13 |
H |
0.038 |
|
|
|
| 14 |
H |
0.075 |
|
|
|
| 15 |
H |
0.065 |
|
|
|
| 16 |
H |
0.044 |
|
|
|
| 17 |
H |
0.054 |
|
|
|
| 18 |
H |
0.193 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.119 |
-0.575 |
0.981 |
1.596 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.033 |
2.125 |
0.211 |
| y |
2.125 |
-39.479 |
1.531 |
| z |
0.211 |
1.531 |
-40.370 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.108 |
2.125 |
0.211 |
| y |
2.125 |
0.722 |
1.531 |
| z |
0.211 |
1.531 |
-0.614 |
|
| Polar |
|---|
| 3z2-r2 | -1.228 |
|---|
| x2-y2 | -0.553 |
|---|
| xy | 2.125 |
|---|
| xz | 0.211 |
|---|
| yz | 1.531 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.670 |
0.089 |
0.040 |
| y |
0.089 |
8.759 |
0.108 |
| z |
0.040 |
0.108 |
8.353 |
<r2> (average value of r
2) Å
2
| <r2> |
224.469 |
| (<r2>)1/2 |
14.982 |