Jump to
S1C2
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -615.100240 |
| Energy at 298.15K | -615.107606 |
| HF Energy | -615.100240 |
| Nuclear repulsion energy | 206.189001 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3322 |
3011 |
7.59 |
|
|
|
| 2 |
A' |
3268 |
2962 |
23.31 |
|
|
|
| 3 |
A' |
3231 |
2929 |
23.29 |
|
|
|
| 4 |
A' |
3220 |
2918 |
22.81 |
|
|
|
| 5 |
A' |
3157 |
2861 |
38.35 |
|
|
|
| 6 |
A' |
1889 |
1712 |
1.62 |
|
|
|
| 7 |
A' |
1609 |
1458 |
11.66 |
|
|
|
| 8 |
A' |
1604 |
1454 |
2.00 |
|
|
|
| 9 |
A' |
1540 |
1396 |
3.28 |
|
|
|
| 10 |
A' |
1456 |
1320 |
30.14 |
|
|
|
| 11 |
A' |
1442 |
1307 |
13.81 |
|
|
|
| 12 |
A' |
1414 |
1281 |
4.74 |
|
|
|
| 13 |
A' |
1199 |
1087 |
0.55 |
|
|
|
| 14 |
A' |
1108 |
1004 |
12.18 |
|
|
|
| 15 |
A' |
975 |
884 |
9.14 |
|
|
|
| 16 |
A' |
758 |
687 |
26.32 |
|
|
|
| 17 |
A' |
625 |
566 |
7.86 |
|
|
|
| 18 |
A' |
358 |
324 |
1.67 |
|
|
|
| 19 |
A' |
184 |
167 |
0.78 |
|
|
|
| 20 |
A" |
3264 |
2958 |
7.08 |
|
|
|
| 21 |
A" |
3201 |
2901 |
29.08 |
|
|
|
| 22 |
A" |
1597 |
1448 |
6.22 |
|
|
|
| 23 |
A" |
1310 |
1187 |
3.00 |
|
|
|
| 24 |
A" |
1180 |
1070 |
4.25 |
|
|
|
| 25 |
A" |
1096 |
994 |
40.26 |
|
|
|
| 26 |
A" |
1046 |
948 |
0.17 |
|
|
|
| 27 |
A" |
814 |
738 |
0.50 |
|
|
|
| 28 |
A" |
260 |
236 |
4.89 |
|
|
|
| 29 |
A" |
221 |
200 |
1.63 |
|
|
|
| 30 |
A" |
121 |
110 |
1.33 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23233.3 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 21058.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.308 |
0.275 |
0.000 |
| C2 |
0.000 |
1.004 |
0.000 |
| C3 |
1.211 |
0.497 |
0.000 |
| C4 |
2.469 |
1.312 |
0.000 |
| Cl5 |
-1.181 |
-1.508 |
0.000 |
| H6 |
-1.886 |
0.532 |
0.874 |
| H7 |
-1.886 |
0.532 |
-0.874 |
| H8 |
-0.126 |
2.075 |
0.000 |
| H9 |
1.334 |
-0.570 |
0.000 |
| H10 |
2.260 |
2.373 |
0.000 |
| H11 |
3.072 |
1.082 |
0.872 |
| H12 |
3.072 |
1.082 |
-0.872 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
Cl5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.4979 | 2.5289 | 3.9173 | 1.7875 | 1.0791 | 1.0791 | 2.1540 | 2.7736 | 4.1397 | 4.5385 | 4.5385 |
C2 | 1.4979 | | 1.3126 | 2.4884 | 2.7760 | 2.1314 | 2.1314 | 1.0786 | 2.0631 | 2.6425 | 3.1942 | 3.1942 | C3 | 2.5289 | 1.3126 | | 1.4990 | 3.1211 | 3.2177 | 3.2177 | 2.0683 | 1.0743 | 2.1495 | 2.1370 | 2.1370 | C4 | 3.9173 | 2.4884 | 1.4990 | | 4.6123 | 4.5098 | 4.5098 | 2.7056 | 2.1976 | 1.0822 | 1.0847 | 1.0847 | Cl5 | 1.7875 | 2.7760 | 3.1211 | 4.6123 | | 2.3292 | 2.3292 | 3.7355 | 2.6837 | 5.1870 | 5.0552 | 5.0552 | H6 | 1.0791 | 2.1314 | 3.2177 | 4.5098 | 2.3292 | | 1.7488 | 2.4983 | 3.5133 | 4.6197 | 4.9879 | 5.2849 | H7 | 1.0791 | 2.1314 | 3.2177 | 4.5098 | 2.3292 | 1.7488 | | 2.4983 | 3.5133 | 4.6197 | 5.2849 | 4.9879 | H8 | 2.1540 | 1.0786 | 2.0683 | 2.7056 | 3.7355 | 2.4983 | 2.4983 | | 3.0214 | 2.4048 | 3.4604 | 3.4604 | H9 | 2.7736 | 2.0631 | 1.0743 | 2.1976 | 2.6837 | 3.5133 | 3.5133 | 3.0214 | | 3.0856 | 2.5519 | 2.5519 | H10 | 4.1397 | 2.6425 | 2.1495 | 1.0822 | 5.1870 | 4.6197 | 4.6197 | 2.4048 | 3.0856 | | 1.7568 | 1.7568 | H11 | 4.5385 | 3.1942 | 2.1370 | 1.0847 | 5.0552 | 4.9879 | 5.2849 | 3.4604 | 2.5519 | 1.7568 | | 1.7450 | H12 | 4.5385 | 3.1942 | 2.1370 | 1.0847 | 5.0552 | 5.2849 | 4.9879 | 3.4604 | 2.5519 | 1.7568 | 1.7450 | |
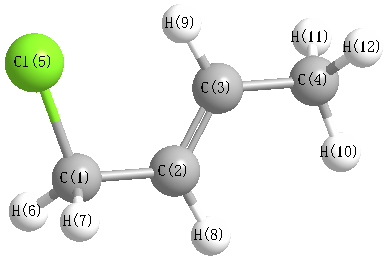 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
128.142 |
|
C1 |
C2 |
H8 |
112.425 |
| C2 |
C1 |
Cl5 |
115.049 |
|
C2 |
C1 |
H6 |
110.555 |
| C2 |
C1 |
H7 |
110.555 |
|
C2 |
C3 |
C4 |
124.387 |
| C2 |
C3 |
H9 |
119.282 |
|
C3 |
C2 |
H8 |
119.433 |
| C3 |
C4 |
H10 |
111.750 |
|
C3 |
C4 |
H11 |
110.584 |
| C3 |
C4 |
H12 |
110.584 |
|
C4 |
C3 |
H9 |
116.331 |
| Cl5 |
C1 |
H6 |
106.040 |
|
Cl5 |
C1 |
H7 |
106.040 |
| H6 |
C1 |
H7 |
108.261 |
|
H10 |
C4 |
H11 |
108.336 |
| H10 |
C4 |
H12 |
108.336 |
|
H11 |
C4 |
H12 |
107.095 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.086 |
|
|
|
| 2 |
C |
-0.188 |
|
|
|
| 3 |
C |
-0.046 |
|
|
|
| 4 |
C |
-0.124 |
|
|
|
| 5 |
Cl |
-0.186 |
|
|
|
| 6 |
H |
0.081 |
|
|
|
| 7 |
H |
0.081 |
|
|
|
| 8 |
H |
0.051 |
|
|
|
| 9 |
H |
0.066 |
|
|
|
| 10 |
H |
0.056 |
|
|
|
| 11 |
H |
0.063 |
|
|
|
| 12 |
H |
0.063 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.330 |
2.061 |
0.000 |
2.087 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.981 |
-2.832 |
0.000 |
| y |
-2.832 |
-39.148 |
0.000 |
| z |
0.000 |
0.000 |
-39.799 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.492 |
-2.832 |
0.000 |
| y |
-2.832 |
-1.258 |
0.000 |
| z |
0.000 |
0.000 |
-2.234 |
|
| Polar |
|---|
| 3z2-r2 | -4.468 |
|---|
| x2-y2 | 3.167 |
|---|
| xy | -2.832 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.608 |
0.400 |
0.000 |
| y |
0.400 |
9.059 |
0.000 |
| z |
0.000 |
0.000 |
6.353 |
<r2> (average value of r
2) Å
2
| <r2> |
209.767 |
| (<r2>)1/2 |
14.483 |
Jump to
S1C1
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -615.102765 |
| Energy at 298.15K | -615.110100 |
| HF Energy | -615.102765 |
| Nuclear repulsion energy | 202.186095 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3308 |
2998 |
20.52 |
|
|
|
| 2 |
A |
3292 |
2984 |
1.53 |
|
|
|
| 3 |
A |
3277 |
2971 |
22.22 |
|
|
|
| 4 |
A |
3239 |
2935 |
17.59 |
|
|
|
| 5 |
A |
3235 |
2932 |
18.75 |
|
|
|
| 6 |
A |
3202 |
2902 |
27.42 |
|
|
|
| 7 |
A |
3158 |
2862 |
36.16 |
|
|
|
| 8 |
A |
1878 |
1702 |
9.93 |
|
|
|
| 9 |
A |
1612 |
1462 |
8.31 |
|
|
|
| 10 |
A |
1606 |
1456 |
7.37 |
|
|
|
| 11 |
A |
1597 |
1448 |
6.82 |
|
|
|
| 12 |
A |
1540 |
1396 |
2.91 |
|
|
|
| 13 |
A |
1460 |
1324 |
10.42 |
|
|
|
| 14 |
A |
1442 |
1307 |
1.09 |
|
|
|
| 15 |
A |
1397 |
1266 |
52.04 |
|
|
|
| 16 |
A |
1300 |
1179 |
1.25 |
|
|
|
| 17 |
A |
1209 |
1096 |
0.98 |
|
|
|
| 18 |
A |
1178 |
1067 |
3.66 |
|
|
|
| 19 |
A |
1148 |
1040 |
2.34 |
|
|
|
| 20 |
A |
1099 |
996 |
44.38 |
|
|
|
| 21 |
A |
1029 |
933 |
5.67 |
|
|
|
| 22 |
A |
966 |
876 |
2.22 |
|
|
|
| 23 |
A |
877 |
795 |
12.50 |
|
|
|
| 24 |
A |
746 |
676 |
74.45 |
|
|
|
| 25 |
A |
526 |
476 |
1.64 |
|
|
|
| 26 |
A |
382 |
346 |
3.74 |
|
|
|
| 27 |
A |
297 |
269 |
2.50 |
|
|
|
| 28 |
A |
227 |
205 |
0.81 |
|
|
|
| 29 |
A |
171 |
155 |
2.30 |
|
|
|
| 30 |
A |
97 |
87 |
0.42 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23246.8 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 21070.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.773 |
0.816 |
0.126 |
| C2 |
0.521 |
0.150 |
0.442 |
| C3 |
1.593 |
0.252 |
-0.313 |
| C4 |
2.917 |
-0.382 |
-0.017 |
| Cl5 |
-2.094 |
-0.390 |
-0.109 |
| H6 |
-1.109 |
1.448 |
0.932 |
| H7 |
-0.720 |
1.391 |
-0.783 |
| H8 |
0.547 |
-0.430 |
1.348 |
| H9 |
1.536 |
0.829 |
-1.222 |
| H10 |
2.890 |
-0.946 |
0.906 |
| H11 |
3.206 |
-1.052 |
-0.819 |
| H12 |
3.693 |
0.372 |
0.065 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
Cl5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.4901 | 2.4720 | 3.8831 | 1.8039 | 1.0776 | 1.0762 | 2.1888 | 2.6738 | 4.1391 | 4.4971 | 4.4890 |
C2 | 1.4901 | | 1.3150 | 2.4970 | 2.7268 | 2.1400 | 2.1402 | 1.0758 | 2.0636 | 2.6505 | 3.2011 | 3.2016 | C3 | 2.4720 | 1.3150 | | 1.4978 | 3.7478 | 3.2060 | 2.6205 | 2.0777 | 1.0777 | 2.1452 | 2.1355 | 2.1374 | C4 | 3.8831 | 2.4970 | 1.4978 | | 5.0121 | 4.5228 | 4.1182 | 2.7354 | 2.1966 | 1.0818 | 1.0847 | 1.0847 | Cl5 | 1.8039 | 2.7268 | 3.7478 | 5.0121 | | 2.3307 | 2.3482 | 3.0163 | 3.9873 | 5.1160 | 5.3885 | 5.8395 | H6 | 1.0776 | 2.1400 | 3.2060 | 4.5228 | 2.3307 | | 1.7593 | 2.5377 | 3.4664 | 4.6599 | 5.2854 | 4.9964 | H7 | 1.0762 | 2.1402 | 2.6205 | 4.1182 | 2.3482 | 1.7593 | | 3.0758 | 2.3660 | 4.6196 | 4.6246 | 4.6079 | H8 | 2.1888 | 1.0758 | 2.0777 | 2.7354 | 3.0163 | 2.5377 | 3.0758 | | 3.0272 | 2.4389 | 3.4862 | 3.4904 | H9 | 2.6738 | 2.0636 | 1.0777 | 2.1966 | 3.9873 | 3.4664 | 2.3660 | 3.0272 | | 3.0834 | 2.5476 | 2.5534 | H10 | 4.1391 | 2.6505 | 2.1452 | 1.0818 | 5.1160 | 4.6599 | 4.6196 | 2.4389 | 3.0834 | | 1.7570 | 1.7571 | H11 | 4.4971 | 3.2011 | 2.1355 | 1.0847 | 5.3885 | 5.2854 | 4.6246 | 3.4862 | 2.5476 | 1.7570 | | 1.7459 | H12 | 4.4890 | 3.2016 | 2.1374 | 1.0847 | 5.8395 | 4.9964 | 4.6079 | 3.4904 | 2.5534 | 1.7571 | 1.7459 | |
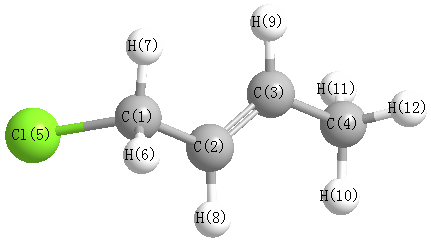 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
123.473 |
|
C1 |
C2 |
H8 |
116.154 |
| C2 |
C1 |
Cl5 |
111.392 |
|
C2 |
C1 |
H6 |
111.903 |
| C2 |
C1 |
H7 |
112.007 |
|
C2 |
C3 |
C4 |
125.052 |
| C2 |
C3 |
H9 |
118.853 |
|
C3 |
C2 |
H8 |
120.372 |
| C3 |
C4 |
H10 |
111.500 |
|
C3 |
C4 |
H11 |
110.547 |
| C3 |
C4 |
H12 |
110.697 |
|
C4 |
C3 |
H9 |
116.094 |
| Cl5 |
C1 |
H6 |
105.163 |
|
Cl5 |
C1 |
H7 |
106.485 |
| H6 |
C1 |
H7 |
109.534 |
|
H10 |
C4 |
H11 |
108.382 |
| H10 |
C4 |
H12 |
108.395 |
|
H11 |
C4 |
H12 |
107.179 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.053 |
|
|
|
| 2 |
C |
-0.121 |
|
|
|
| 3 |
C |
-0.064 |
|
|
|
| 4 |
C |
-0.130 |
|
|
|
| 5 |
Cl |
-0.201 |
|
|
|
| 6 |
H |
0.082 |
|
|
|
| 7 |
H |
0.082 |
|
|
|
| 8 |
H |
0.058 |
|
|
|
| 9 |
H |
0.052 |
|
|
|
| 10 |
H |
0.061 |
|
|
|
| 11 |
H |
0.065 |
|
|
|
| 12 |
H |
0.062 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.222 |
1.276 |
0.171 |
2.568 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.612 |
-1.986 |
-1.173 |
| y |
-1.986 |
-38.088 |
-1.008 |
| z |
-1.173 |
-1.008 |
-37.569 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.784 |
-1.986 |
-1.173 |
| y |
-1.986 |
1.003 |
-1.008 |
| z |
-1.173 |
-1.008 |
1.781 |
|
| Polar |
|---|
| 3z2-r2 | 3.561 |
|---|
| x2-y2 | -2.525 |
|---|
| xy | -1.986 |
|---|
| xz | -1.173 |
|---|
| yz | -1.008 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.127 |
0.415 |
-0.827 |
| y |
0.415 |
7.315 |
-0.142 |
| z |
-0.827 |
-0.142 |
7.294 |
<r2> (average value of r
2) Å
2
| <r2> |
230.662 |
| (<r2>)1/2 |
15.188 |
