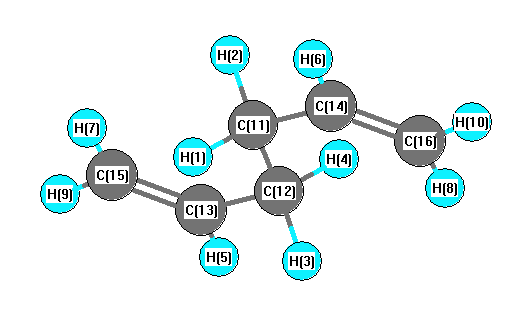
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3357 |
3043 |
10.28 |
|
|
|
| 2 |
A |
3281 |
2974 |
0.06 |
|
|
|
| 3 |
A |
3268 |
2962 |
45.72 |
|
|
|
| 4 |
A |
3212 |
2911 |
30.08 |
|
|
|
| 5 |
A |
3146 |
2852 |
14.42 |
|
|
|
| 6 |
A |
1841 |
1669 |
7.59 |
|
|
|
| 7 |
A |
1604 |
1454 |
4.82 |
|
|
|
| 8 |
A |
1574 |
1427 |
3.06 |
|
|
|
| 9 |
A |
1497 |
1357 |
0.38 |
|
|
|
| 10 |
A |
1429 |
1295 |
0.36 |
|
|
|
| 11 |
A |
1344 |
1219 |
0.00 |
|
|
|
| 12 |
A |
1180 |
1069 |
0.76 |
|
|
|
| 13 |
A |
1127 |
1021 |
5.18 |
|
|
|
| 14 |
A |
1099 |
997 |
2.13 |
|
|
|
| 15 |
A |
1069 |
969 |
28.37 |
|
|
|
| 16 |
A |
1020 |
924 |
1.40 |
|
|
|
| 17 |
A |
843 |
764 |
0.13 |
|
|
|
| 18 |
A |
683 |
619 |
7.94 |
|
|
|
| 19 |
A |
446 |
405 |
0.01 |
|
|
|
| 20 |
A |
375 |
340 |
0.05 |
|
|
|
| 21 |
A |
103 |
94 |
0.04 |
|
|
|
| 22 |
A |
82 |
75 |
0.04 |
|
|
|
| 23 |
B |
3357 |
3043 |
32.49 |
|
|
|
| 24 |
B |
3281 |
2974 |
9.51 |
|
|
|
| 25 |
B |
3267 |
2961 |
12.30 |
|
|
|
| 26 |
B |
3193 |
2894 |
12.26 |
|
|
|
| 27 |
B |
3146 |
2851 |
43.48 |
|
|
|
| 28 |
B |
1844 |
1672 |
25.04 |
|
|
|
| 29 |
B |
1617 |
1466 |
8.34 |
|
|
|
| 30 |
B |
1574 |
1427 |
0.51 |
|
|
|
| 31 |
B |
1437 |
1303 |
0.15 |
|
|
|
| 32 |
B |
1416 |
1283 |
0.11 |
|
|
|
| 33 |
B |
1386 |
1256 |
8.72 |
|
|
|
| 34 |
B |
1274 |
1155 |
1.01 |
|
|
|
| 35 |
B |
1137 |
1030 |
16.88 |
|
|
|
| 36 |
B |
1070 |
969 |
86.21 |
|
|
|
| 37 |
B |
1038 |
941 |
8.32 |
|
|
|
| 38 |
B |
988 |
895 |
0.33 |
|
|
|
| 39 |
B |
723 |
655 |
20.77 |
|
|
|
| 40 |
B |
470 |
426 |
2.35 |
|
|
|
| 41 |
B |
232 |
211 |
0.28 |
|
|
|
| 42 |
B |
123 |
111 |
0.11 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 33075.7 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 29979.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.014 |
|
|
|
| 2 |
C |
0.014 |
|
|
|
| 3 |
C |
-0.142 |
|
|
|
| 4 |
C |
-0.142 |
|
|
|
| 5 |
C |
-0.159 |
|
|
|
| 6 |
C |
-0.159 |
|
|
|
| 7 |
H |
0.046 |
|
|
|
| 8 |
H |
0.053 |
|
|
|
| 9 |
H |
0.046 |
|
|
|
| 10 |
H |
0.053 |
|
|
|
| 11 |
H |
0.064 |
|
|
|
| 12 |
H |
0.064 |
|
|
|
| 13 |
H |
0.071 |
|
|
|
| 14 |
H |
0.053 |
|
|
|
| 15 |
H |
0.071 |
|
|
|
| 16 |
H |
0.053 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.268 |
0.268 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.025 |
1.543 |
0.000 |
| y |
1.543 |
-40.378 |
0.000 |
| z |
0.000 |
0.000 |
-37.463 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.104 |
1.543 |
0.000 |
| y |
1.543 |
-1.635 |
0.000 |
| z |
0.000 |
0.000 |
2.739 |
|
| Polar |
|---|
| 3z2-r2 | 5.478 |
|---|
| x2-y2 | 0.354 |
|---|
| xy | 1.543 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.276 |
0.963 |
0.000 |
| y |
0.963 |
13.606 |
0.000 |
| z |
0.000 |
0.000 |
9.908 |
<r2> (average value of r
2) Å
2
| <r2> |
253.296 |
| (<r2>)1/2 |
15.915 |