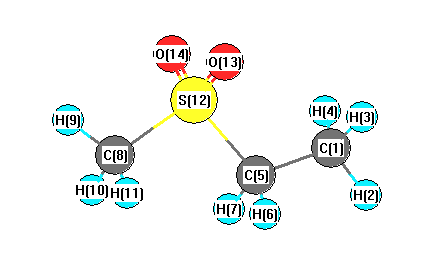
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3302 |
2993 |
4.05 |
|
|
|
| 2 |
A |
3288 |
2981 |
2.67 |
|
|
|
| 3 |
A |
3268 |
2962 |
16.82 |
|
|
|
| 4 |
A |
3248 |
2944 |
5.54 |
|
|
|
| 5 |
A |
3234 |
2931 |
23.99 |
|
|
|
| 6 |
A |
3202 |
2902 |
3.20 |
|
|
|
| 7 |
A |
3198 |
2899 |
2.73 |
|
|
|
| 8 |
A |
3180 |
2882 |
22.30 |
|
|
|
| 9 |
A |
1622 |
1470 |
6.67 |
|
|
|
| 10 |
A |
1616 |
1465 |
6.92 |
|
|
|
| 11 |
A |
1584 |
1435 |
5.35 |
|
|
|
| 12 |
A |
1579 |
1431 |
2.46 |
|
|
|
| 13 |
A |
1572 |
1425 |
2.16 |
|
|
|
| 14 |
A |
1542 |
1398 |
2.09 |
|
|
|
| 15 |
A |
1492 |
1352 |
59.77 |
|
|
|
| 16 |
A |
1484 |
1345 |
239.36 |
|
|
|
| 17 |
A |
1447 |
1311 |
34.02 |
|
|
|
| 18 |
A |
1375 |
1246 |
8.16 |
|
|
|
| 19 |
A |
1282 |
1162 |
206.34 |
|
|
|
| 20 |
A |
1164 |
1055 |
1.42 |
|
|
|
| 21 |
A |
1159 |
1050 |
5.55 |
|
|
|
| 22 |
A |
1068 |
968 |
36.07 |
|
|
|
| 23 |
A |
1058 |
959 |
4.27 |
|
|
|
| 24 |
A |
1042 |
944 |
16.42 |
|
|
|
| 25 |
A |
872 |
790 |
62.78 |
|
|
|
| 26 |
A |
799 |
725 |
11.08 |
|
|
|
| 27 |
A |
731 |
662 |
14.10 |
|
|
|
| 28 |
A |
568 |
515 |
34.26 |
|
|
|
| 29 |
A |
504 |
457 |
49.09 |
|
|
|
| 30 |
A |
444 |
403 |
5.95 |
|
|
|
| 31 |
A |
354 |
321 |
0.73 |
|
|
|
| 32 |
A |
311 |
282 |
2.21 |
|
|
|
| 33 |
A |
242 |
220 |
1.81 |
|
|
|
| 34 |
A |
229 |
207 |
0.91 |
|
|
|
| 35 |
A |
209 |
189 |
0.56 |
|
|
|
| 36 |
A |
94 |
85 |
2.78 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26680.4 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 24183.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.200 |
|
|
|
| 2 |
H |
0.099 |
|
|
|
| 3 |
H |
0.073 |
|
|
|
| 4 |
H |
0.051 |
|
|
|
| 5 |
C |
-0.136 |
|
|
|
| 6 |
H |
0.092 |
|
|
|
| 7 |
H |
0.077 |
|
|
|
| 8 |
C |
-0.277 |
|
|
|
| 9 |
H |
0.098 |
|
|
|
| 10 |
H |
0.115 |
|
|
|
| 11 |
H |
0.100 |
|
|
|
| 12 |
S |
1.130 |
|
|
|
| 13 |
O |
-0.611 |
|
|
|
| 14 |
O |
-0.612 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.310 |
3.046 |
-2.179 |
4.998 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-44.241 |
-5.512 |
-1.438 |
| y |
-5.512 |
-41.746 |
-0.607 |
| z |
-1.438 |
-0.607 |
-48.613 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.938 |
-5.512 |
-1.438 |
| y |
-5.512 |
4.681 |
-0.607 |
| z |
-1.438 |
-0.607 |
-5.619 |
|
| Polar |
|---|
| 3z2-r2 | -11.239 |
|---|
| x2-y2 | -2.496 |
|---|
| xy | -5.512 |
|---|
| xz | -1.438 |
|---|
| yz | -0.607 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.922 |
-0.173 |
0.086 |
| y |
-0.173 |
8.108 |
0.063 |
| z |
0.086 |
0.063 |
7.712 |
<r2> (average value of r
2) Å
2
| <r2> |
187.380 |
| (<r2>)1/2 |
13.689 |