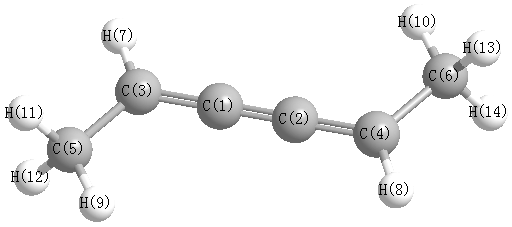
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3284 |
2976 |
0.00 |
|
|
|
| 2 |
Ag |
3249 |
2945 |
0.00 |
|
|
|
| 3 |
Ag |
3156 |
2861 |
0.00 |
|
|
|
| 4 |
Ag |
2348 |
2129 |
0.00 |
|
|
|
| 5 |
Ag |
1611 |
1460 |
0.00 |
|
|
|
| 6 |
Ag |
1534 |
1391 |
0.00 |
|
|
|
| 7 |
Ag |
1498 |
1358 |
0.00 |
|
|
|
| 8 |
Ag |
1192 |
1080 |
0.00 |
|
|
|
| 9 |
Ag |
1150 |
1042 |
0.00 |
|
|
|
| 10 |
Ag |
815 |
738 |
0.00 |
|
|
|
| 11 |
Ag |
582 |
527 |
0.00 |
|
|
|
| 12 |
Ag |
225 |
204 |
0.00 |
|
|
|
| 13 |
Au |
3199 |
2900 |
55.82 |
|
|
|
| 14 |
Au |
1598 |
1449 |
11.42 |
|
|
|
| 15 |
Au |
1160 |
1052 |
4.93 |
|
|
|
| 16 |
Au |
953 |
864 |
42.17 |
|
|
|
| 17 |
Au |
358 |
325 |
0.63 |
|
|
|
| 18 |
Au |
177 |
161 |
0.13 |
|
|
|
| 19 |
Au |
123 |
112 |
4.77 |
|
|
|
| 20 |
Bg |
3199 |
2900 |
0.00 |
|
|
|
| 21 |
Bg |
1598 |
1449 |
0.00 |
|
|
|
| 22 |
Bg |
1159 |
1051 |
0.00 |
|
|
|
| 23 |
Bg |
799 |
724 |
0.00 |
|
|
|
| 24 |
Bg |
586 |
531 |
0.00 |
|
|
|
| 25 |
Bg |
179 |
162 |
0.00 |
|
|
|
| 26 |
Bu |
3284 |
2977 |
35.81 |
|
|
|
| 27 |
Bu |
3249 |
2945 |
21.23 |
|
|
|
| 28 |
Bu |
3156 |
2860 |
115.97 |
|
|
|
| 29 |
Bu |
1851 |
1678 |
4.03 |
|
|
|
| 30 |
Bu |
1601 |
1451 |
16.51 |
|
|
|
| 31 |
Bu |
1530 |
1387 |
25.01 |
|
|
|
| 32 |
Bu |
1392 |
1262 |
30.75 |
|
|
|
| 33 |
Bu |
1197 |
1085 |
4.95 |
|
|
|
| 34 |
Bu |
990 |
897 |
31.15 |
|
|
|
| 35 |
Bu |
573 |
519 |
12.67 |
|
|
|
| 36 |
Bu |
117 |
106 |
2.59 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 27336.7 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 24778.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.066 |
|
|
|
| 2 |
C |
-0.066 |
|
|
|
| 3 |
C |
-0.079 |
|
|
|
| 4 |
C |
-0.079 |
|
|
|
| 5 |
C |
-0.126 |
|
|
|
| 6 |
C |
-0.126 |
|
|
|
| 7 |
H |
0.070 |
|
|
|
| 8 |
H |
0.070 |
|
|
|
| 9 |
H |
0.074 |
|
|
|
| 10 |
H |
0.074 |
|
|
|
| 11 |
H |
0.063 |
|
|
|
| 12 |
H |
0.063 |
|
|
|
| 13 |
H |
0.063 |
|
|
|
| 14 |
H |
0.063 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.513 |
-0.634 |
0.000 |
| y |
-0.634 |
-29.643 |
0.000 |
| z |
0.000 |
0.000 |
-39.705 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.839 |
-0.634 |
0.000 |
| y |
-0.634 |
8.466 |
0.000 |
| z |
0.000 |
0.000 |
-6.627 |
|
| Polar |
|---|
| 3z2-r2 | -13.254 |
|---|
| x2-y2 | -6.870 |
|---|
| xy | -0.634 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.250 |
-2.026 |
0.000 |
| y |
-2.026 |
22.500 |
0.000 |
| z |
0.000 |
0.000 |
7.130 |
<r2> (average value of r
2) Å
2
| <r2> |
271.794 |
| (<r2>)1/2 |
16.486 |