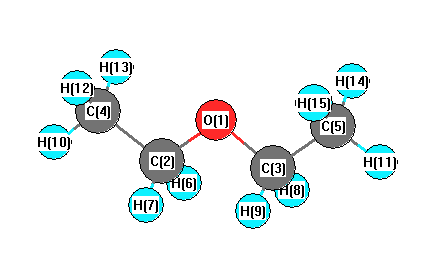
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3227 |
2925 |
66.05 |
|
|
|
| 2 |
A1 |
3167 |
2871 |
3.28 |
|
|
|
| 3 |
A1 |
3122 |
2830 |
122.52 |
|
|
|
| 4 |
A1 |
1661 |
1505 |
1.40 |
|
|
|
| 5 |
A1 |
1618 |
1467 |
5.47 |
|
|
|
| 6 |
A1 |
1588 |
1439 |
3.39 |
|
|
|
| 7 |
A1 |
1525 |
1383 |
0.27 |
|
|
|
| 8 |
A1 |
1288 |
1167 |
31.28 |
|
|
|
| 9 |
A1 |
1133 |
1027 |
10.09 |
|
|
|
| 10 |
A1 |
934 |
846 |
8.95 |
|
|
|
| 11 |
A1 |
468 |
424 |
0.18 |
|
|
|
| 12 |
A1 |
205 |
186 |
0.67 |
|
|
|
| 13 |
A2 |
3236 |
2933 |
0.00 |
|
|
|
| 14 |
A2 |
3132 |
2839 |
0.00 |
|
|
|
| 15 |
A2 |
1599 |
1449 |
0.00 |
|
|
|
| 16 |
A2 |
1409 |
1277 |
0.00 |
|
|
|
| 17 |
A2 |
1274 |
1155 |
0.00 |
|
|
|
| 18 |
A2 |
873 |
792 |
0.00 |
|
|
|
| 19 |
A2 |
270 |
244 |
0.00 |
|
|
|
| 20 |
A2 |
98 |
89 |
0.00 |
|
|
|
| 21 |
B1 |
3237 |
2934 |
100.98 |
|
|
|
| 22 |
B1 |
3136 |
2843 |
122.89 |
|
|
|
| 23 |
B1 |
1599 |
1450 |
9.45 |
|
|
|
| 24 |
B1 |
1421 |
1288 |
6.49 |
|
|
|
| 25 |
B1 |
1304 |
1182 |
11.90 |
|
|
|
| 26 |
B1 |
888 |
805 |
0.04 |
|
|
|
| 27 |
B1 |
278 |
252 |
1.81 |
|
|
|
| 28 |
B1 |
116 |
105 |
4.96 |
|
|
|
| 29 |
B2 |
3226 |
2924 |
29.31 |
|
|
|
| 30 |
B2 |
3166 |
2870 |
49.30 |
|
|
|
| 31 |
B2 |
3107 |
2816 |
5.44 |
|
|
|
| 32 |
B2 |
1642 |
1489 |
3.24 |
|
|
|
| 33 |
B2 |
1615 |
1464 |
0.05 |
|
|
|
| 34 |
B2 |
1544 |
1400 |
53.41 |
|
|
|
| 35 |
B2 |
1508 |
1367 |
35.19 |
|
|
|
| 36 |
B2 |
1284 |
1164 |
258.19 |
|
|
|
| 37 |
B2 |
1184 |
1073 |
4.86 |
|
|
|
| 38 |
B2 |
1007 |
912 |
3.77 |
|
|
|
| 39 |
B2 |
464 |
421 |
2.36 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31776.7 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 28802.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.397 |
|
|
|
| 2 |
C |
0.154 |
|
|
|
| 3 |
C |
0.154 |
|
|
|
| 4 |
C |
-0.159 |
|
|
|
| 5 |
C |
-0.159 |
|
|
|
| 6 |
H |
0.024 |
|
|
|
| 7 |
H |
0.024 |
|
|
|
| 8 |
H |
0.024 |
|
|
|
| 9 |
H |
0.024 |
|
|
|
| 10 |
H |
0.042 |
|
|
|
| 11 |
H |
0.042 |
|
|
|
| 12 |
H |
0.057 |
|
|
|
| 13 |
H |
0.057 |
|
|
|
| 14 |
H |
0.057 |
|
|
|
| 15 |
H |
0.057 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-1.178 |
1.178 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.946 |
0.000 |
0.000 |
| y |
0.000 |
-31.062 |
0.000 |
| z |
0.000 |
0.000 |
-33.716 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.558 |
0.000 |
0.000 |
| y |
0.000 |
2.269 |
0.000 |
| z |
0.000 |
0.000 |
-1.712 |
|
| Polar |
|---|
| 3z2-r2 | -3.423 |
|---|
| x2-y2 | -1.885 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.963 |
0.000 |
0.000 |
| y |
0.000 |
9.034 |
0.000 |
| z |
0.000 |
0.000 |
7.293 |
<r2> (average value of r
2) Å
2
| <r2> |
175.024 |
| (<r2>)1/2 |
13.230 |