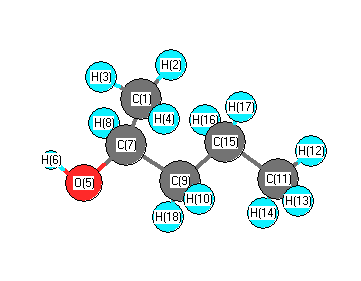
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
4176 |
3785 |
44.25 |
|
|
|
| 2 |
A |
3237 |
2934 |
44.97 |
|
|
|
| 3 |
A |
3219 |
2917 |
138.66 |
|
|
|
| 4 |
A |
3213 |
2913 |
63.47 |
|
|
|
| 5 |
A |
3209 |
2909 |
31.69 |
|
|
|
| 6 |
A |
3197 |
2898 |
11.36 |
|
|
|
| 7 |
A |
3173 |
2876 |
2.56 |
|
|
|
| 8 |
A |
3161 |
2865 |
45.83 |
|
|
|
| 9 |
A |
3155 |
2860 |
48.66 |
|
|
|
| 10 |
A |
3154 |
2859 |
22.04 |
|
|
|
| 11 |
A |
3145 |
2851 |
22.64 |
|
|
|
| 12 |
A |
3113 |
2822 |
42.19 |
|
|
|
| 13 |
A |
1630 |
1478 |
5.22 |
|
|
|
| 14 |
A |
1620 |
1469 |
0.57 |
|
|
|
| 15 |
A |
1616 |
1464 |
1.68 |
|
|
|
| 16 |
A |
1614 |
1463 |
9.09 |
|
|
|
| 17 |
A |
1612 |
1461 |
2.19 |
|
|
|
| 18 |
A |
1604 |
1454 |
0.29 |
|
|
|
| 19 |
A |
1557 |
1411 |
14.40 |
|
|
|
| 20 |
A |
1544 |
1400 |
26.13 |
|
|
|
| 21 |
A |
1538 |
1394 |
0.34 |
|
|
|
| 22 |
A |
1517 |
1375 |
6.05 |
|
|
|
| 23 |
A |
1500 |
1360 |
4.74 |
|
|
|
| 24 |
A |
1436 |
1301 |
2.72 |
|
|
|
| 25 |
A |
1427 |
1293 |
15.05 |
|
|
|
| 26 |
A |
1389 |
1259 |
11.52 |
|
|
|
| 27 |
A |
1347 |
1221 |
27.80 |
|
|
|
| 28 |
A |
1284 |
1164 |
26.69 |
|
|
|
| 29 |
A |
1247 |
1130 |
45.41 |
|
|
|
| 30 |
A |
1190 |
1079 |
43.40 |
|
|
|
| 31 |
A |
1152 |
1044 |
1.28 |
|
|
|
| 32 |
A |
1104 |
1001 |
3.96 |
|
|
|
| 33 |
A |
1082 |
981 |
2.52 |
|
|
|
| 34 |
A |
1042 |
945 |
19.11 |
|
|
|
| 35 |
A |
975 |
884 |
15.19 |
|
|
|
| 36 |
A |
941 |
853 |
0.52 |
|
|
|
| 37 |
A |
890 |
807 |
0.22 |
|
|
|
| 38 |
A |
799 |
724 |
1.65 |
|
|
|
| 39 |
A |
529 |
479 |
8.72 |
|
|
|
| 40 |
A |
482 |
437 |
2.70 |
|
|
|
| 41 |
A |
441 |
400 |
7.97 |
|
|
|
| 42 |
A |
342 |
310 |
0.86 |
|
|
|
| 43 |
A |
299 |
271 |
119.19 |
|
|
|
| 44 |
A |
276 |
250 |
3.15 |
|
|
|
| 45 |
A |
258 |
234 |
1.55 |
|
|
|
| 46 |
A |
199 |
181 |
1.83 |
|
|
|
| 47 |
A |
118 |
107 |
0.03 |
|
|
|
| 48 |
A |
81 |
73 |
0.60 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 38416.5 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 34820.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.216 |
|
|
|
| 2 |
H |
0.049 |
|
|
|
| 3 |
H |
0.043 |
|
|
|
| 4 |
H |
0.063 |
|
|
|
| 5 |
O |
-0.421 |
|
|
|
| 6 |
H |
0.190 |
|
|
|
| 7 |
C |
0.273 |
|
|
|
| 8 |
H |
0.008 |
|
|
|
| 9 |
C |
-0.060 |
|
|
|
| 10 |
H |
0.038 |
|
|
|
| 11 |
C |
-0.188 |
|
|
|
| 12 |
H |
0.059 |
|
|
|
| 13 |
H |
0.050 |
|
|
|
| 14 |
H |
0.050 |
|
|
|
| 15 |
C |
-0.050 |
|
|
|
| 16 |
H |
0.029 |
|
|
|
| 17 |
H |
0.029 |
|
|
|
| 18 |
H |
0.054 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.334 |
1.368 |
-0.829 |
1.634 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.606 |
-1.730 |
1.936 |
| y |
-1.730 |
-42.757 |
0.919 |
| z |
1.936 |
0.919 |
-39.905 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.724 |
-1.730 |
1.936 |
| y |
-1.730 |
-4.501 |
0.919 |
| z |
1.936 |
0.919 |
-0.223 |
|
| Polar |
|---|
| 3z2-r2 | -0.446 |
|---|
| x2-y2 | 6.151 |
|---|
| xy | -1.730 |
|---|
| xz | 1.936 |
|---|
| yz | 0.919 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.643 |
-0.140 |
0.046 |
| y |
-0.140 |
8.841 |
-0.072 |
| z |
0.046 |
-0.072 |
8.308 |
<r2> (average value of r
2) Å
2
| <r2> |
238.516 |
| (<r2>)1/2 |
15.444 |