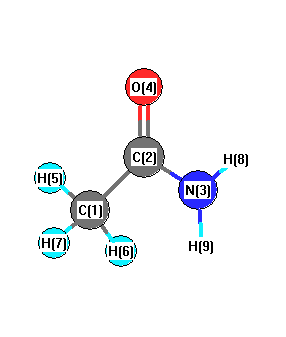
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3968 |
3597 |
54.25 |
|
|
|
| 2 |
A |
3830 |
3472 |
52.79 |
|
|
|
| 3 |
A |
3287 |
2980 |
11.54 |
|
|
|
| 4 |
A |
3236 |
2933 |
20.35 |
|
|
|
| 5 |
A |
3178 |
2881 |
9.09 |
|
|
|
| 6 |
A |
1937 |
1755 |
476.19 |
|
|
|
| 7 |
A |
1771 |
1605 |
114.51 |
|
|
|
| 8 |
A |
1606 |
1456 |
8.61 |
|
|
|
| 9 |
A |
1588 |
1439 |
10.08 |
|
|
|
| 10 |
A |
1535 |
1391 |
73.59 |
|
|
|
| 11 |
A |
1443 |
1308 |
150.19 |
|
|
|
| 12 |
A |
1225 |
1110 |
0.98 |
|
|
|
| 13 |
A |
1158 |
1049 |
6.24 |
|
|
|
| 14 |
A |
1068 |
968 |
18.20 |
|
|
|
| 15 |
A |
905 |
820 |
1.04 |
|
|
|
| 16 |
A |
699 |
633 |
14.46 |
|
|
|
| 17 |
A |
601 |
545 |
20.37 |
|
|
|
| 18 |
A |
558 |
506 |
3.46 |
|
|
|
| 19 |
A |
453 |
411 |
4.32 |
|
|
|
| 20 |
A |
151 |
137 |
229.85 |
|
|
|
| 21 |
A |
46 |
42 |
2.58 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17120.7 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 15518.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.193 |
|
|
|
| 2 |
C |
0.464 |
|
|
|
| 3 |
N |
-0.425 |
|
|
|
| 4 |
O |
-0.487 |
|
|
|
| 5 |
H |
0.104 |
|
|
|
| 6 |
H |
0.058 |
|
|
|
| 7 |
H |
0.078 |
|
|
|
| 8 |
H |
0.212 |
|
|
|
| 9 |
H |
0.191 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.363 |
-4.060 |
0.181 |
4.081 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.516 |
-2.298 |
0.267 |
| y |
-2.298 |
-26.457 |
-0.248 |
| z |
0.267 |
-0.248 |
-24.938 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.182 |
-2.298 |
0.267 |
| y |
-2.298 |
-3.730 |
-0.248 |
| z |
0.267 |
-0.248 |
-1.451 |
|
| Polar |
|---|
| 3z2-r2 | -2.903 |
|---|
| x2-y2 | 5.941 |
|---|
| xy | -2.298 |
|---|
| xz | 0.267 |
|---|
| yz | -0.248 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.171 |
-0.042 |
-0.009 |
| y |
-0.042 |
5.522 |
-0.001 |
| z |
-0.009 |
-0.001 |
3.764 |
<r2> (average value of r
2) Å
2
| <r2> |
73.950 |
| (<r2>)1/2 |
8.599 |