Jump to
S1C2
Energy calculated at QCISD(T)=FULL/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -190.421721 |
| Energy at 298.15K | |
| HF Energy | -189.612438 |
| Nuclear repulsion energy | 88.172532 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)=FULL/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3098 |
3098 |
|
|
|
|
| 2 |
A1 |
2275 |
2275 |
|
|
|
|
| 3 |
A1 |
1750 |
1750 |
|
|
|
|
| 4 |
A1 |
1497 |
1497 |
|
|
|
|
| 5 |
A1 |
904 |
904 |
|
|
|
|
| 6 |
B1 |
999 |
999 |
|
|
|
|
| 7 |
B1 |
628 |
628 |
|
|
|
|
| 8 |
B1 |
215 |
215 |
|
|
|
|
| 9 |
B2 |
3174 |
3174 |
|
|
|
|
| 10 |
B2 |
1074 |
1074 |
|
|
|
|
| 11 |
B2 |
435 |
435 |
|
|
|
|
| 12 |
B2 |
152i |
152i |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 7948.6 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 7948.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)=FULL/Def2TZVPP
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.882 |
| C2 |
0.000 |
0.000 |
-0.558 |
| C3 |
0.000 |
0.000 |
0.723 |
| O4 |
0.000 |
0.000 |
1.901 |
| H5 |
0.000 |
0.923 |
-2.452 |
| H6 |
0.000 |
-0.923 |
-2.452 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
| C1 | | 1.3240 | 2.6042 | 3.7822 | 1.0850 | 1.0850 |
C2 | 1.3240 | | 1.2802 | 2.4582 | 2.1069 | 2.1069 | C3 | 2.6042 | 1.2802 | | 1.1780 | 3.3056 | 3.3056 | O4 | 3.7822 | 2.4582 | 1.1780 | | 4.4489 | 4.4489 | H5 | 1.0850 | 2.1069 | 3.3056 | 4.4489 | | 1.8466 | H6 | 1.0850 | 2.1069 | 3.3056 | 4.4489 | 1.8466 | |
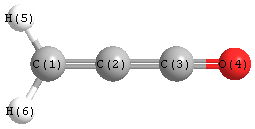 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
180.000 |
|
C2 |
C1 |
H5 |
121.684 |
| C2 |
C1 |
H6 |
121.684 |
|
C2 |
C3 |
O4 |
180.000 |
| H5 |
C1 |
H6 |
116.633 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD(T)=FULL/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -190.423808 |
| Energy at 298.15K | |
| HF Energy | -189.611457 |
| Nuclear repulsion energy | 88.499577 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)=FULL/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3193 |
3193 |
|
|
|
|
| 2 |
A' |
3110 |
3110 |
|
|
|
|
| 3 |
A' |
2156 |
2156 |
|
|
|
|
| 4 |
A' |
1707 |
1707 |
|
|
|
|
| 5 |
A' |
1495 |
1495 |
|
|
|
|
| 6 |
A' |
1083 |
1083 |
|
|
|
|
| 7 |
A' |
933 |
933 |
|
|
|
|
| 8 |
A' |
509 |
509 |
|
|
|
|
| 9 |
A' |
186 |
186 |
|
|
|
|
| 10 |
A" |
1011 |
1011 |
|
|
|
|
| 11 |
A" |
703 |
703 |
|
|
|
|
| 12 |
A" |
275 |
275 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 8179.8 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 8179.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)=FULL/Def2TZVPP
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.101 |
-1.438 |
0.000 |
| C2 |
0.000 |
-0.683 |
0.000 |
| C3 |
-0.370 |
0.578 |
0.000 |
| O4 |
-0.940 |
1.599 |
0.000 |
| H5 |
2.101 |
-1.018 |
0.000 |
| H6 |
1.029 |
-2.519 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
| C1 | | 1.3354 | 2.4959 | 3.6592 | 1.0846 | 1.0829 |
C2 | 1.3354 | | 1.3142 | 2.4677 | 2.1278 | 2.1047 | C3 | 2.4959 | 1.3142 | | 1.1691 | 2.9417 | 3.3983 | O4 | 3.6592 | 2.4677 | 1.1691 | | 4.0121 | 4.5642 | H5 | 1.0846 | 2.1278 | 2.9417 | 4.0121 | | 1.8441 | H6 | 1.0829 | 2.1047 | 3.3983 | 4.5642 | 1.8441 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
140.778 |
|
C2 |
C1 |
H5 |
122.770 |
| C2 |
C1 |
H6 |
120.639 |
|
C2 |
C3 |
O4 |
167.128 |
| H5 |
C1 |
H6 |
116.591 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
