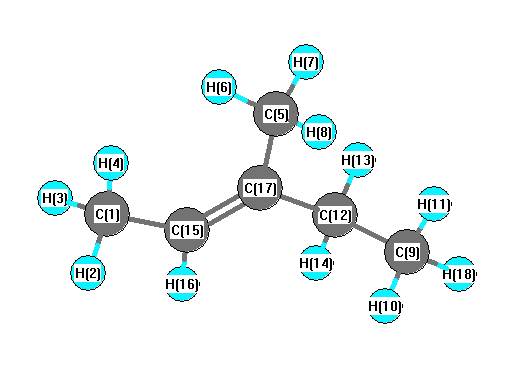
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3280 |
2973 |
64.87 |
|
|
|
| 2 |
A |
3269 |
2963 |
21.27 |
|
|
|
| 3 |
A |
3248 |
2944 |
10.44 |
|
|
|
| 4 |
A |
3228 |
2926 |
62.56 |
|
|
|
| 5 |
A |
3216 |
2915 |
61.73 |
|
|
|
| 6 |
A |
3200 |
2900 |
21.89 |
|
|
|
| 7 |
A |
3194 |
2895 |
19.90 |
|
|
|
| 8 |
A |
3188 |
2889 |
32.09 |
|
|
|
| 9 |
A |
3162 |
2866 |
37.43 |
|
|
|
| 10 |
A |
3158 |
2862 |
23.26 |
|
|
|
| 11 |
A |
3153 |
2858 |
65.01 |
|
|
|
| 12 |
A |
3141 |
2847 |
34.06 |
|
|
|
| 13 |
A |
1877 |
1701 |
3.13 |
|
|
|
| 14 |
A |
1626 |
1474 |
3.27 |
|
|
|
| 15 |
A |
1619 |
1468 |
10.21 |
|
|
|
| 16 |
A |
1618 |
1467 |
3.60 |
|
|
|
| 17 |
A |
1609 |
1459 |
5.30 |
|
|
|
| 18 |
A |
1607 |
1456 |
8.51 |
|
|
|
| 19 |
A |
1602 |
1452 |
3.79 |
|
|
|
| 20 |
A |
1598 |
1448 |
0.36 |
|
|
|
| 21 |
A |
1548 |
1403 |
2.49 |
|
|
|
| 22 |
A |
1544 |
1399 |
0.82 |
|
|
|
| 23 |
A |
1531 |
1388 |
1.46 |
|
|
|
| 24 |
A |
1496 |
1356 |
1.54 |
|
|
|
| 25 |
A |
1478 |
1340 |
8.19 |
|
|
|
| 26 |
A |
1390 |
1260 |
0.13 |
|
|
|
| 27 |
A |
1329 |
1204 |
3.01 |
|
|
|
| 28 |
A |
1231 |
1115 |
4.27 |
|
|
|
| 29 |
A |
1192 |
1080 |
7.20 |
|
|
|
| 30 |
A |
1167 |
1057 |
5.25 |
|
|
|
| 31 |
A |
1166 |
1057 |
0.21 |
|
|
|
| 32 |
A |
1131 |
1025 |
1.65 |
|
|
|
| 33 |
A |
1083 |
982 |
1.95 |
|
|
|
| 34 |
A |
1054 |
955 |
6.19 |
|
|
|
| 35 |
A |
984 |
892 |
1.47 |
|
|
|
| 36 |
A |
941 |
853 |
12.32 |
|
|
|
| 37 |
A |
846 |
767 |
1.04 |
|
|
|
| 38 |
A |
793 |
718 |
1.28 |
|
|
|
| 39 |
A |
598 |
542 |
5.19 |
|
|
|
| 40 |
A |
529 |
480 |
2.78 |
|
|
|
| 41 |
A |
431 |
390 |
0.36 |
|
|
|
| 42 |
A |
345 |
312 |
0.73 |
|
|
|
| 43 |
A |
330 |
299 |
0.22 |
|
|
|
| 44 |
A |
253 |
230 |
0.24 |
|
|
|
| 45 |
A |
201 |
182 |
1.09 |
|
|
|
| 46 |
A |
125 |
113 |
0.61 |
|
|
|
| 47 |
A |
120 |
109 |
0.12 |
|
|
|
| 48 |
A |
64 |
58 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 38244.3 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 34664.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.154 |
|
|
|
| 2 |
H |
0.060 |
|
|
|
| 3 |
H |
0.060 |
|
|
|
| 4 |
H |
0.052 |
|
|
|
| 5 |
C |
-0.204 |
|
|
|
| 6 |
H |
0.066 |
|
|
|
| 7 |
H |
0.062 |
|
|
|
| 8 |
H |
0.067 |
|
|
|
| 9 |
C |
-0.184 |
|
|
|
| 10 |
H |
0.056 |
|
|
|
| 11 |
H |
0.048 |
|
|
|
| 12 |
C |
-0.011 |
|
|
|
| 13 |
H |
0.035 |
|
|
|
| 14 |
H |
0.045 |
|
|
|
| 15 |
C |
-0.055 |
|
|
|
| 16 |
H |
0.042 |
|
|
|
| 17 |
C |
-0.045 |
|
|
|
| 18 |
H |
0.060 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.059 |
0.171 |
0.005 |
0.181 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.632 |
0.162 |
-0.699 |
| y |
0.162 |
-39.216 |
0.081 |
| z |
-0.699 |
0.081 |
-41.537 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.745 |
0.162 |
-0.699 |
| y |
0.162 |
1.368 |
0.081 |
| z |
-0.699 |
0.081 |
-2.113 |
|
| Polar |
|---|
| 3z2-r2 | -4.226 |
|---|
| x2-y2 | -0.415 |
|---|
| xy | 0.162 |
|---|
| xz | -0.699 |
|---|
| yz | 0.081 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.987 |
1.089 |
-0.276 |
| y |
1.089 |
10.282 |
-0.037 |
| z |
-0.276 |
-0.037 |
8.528 |
<r2> (average value of r
2) Å
2
| <r2> |
222.422 |
| (<r2>)1/2 |
14.914 |