Jump to
S1C2
Energy calculated at B2PLYP/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -278.231135 |
| Energy at 298.15K | |
| HF Energy | -277.961818 |
| Nuclear repulsion energy | 125.654441 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3084 |
3084 |
0.00 |
233.02 |
0.06 |
0.12 |
| 2 |
Ag |
1530 |
1530 |
0.00 |
9.90 |
0.74 |
0.85 |
| 3 |
Ag |
1459 |
1459 |
0.00 |
2.72 |
0.42 |
0.59 |
| 4 |
Ag |
1098 |
1098 |
0.00 |
7.05 |
0.40 |
0.57 |
| 5 |
Ag |
1064 |
1064 |
0.00 |
8.74 |
0.49 |
0.66 |
| 6 |
Ag |
460 |
460 |
0.00 |
2.81 |
0.44 |
0.61 |
| 7 |
Au |
3157 |
3157 |
44.20 |
0.00 |
0.00 |
0.00 |
| 8 |
Au |
1244 |
1244 |
4.30 |
0.00 |
0.00 |
0.00 |
| 9 |
Au |
822 |
822 |
0.01 |
0.00 |
0.00 |
0.00 |
| 10 |
Au |
118 |
118 |
13.25 |
0.00 |
0.00 |
0.00 |
| 11 |
Bg |
3131 |
3131 |
0.00 |
113.45 |
0.75 |
0.86 |
| 12 |
Bg |
1314 |
1314 |
0.00 |
10.19 |
0.75 |
0.86 |
| 13 |
Bg |
1186 |
1186 |
0.00 |
2.69 |
0.75 |
0.86 |
| 14 |
Bu |
3090 |
3090 |
54.85 |
0.00 |
0.07 |
0.00 |
| 15 |
Bu |
1541 |
1541 |
3.19 |
0.00 |
0.00 |
0.00 |
| 16 |
Bu |
1375 |
1375 |
8.22 |
0.00 |
0.00 |
0.00 |
| 17 |
Bu |
1072 |
1072 |
209.23 |
0.00 |
0.00 |
0.00 |
| 18 |
Bu |
282 |
282 |
19.47 |
0.00 |
0.00 |
0.00 |
Unscaled Zero Point Vibrational Energy (zpe) 13512.9 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 13512.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/Def2TZVPP
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.425 |
0.626 |
0.000 |
| C2 |
-0.425 |
-0.626 |
0.000 |
| F3 |
-0.425 |
1.726 |
0.000 |
| F4 |
0.425 |
-1.726 |
0.000 |
| H5 |
1.050 |
0.673 |
0.890 |
| H6 |
1.050 |
0.673 |
-0.890 |
| H7 |
-1.050 |
-0.673 |
0.890 |
| H8 |
-1.050 |
-0.673 |
-0.890 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5132 | 1.3903 | 2.3514 | 1.0882 | 1.0882 | 2.1573 | 2.1573 |
C2 | 1.5132 | | 2.3514 | 1.3903 | 2.1573 | 2.1573 | 1.0882 | 1.0882 | F3 | 1.3903 | 2.3514 | | 3.5545 | 2.0191 | 2.0191 | 2.6332 | 2.6332 | F4 | 2.3514 | 1.3903 | 3.5545 | | 2.6332 | 2.6332 | 2.0191 | 2.0191 | H5 | 1.0882 | 2.1573 | 2.0191 | 2.6332 | | 1.7791 | 2.4943 | 3.0638 | H6 | 1.0882 | 2.1573 | 2.0191 | 2.6332 | 1.7791 | | 3.0638 | 2.4943 | H7 | 2.1573 | 1.0882 | 2.6332 | 2.0191 | 2.4943 | 3.0638 | | 1.7791 | H8 | 2.1573 | 1.0882 | 2.6332 | 2.0191 | 3.0638 | 2.4943 | 1.7791 | |
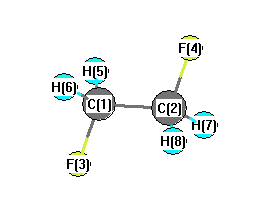 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
108.090 |
|
C1 |
C2 |
H7 |
111.005 |
| C1 |
C2 |
H8 |
111.005 |
|
C2 |
C1 |
F3 |
108.090 |
| C2 |
C1 |
H5 |
111.005 |
|
C2 |
C1 |
H6 |
111.005 |
| F3 |
C1 |
H5 |
108.494 |
|
F3 |
C1 |
H6 |
108.494 |
| F4 |
C2 |
H7 |
108.494 |
|
F4 |
C2 |
H8 |
108.494 |
| H5 |
C1 |
H6 |
109.670 |
|
H7 |
C2 |
H8 |
109.670 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.085 |
|
|
|
| 2 |
C |
0.085 |
|
|
|
| 3 |
F |
-0.253 |
|
|
|
| 4 |
F |
-0.253 |
|
|
|
| 5 |
H |
0.084 |
|
|
|
| 6 |
H |
0.084 |
|
|
|
| 7 |
H |
0.084 |
|
|
|
| 8 |
H |
0.084 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.256 |
3.056 |
0.000 |
| y |
3.056 |
-29.412 |
0.000 |
| z |
0.000 |
0.000 |
-21.690 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.295 |
3.056 |
0.000 |
| y |
3.056 |
-7.439 |
0.000 |
| z |
0.000 |
0.000 |
4.144 |
|
| Polar |
|---|
| 3z2-r2 | 8.289 |
|---|
| x2-y2 | 7.156 |
|---|
| xy | 3.056 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.029 |
-0.044 |
0.000 |
| y |
-0.044 |
4.198 |
0.000 |
| z |
0.000 |
0.000 |
3.730 |
<r2> (average value of r
2) Å
2
| <r2> |
88.382 |
| (<r2>)1/2 |
9.401 |
Jump to
S1C1
Energy calculated at B2PLYP/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -278.232499 |
| Energy at 298.15K | |
| HF Energy | -277.962887 |
| Nuclear repulsion energy | 127.737194 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3124 |
3124 |
13.09 |
90.79 |
0.73 |
0.85 |
| 2 |
A |
3076 |
3076 |
37.45 |
212.88 |
0.02 |
0.03 |
| 3 |
A |
1505 |
1505 |
0.52 |
1.55 |
0.70 |
0.82 |
| 4 |
A |
1451 |
1451 |
11.22 |
2.34 |
0.28 |
0.43 |
| 5 |
A |
1316 |
1316 |
2.13 |
12.10 |
0.69 |
0.82 |
| 6 |
A |
1139 |
1139 |
0.21 |
1.13 |
0.05 |
0.10 |
| 7 |
A |
1120 |
1120 |
88.98 |
3.60 |
0.70 |
0.82 |
| 8 |
A |
878 |
878 |
26.37 |
5.34 |
0.19 |
0.32 |
| 9 |
A |
326 |
326 |
0.36 |
0.41 |
0.38 |
0.55 |
| 10 |
A |
152 |
152 |
3.74 |
0.13 |
0.74 |
0.85 |
| 11 |
B |
3137 |
3137 |
33.77 |
12.08 |
0.75 |
0.86 |
| 12 |
B |
3066 |
3066 |
8.77 |
35.49 |
0.75 |
0.86 |
| 13 |
B |
1503 |
1503 |
7.75 |
8.22 |
0.75 |
0.86 |
| 14 |
B |
1418 |
1418 |
8.38 |
0.21 |
0.75 |
0.86 |
| 15 |
B |
1273 |
1273 |
5.70 |
3.21 |
0.75 |
0.86 |
| 16 |
B |
1092 |
1092 |
49.64 |
3.46 |
0.75 |
0.86 |
| 17 |
B |
912 |
912 |
46.37 |
3.03 |
0.75 |
0.86 |
| 18 |
B |
500 |
500 |
16.41 |
0.62 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13492.7 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 13492.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/Def2TZVPP
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.267 |
0.701 |
0.510 |
| C2 |
-0.267 |
-0.701 |
0.510 |
| F3 |
-0.267 |
1.427 |
-0.545 |
| F4 |
0.267 |
-1.427 |
-0.545 |
| H5 |
-0.011 |
1.200 |
1.440 |
| H6 |
1.351 |
0.700 |
0.404 |
| H7 |
0.011 |
-1.200 |
1.440 |
| H8 |
-1.351 |
-0.700 |
0.404 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5006 | 1.3880 | 2.3758 | 1.0907 | 1.0889 | 2.1314 | 2.1430 |
C2 | 1.5006 | | 2.3758 | 1.3880 | 2.1314 | 2.1430 | 1.0907 | 1.0889 | F3 | 1.3880 | 2.3758 | | 2.9044 | 2.0139 | 2.0117 | 3.3043 | 2.5693 | F4 | 2.3758 | 1.3880 | 2.9044 | | 3.3043 | 2.5693 | 2.0139 | 2.0117 | H5 | 1.0907 | 2.1314 | 2.0139 | 3.3043 | | 1.7826 | 2.3998 | 2.5450 | H6 | 1.0889 | 2.1430 | 2.0117 | 2.5693 | 1.7826 | | 2.5450 | 3.0429 | H7 | 2.1314 | 1.0907 | 3.3043 | 2.0139 | 2.3998 | 2.5450 | | 1.7826 | H8 | 2.1430 | 1.0889 | 2.5693 | 2.0117 | 2.5450 | 3.0429 | 1.7826 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
110.605 |
|
C1 |
C2 |
H7 |
109.655 |
| C1 |
C2 |
H8 |
110.697 |
|
C2 |
C1 |
F3 |
110.605 |
| C2 |
C1 |
H5 |
109.655 |
|
C2 |
C1 |
H6 |
110.697 |
| F3 |
C1 |
H5 |
108.076 |
|
F3 |
C1 |
H6 |
108.012 |
| F4 |
C2 |
H7 |
108.076 |
|
F4 |
C2 |
H8 |
108.012 |
| H5 |
C1 |
H6 |
109.740 |
|
H7 |
C2 |
H8 |
109.740 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.100 |
|
|
|
| 2 |
C |
0.100 |
|
|
|
| 3 |
F |
-0.248 |
|
|
|
| 4 |
F |
-0.248 |
|
|
|
| 5 |
H |
0.065 |
|
|
|
| 6 |
H |
0.082 |
|
|
|
| 7 |
H |
0.065 |
|
|
|
| 8 |
H |
0.082 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
2.709 |
2.709 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-21.884 |
1.672 |
0.000 |
| y |
1.672 |
-26.368 |
0.000 |
| z |
0.000 |
0.000 |
-22.527 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.563 |
1.672 |
0.000 |
| y |
1.672 |
-4.162 |
0.000 |
| z |
0.000 |
0.000 |
1.599 |
|
| Polar |
|---|
| 3z2-r2 | 3.199 |
|---|
| x2-y2 | 4.483 |
|---|
| xy | 1.672 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.868 |
0.031 |
0.000 |
| y |
0.031 |
4.170 |
0.000 |
| z |
0.000 |
0.000 |
4.027 |
<r2> (average value of r
2) Å
2
| <r2> |
79.907 |
| (<r2>)1/2 |
8.939 |
