Jump to
S1C2
Energy calculated at QCISD(T)=FULL/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -278.018707 |
| Energy at 298.15K | |
| HF Energy | -277.049214 |
| Nuclear repulsion energy | 125.866390 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)=FULL/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3071 |
3071 |
|
|
|
|
| 2 |
Ag |
1524 |
1524 |
|
|
|
|
| 3 |
Ag |
1464 |
1464 |
|
|
|
|
| 4 |
Ag |
1108 |
1108 |
|
|
|
|
| 5 |
Ag |
1084 |
1084 |
|
|
|
|
| 6 |
Ag |
462 |
462 |
|
|
|
|
| 7 |
Au |
3141 |
3141 |
|
|
|
|
| 8 |
Au |
1248 |
1248 |
|
|
|
|
| 9 |
Au |
816 |
816 |
|
|
|
|
| 10 |
Au |
117 |
117 |
|
|
|
|
| 11 |
Bg |
3117 |
3117 |
|
|
|
|
| 12 |
Bg |
1315 |
1315 |
|
|
|
|
| 13 |
Bg |
1187 |
1187 |
|
|
|
|
| 14 |
Bu |
3076 |
3076 |
|
|
|
|
| 15 |
Bu |
1534 |
1534 |
|
|
|
|
| 16 |
Bu |
1372 |
1372 |
|
|
|
|
| 17 |
Bu |
1095 |
1095 |
|
|
|
|
| 18 |
Bu |
279 |
279 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13504.0 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 13504.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)=FULL/Def2TZVPP
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.425 |
0.627 |
0.000 |
| C2 |
-0.425 |
-0.627 |
0.000 |
| F3 |
-0.425 |
1.722 |
0.000 |
| F4 |
0.425 |
-1.722 |
0.000 |
| H5 |
1.050 |
0.670 |
0.889 |
| H6 |
1.050 |
0.670 |
-0.889 |
| H7 |
-1.050 |
-0.670 |
0.889 |
| H8 |
-1.050 |
-0.670 |
-0.889 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5147 | 1.3853 | 2.3489 | 1.0883 | 1.0883 | 2.1559 | 2.1559 |
C2 | 1.5147 | | 2.3489 | 1.3853 | 2.1559 | 2.1559 | 1.0883 | 1.0883 | F3 | 1.3853 | 2.3489 | | 3.5466 | 2.0183 | 2.0183 | 2.6270 | 2.6270 | F4 | 2.3489 | 1.3853 | 3.5466 | | 2.6270 | 2.6270 | 2.0183 | 2.0183 | H5 | 1.0883 | 2.1559 | 2.0183 | 2.6270 | | 1.7787 | 2.4913 | 3.0611 | H6 | 1.0883 | 2.1559 | 2.0183 | 2.6270 | 1.7787 | | 3.0611 | 2.4913 | H7 | 2.1559 | 1.0883 | 2.6270 | 2.0183 | 2.4913 | 3.0611 | | 1.7787 | H8 | 2.1559 | 1.0883 | 2.6270 | 2.0183 | 3.0611 | 2.4913 | 1.7787 | |
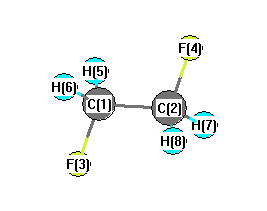 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
108.103 |
|
C1 |
C2 |
H7 |
110.770 |
| C1 |
C2 |
H8 |
110.770 |
|
C2 |
C1 |
F3 |
108.103 |
| C2 |
C1 |
H5 |
110.770 |
|
C2 |
C1 |
H6 |
110.770 |
| F3 |
C1 |
H5 |
108.766 |
|
F3 |
C1 |
H6 |
108.766 |
| F4 |
C2 |
H7 |
108.766 |
|
F4 |
C2 |
H8 |
108.766 |
| H5 |
C1 |
H6 |
109.608 |
|
H7 |
C2 |
H8 |
109.608 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD(T)=FULL/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -278.019885 |
| Energy at 298.15K | |
| HF Energy | -277.049716 |
| Nuclear repulsion energy | 128.008189 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)=FULL/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3112 |
3112 |
|
|
|
|
| 2 |
A |
3064 |
3064 |
|
|
|
|
| 3 |
A |
1501 |
1501 |
|
|
|
|
| 4 |
A |
1458 |
1458 |
|
|
|
|
| 5 |
A |
1318 |
1318 |
|
|
|
|
| 6 |
A |
1142 |
1142 |
|
|
|
|
| 7 |
A |
1140 |
1140 |
|
|
|
|
| 8 |
A |
883 |
883 |
|
|
|
|
| 9 |
A |
324 |
324 |
|
|
|
|
| 10 |
A |
151 |
151 |
|
|
|
|
| 11 |
B |
3124 |
3124 |
|
|
|
|
| 12 |
B |
3054 |
3054 |
|
|
|
|
| 13 |
B |
1500 |
1500 |
|
|
|
|
| 14 |
B |
1418 |
1418 |
|
|
|
|
| 15 |
B |
1277 |
1277 |
|
|
|
|
| 16 |
B |
1108 |
1108 |
|
|
|
|
| 17 |
B |
916 |
916 |
|
|
|
|
| 18 |
B |
499 |
499 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13494.3 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 13494.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)=FULL/Def2TZVPP
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.265 |
0.703 |
0.511 |
| C2 |
-0.265 |
-0.703 |
0.511 |
| F3 |
-0.265 |
1.419 |
-0.546 |
| F4 |
0.265 |
-1.419 |
-0.546 |
| H5 |
-0.018 |
1.201 |
1.439 |
| H6 |
1.350 |
0.700 |
0.410 |
| H7 |
0.018 |
-1.201 |
1.439 |
| H8 |
-1.350 |
-0.700 |
0.410 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5029 | 1.3833 | 2.3712 | 1.0906 | 1.0889 | 2.1322 | 2.1419 |
C2 | 1.5029 | | 2.3712 | 1.3833 | 2.1322 | 2.1419 | 1.0906 | 1.0889 | F3 | 1.3833 | 2.3712 | | 2.8875 | 2.0131 | 2.0102 | 3.2995 | 2.5657 | F4 | 2.3712 | 1.3833 | 2.8875 | | 3.2995 | 2.5657 | 2.0131 | 2.0102 | H5 | 1.0906 | 2.1322 | 2.0131 | 3.2995 | | 1.7833 | 2.4014 | 2.5386 | H6 | 1.0889 | 2.1419 | 2.0102 | 2.5657 | 1.7833 | | 2.5386 | 3.0410 | H7 | 2.1322 | 1.0906 | 3.2995 | 2.0131 | 2.4014 | 2.5386 | | 1.7833 | H8 | 2.1419 | 1.0889 | 2.5657 | 2.0102 | 2.5386 | 3.0410 | 1.7833 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
110.410 |
|
C1 |
C2 |
H7 |
109.568 |
| C1 |
C2 |
H8 |
110.446 |
|
C2 |
C1 |
F3 |
110.410 |
| C2 |
C1 |
H5 |
109.568 |
|
C2 |
C1 |
H6 |
110.446 |
| F3 |
C1 |
H5 |
108.346 |
|
F3 |
C1 |
H6 |
108.216 |
| F4 |
C2 |
H7 |
108.346 |
|
F4 |
C2 |
H8 |
108.216 |
| H5 |
C1 |
H6 |
109.815 |
|
H7 |
C2 |
H8 |
109.815 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
