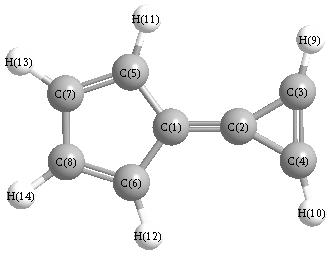
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3460 |
3136 |
19.67 |
|
|
|
| 2 |
A1 |
3374 |
3058 |
17.37 |
|
|
|
| 3 |
A1 |
3348 |
3035 |
6.04 |
|
|
|
| 4 |
A1 |
1894 |
1717 |
825.54 |
|
|
|
| 5 |
A1 |
1678 |
1521 |
232.63 |
|
|
|
| 6 |
A1 |
1601 |
1451 |
260.49 |
|
|
|
| 7 |
A1 |
1513 |
1372 |
328.92 |
|
|
|
| 8 |
A1 |
1303 |
1181 |
31.90 |
|
|
|
| 9 |
A1 |
1158 |
1049 |
32.22 |
|
|
|
| 10 |
A1 |
1070 |
970 |
24.44 |
|
|
|
| 11 |
A1 |
1028 |
932 |
3.60 |
|
|
|
| 12 |
A1 |
943 |
855 |
115.11 |
|
|
|
| 13 |
A1 |
519 |
471 |
0.26 |
|
|
|
| 14 |
A2 |
1049 |
951 |
0.00 |
|
|
|
| 15 |
A2 |
1033 |
936 |
0.00 |
|
|
|
| 16 |
A2 |
804 |
729 |
0.00 |
|
|
|
| 17 |
A2 |
641 |
581 |
0.00 |
|
|
|
| 18 |
A2 |
233 |
211 |
0.00 |
|
|
|
| 19 |
B1 |
1024 |
928 |
0.21 |
|
|
|
| 20 |
B1 |
845 |
766 |
31.44 |
|
|
|
| 21 |
B1 |
835 |
757 |
130.51 |
|
|
|
| 22 |
B1 |
676 |
612 |
17.12 |
|
|
|
| 23 |
B1 |
501 |
454 |
10.47 |
|
|
|
| 24 |
B1 |
148 |
134 |
8.21 |
|
|
|
| 25 |
B2 |
3419 |
3099 |
8.13 |
|
|
|
| 26 |
B2 |
3364 |
3049 |
32.76 |
|
|
|
| 27 |
B2 |
3339 |
3027 |
0.76 |
|
|
|
| 28 |
B2 |
1717 |
1556 |
0.60 |
|
|
|
| 29 |
B2 |
1461 |
1324 |
1.07 |
|
|
|
| 30 |
B2 |
1331 |
1206 |
5.24 |
|
|
|
| 31 |
B2 |
1252 |
1135 |
17.68 |
|
|
|
| 32 |
B2 |
1188 |
1077 |
7.52 |
|
|
|
| 33 |
B2 |
1075 |
974 |
5.45 |
|
|
|
| 34 |
B2 |
896 |
812 |
2.45 |
|
|
|
| 35 |
B2 |
537 |
487 |
0.51 |
|
|
|
| 36 |
B2 |
163 |
147 |
2.05 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25209.2 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 22849.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.103 |
|
|
|
| 2 |
C |
0.340 |
|
|
|
| 3 |
C |
-0.031 |
|
|
|
| 4 |
C |
-0.031 |
|
|
|
| 5 |
C |
-0.309 |
|
|
|
| 6 |
C |
-0.309 |
|
|
|
| 7 |
C |
-0.110 |
|
|
|
| 8 |
C |
-0.110 |
|
|
|
| 9 |
H |
0.183 |
|
|
|
| 10 |
H |
0.183 |
|
|
|
| 11 |
H |
0.068 |
|
|
|
| 12 |
H |
0.068 |
|
|
|
| 13 |
H |
0.081 |
|
|
|
| 14 |
H |
0.081 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
5.058 |
5.058 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-51.842 |
0.000 |
0.000 |
| y |
0.000 |
-41.592 |
0.000 |
| z |
0.000 |
0.000 |
-35.096 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-13.498 |
0.000 |
0.000 |
| y |
0.000 |
1.877 |
0.000 |
| z |
0.000 |
0.000 |
11.621 |
|
| Polar |
|---|
| 3z2-r2 | 23.242 |
|---|
| x2-y2 | -10.250 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.288 |
0.000 |
0.000 |
| y |
0.000 |
12.092 |
0.000 |
| z |
0.000 |
0.000 |
21.989 |
<r2> (average value of r
2) Å
2
| <r2> |
265.886 |
| (<r2>)1/2 |
16.306 |