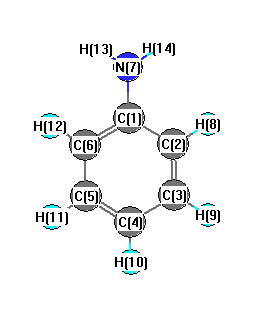
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3788 |
3434 |
23.00 |
|
|
|
| 2 |
A' |
3349 |
3035 |
15.45 |
|
|
|
| 3 |
A' |
3325 |
3013 |
7.61 |
|
|
|
| 4 |
A' |
3307 |
2997 |
16.25 |
|
|
|
| 5 |
A' |
1805 |
1636 |
136.83 |
|
|
|
| 6 |
A' |
1779 |
1613 |
37.51 |
|
|
|
| 7 |
A' |
1655 |
1500 |
68.89 |
|
|
|
| 8 |
A' |
1379 |
1250 |
68.13 |
|
|
|
| 9 |
A' |
1287 |
1166 |
11.57 |
|
|
|
| 10 |
A' |
1118 |
1014 |
2.33 |
|
|
|
| 11 |
A' |
1098 |
995 |
0.28 |
|
|
|
| 12 |
A' |
1078 |
977 |
2.14 |
|
|
|
| 13 |
A' |
992 |
899 |
15.22 |
|
|
|
| 14 |
A' |
885 |
802 |
8.94 |
|
|
|
| 15 |
A' |
845 |
766 |
94.29 |
|
|
|
| 16 |
A' |
770 |
698 |
23.98 |
|
|
|
| 17 |
A' |
677 |
613 |
243.99 |
|
|
|
| 18 |
A' |
572 |
519 |
2.78 |
|
|
|
| 19 |
A' |
553 |
502 |
72.81 |
|
|
|
| 20 |
A' |
244 |
221 |
6.38 |
|
|
|
| 21 |
A" |
3885 |
3522 |
18.23 |
|
|
|
| 22 |
A" |
3331 |
3019 |
43.64 |
|
|
|
| 23 |
A" |
3309 |
3000 |
2.23 |
|
|
|
| 24 |
A" |
1768 |
1602 |
11.95 |
|
|
|
| 25 |
A" |
1622 |
1470 |
3.28 |
|
|
|
| 26 |
A" |
1486 |
1347 |
0.07 |
|
|
|
| 27 |
A" |
1344 |
1218 |
4.31 |
|
|
|
| 28 |
A" |
1241 |
1125 |
1.81 |
|
|
|
| 29 |
A" |
1175 |
1065 |
18.39 |
|
|
|
| 30 |
A" |
1133 |
1027 |
0.85 |
|
|
|
| 31 |
A" |
1090 |
988 |
0.02 |
|
|
|
| 32 |
A" |
923 |
837 |
0.01 |
|
|
|
| 33 |
A" |
679 |
616 |
0.32 |
|
|
|
| 34 |
A" |
457 |
415 |
0.18 |
|
|
|
| 35 |
A" |
411 |
372 |
0.43 |
|
|
|
| 36 |
A" |
276 |
251 |
21.84 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 27316.6 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 24759.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.288 |
|
|
|
| 2 |
C |
-0.217 |
|
|
|
| 3 |
C |
-0.020 |
|
|
|
| 4 |
C |
-0.166 |
|
|
|
| 5 |
C |
-0.020 |
|
|
|
| 6 |
C |
-0.217 |
|
|
|
| 7 |
N |
-0.354 |
|
|
|
| 8 |
H |
0.063 |
|
|
|
| 9 |
H |
0.089 |
|
|
|
| 10 |
H |
0.087 |
|
|
|
| 11 |
H |
0.089 |
|
|
|
| 12 |
H |
0.063 |
|
|
|
| 13 |
H |
0.159 |
|
|
|
| 14 |
H |
0.159 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.033 |
1.022 |
0.000 |
1.453 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.200 |
-3.381 |
0.000 |
| y |
-3.381 |
-35.737 |
0.000 |
| z |
0.000 |
0.000 |
-36.568 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-11.048 |
-3.381 |
0.000 |
| y |
-3.381 |
6.148 |
0.000 |
| z |
0.000 |
0.000 |
4.900 |
|
| Polar |
|---|
| 3z2-r2 | 9.800 |
|---|
| x2-y2 | -11.463 |
|---|
| xy | -3.381 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.532 |
-0.079 |
0.000 |
| y |
-0.079 |
13.579 |
0.000 |
| z |
0.000 |
0.000 |
11.804 |
<r2> (average value of r
2) Å
2
| <r2> |
189.612 |
| (<r2>)1/2 |
13.770 |