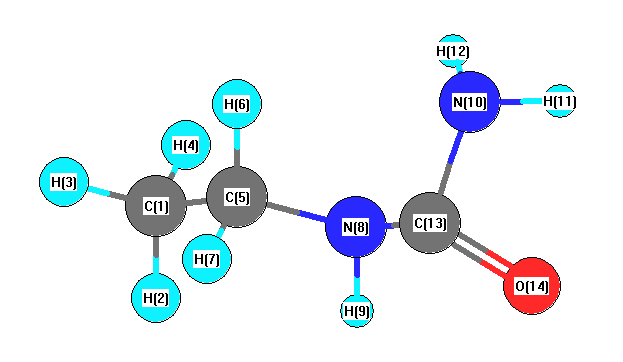
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3945 |
3576 |
47.56 |
|
|
|
| 2 |
A |
3864 |
3502 |
40.25 |
|
|
|
| 3 |
A |
3825 |
3467 |
41.92 |
|
|
|
| 4 |
A |
3238 |
2935 |
39.89 |
|
|
|
| 5 |
A |
3228 |
2926 |
51.84 |
|
|
|
| 6 |
A |
3171 |
2875 |
28.94 |
|
|
|
| 7 |
A |
3166 |
2869 |
19.99 |
|
|
|
| 8 |
A |
3150 |
2855 |
49.62 |
|
|
|
| 9 |
A |
1920 |
1740 |
703.66 |
|
|
|
| 10 |
A |
1775 |
1609 |
175.57 |
|
|
|
| 11 |
A |
1658 |
1503 |
3.82 |
|
|
|
| 12 |
A |
1625 |
1473 |
4.43 |
|
|
|
| 13 |
A |
1611 |
1460 |
7.08 |
|
|
|
| 14 |
A |
1606 |
1456 |
20.81 |
|
|
|
| 15 |
A |
1546 |
1402 |
166.56 |
|
|
|
| 16 |
A |
1542 |
1397 |
160.07 |
|
|
|
| 17 |
A |
1491 |
1351 |
37.50 |
|
|
|
| 18 |
A |
1415 |
1283 |
3.33 |
|
|
|
| 19 |
A |
1297 |
1176 |
18.44 |
|
|
|
| 20 |
A |
1267 |
1148 |
2.32 |
|
|
|
| 21 |
A |
1217 |
1103 |
44.86 |
|
|
|
| 22 |
A |
1150 |
1042 |
13.55 |
|
|
|
| 23 |
A |
1055 |
956 |
3.52 |
|
|
|
| 24 |
A |
964 |
874 |
3.18 |
|
|
|
| 25 |
A |
888 |
805 |
6.65 |
|
|
|
| 26 |
A |
871 |
789 |
55.34 |
|
|
|
| 27 |
A |
637 |
577 |
39.60 |
|
|
|
| 28 |
A |
602 |
545 |
14.28 |
|
|
|
| 29 |
A |
554 |
502 |
109.53 |
|
|
|
| 30 |
A |
506 |
458 |
17.91 |
|
|
|
| 31 |
A |
399 |
362 |
176.38 |
|
|
|
| 32 |
A |
387 |
351 |
8.47 |
|
|
|
| 33 |
A |
289 |
262 |
1.67 |
|
|
|
| 34 |
A |
215 |
194 |
4.23 |
|
|
|
| 35 |
A |
106 |
96 |
5.29 |
|
|
|
| 36 |
A |
66 |
60 |
0.63 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28120.8 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 25488.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.196 |
|
|
|
| 2 |
H |
0.059 |
|
|
|
| 3 |
H |
0.061 |
|
|
|
| 4 |
H |
0.069 |
|
|
|
| 5 |
C |
0.062 |
|
|
|
| 6 |
H |
0.037 |
|
|
|
| 7 |
H |
0.067 |
|
|
|
| 8 |
N |
-0.334 |
|
|
|
| 9 |
H |
0.184 |
|
|
|
| 10 |
N |
-0.409 |
|
|
|
| 11 |
H |
0.201 |
|
|
|
| 12 |
H |
0.173 |
|
|
|
| 13 |
C |
0.556 |
|
|
|
| 14 |
O |
-0.530 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.443 |
3.034 |
-0.521 |
4.619 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.306 |
7.512 |
-1.465 |
| y |
7.512 |
-35.437 |
-1.608 |
| z |
-1.465 |
-1.608 |
-38.136 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.520 |
7.512 |
-1.465 |
| y |
7.512 |
3.284 |
-1.608 |
| z |
-1.465 |
-1.608 |
-0.764 |
|
| Polar |
|---|
| 3z2-r2 | -1.528 |
|---|
| x2-y2 | -3.869 |
|---|
| xy | 7.512 |
|---|
| xz | -1.465 |
|---|
| yz | -1.608 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.132 |
-0.192 |
0.056 |
| y |
-0.192 |
7.670 |
-0.030 |
| z |
0.056 |
-0.030 |
5.882 |
<r2> (average value of r
2) Å
2
| <r2> |
200.353 |
| (<r2>)1/2 |
14.155 |