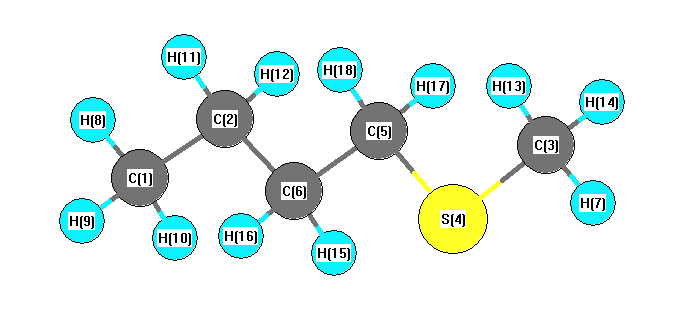
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3261 |
2955 |
14.02 |
|
|
|
| 2 |
A' |
3215 |
2914 |
61.37 |
|
|
|
| 3 |
A' |
3173 |
2876 |
51.96 |
|
|
|
| 4 |
A' |
3171 |
2875 |
50.34 |
|
|
|
| 5 |
A' |
3160 |
2864 |
34.53 |
|
|
|
| 6 |
A' |
3154 |
2859 |
15.00 |
|
|
|
| 7 |
A' |
3145 |
2850 |
23.96 |
|
|
|
| 8 |
A' |
1630 |
1477 |
5.10 |
|
|
|
| 9 |
A' |
1617 |
1466 |
0.72 |
|
|
|
| 10 |
A' |
1610 |
1459 |
1.26 |
|
|
|
| 11 |
A' |
1607 |
1457 |
0.61 |
|
|
|
| 12 |
A' |
1603 |
1453 |
10.02 |
|
|
|
| 13 |
A' |
1539 |
1395 |
2.03 |
|
|
|
| 14 |
A' |
1530 |
1386 |
2.54 |
|
|
|
| 15 |
A' |
1483 |
1345 |
6.43 |
|
|
|
| 16 |
A' |
1459 |
1322 |
4.99 |
|
|
|
| 17 |
A' |
1364 |
1236 |
19.77 |
|
|
|
| 18 |
A' |
1210 |
1097 |
1.29 |
|
|
|
| 19 |
A' |
1127 |
1022 |
1.13 |
|
|
|
| 20 |
A' |
1092 |
990 |
1.47 |
|
|
|
| 21 |
A' |
1055 |
956 |
1.94 |
|
|
|
| 22 |
A' |
961 |
871 |
0.34 |
|
|
|
| 23 |
A' |
828 |
750 |
3.45 |
|
|
|
| 24 |
A' |
776 |
703 |
2.16 |
|
|
|
| 25 |
A' |
451 |
409 |
0.44 |
|
|
|
| 26 |
A' |
336 |
304 |
0.59 |
|
|
|
| 27 |
A' |
255 |
231 |
0.00 |
|
|
|
| 28 |
A' |
125 |
114 |
0.89 |
|
|
|
| 29 |
A" |
3245 |
2941 |
17.71 |
|
|
|
| 30 |
A" |
3221 |
2920 |
125.29 |
|
|
|
| 31 |
A" |
3211 |
2910 |
13.08 |
|
|
|
| 32 |
A" |
3187 |
2889 |
2.96 |
|
|
|
| 33 |
A" |
3163 |
2867 |
4.39 |
|
|
|
| 34 |
A" |
1614 |
1463 |
6.27 |
|
|
|
| 35 |
A" |
1588 |
1440 |
7.52 |
|
|
|
| 36 |
A" |
1440 |
1305 |
0.33 |
|
|
|
| 37 |
A" |
1414 |
1282 |
0.29 |
|
|
|
| 38 |
A" |
1335 |
1210 |
0.03 |
|
|
|
| 39 |
A" |
1167 |
1058 |
0.36 |
|
|
|
| 40 |
A" |
1054 |
955 |
1.47 |
|
|
|
| 41 |
A" |
996 |
903 |
0.42 |
|
|
|
| 42 |
A" |
851 |
772 |
0.01 |
|
|
|
| 43 |
A" |
788 |
714 |
2.53 |
|
|
|
| 44 |
A" |
263 |
238 |
0.03 |
|
|
|
| 45 |
A" |
194 |
176 |
0.56 |
|
|
|
| 46 |
A" |
138 |
125 |
0.05 |
|
|
|
| 47 |
A" |
98 |
89 |
0.75 |
|
|
|
| 48 |
A" |
60 |
54 |
0.93 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 37482.0 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 33973.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.189 |
|
|
|
| 2 |
C |
-0.028 |
|
|
|
| 3 |
C |
-0.220 |
|
|
|
| 4 |
S |
-0.072 |
|
|
|
| 5 |
C |
-0.089 |
|
|
|
| 6 |
C |
-0.058 |
|
|
|
| 7 |
H |
0.090 |
|
|
|
| 8 |
H |
0.061 |
|
|
|
| 9 |
H |
0.050 |
|
|
|
| 10 |
H |
0.050 |
|
|
|
| 11 |
H |
0.029 |
|
|
|
| 12 |
H |
0.029 |
|
|
|
| 13 |
H |
0.074 |
|
|
|
| 14 |
H |
0.074 |
|
|
|
| 15 |
H |
0.047 |
|
|
|
| 16 |
H |
0.047 |
|
|
|
| 17 |
H |
0.052 |
|
|
|
| 18 |
H |
0.052 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.196 |
1.275 |
0.000 |
1.749 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-48.644 |
3.107 |
0.000 |
| y |
3.107 |
-45.240 |
0.000 |
| z |
0.000 |
0.000 |
-48.298 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.875 |
3.107 |
0.000 |
| y |
3.107 |
3.231 |
0.000 |
| z |
0.000 |
0.000 |
-1.356 |
|
| Polar |
|---|
| 3z2-r2 | -2.712 |
|---|
| x2-y2 | -3.404 |
|---|
| xy | 3.107 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.447 |
2.035 |
0.000 |
| y |
2.035 |
12.032 |
0.000 |
| z |
0.000 |
0.000 |
9.360 |
<r2> (average value of r
2) Å
2
| <r2> |
369.996 |
| (<r2>)1/2 |
19.235 |