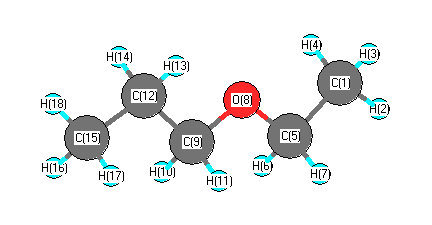
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3227 |
2925 |
49.06 |
|
|
|
| 2 |
A' |
3220 |
2919 |
55.06 |
|
|
|
| 3 |
A' |
3170 |
2873 |
26.82 |
|
|
|
| 4 |
A' |
3167 |
2870 |
24.21 |
|
|
|
| 5 |
A' |
3155 |
2859 |
49.29 |
|
|
|
| 6 |
A' |
3119 |
2827 |
103.13 |
|
|
|
| 7 |
A' |
3101 |
2811 |
11.85 |
|
|
|
| 8 |
A' |
1660 |
1504 |
1.66 |
|
|
|
| 9 |
A' |
1642 |
1488 |
5.84 |
|
|
|
| 10 |
A' |
1627 |
1475 |
4.75 |
|
|
|
| 11 |
A' |
1616 |
1465 |
1.94 |
|
|
|
| 12 |
A' |
1611 |
1460 |
0.83 |
|
|
|
| 13 |
A' |
1591 |
1442 |
1.49 |
|
|
|
| 14 |
A' |
1545 |
1400 |
43.74 |
|
|
|
| 15 |
A' |
1538 |
1394 |
8.37 |
|
|
|
| 16 |
A' |
1517 |
1375 |
17.34 |
|
|
|
| 17 |
A' |
1452 |
1316 |
14.97 |
|
|
|
| 18 |
A' |
1288 |
1167 |
206.45 |
|
|
|
| 19 |
A' |
1273 |
1154 |
94.91 |
|
|
|
| 20 |
A' |
1202 |
1090 |
5.58 |
|
|
|
| 21 |
A' |
1135 |
1029 |
9.60 |
|
|
|
| 22 |
A' |
1094 |
992 |
4.72 |
|
|
|
| 23 |
A' |
980 |
888 |
9.85 |
|
|
|
| 24 |
A' |
960 |
871 |
1.15 |
|
|
|
| 25 |
A' |
529 |
479 |
2.92 |
|
|
|
| 26 |
A' |
433 |
392 |
0.85 |
|
|
|
| 27 |
A' |
328 |
297 |
0.61 |
|
|
|
| 28 |
A' |
150 |
136 |
0.41 |
|
|
|
| 29 |
A" |
3236 |
2933 |
52.55 |
|
|
|
| 30 |
A" |
3219 |
2917 |
103.84 |
|
|
|
| 31 |
A" |
3191 |
2892 |
0.03 |
|
|
|
| 32 |
A" |
3134 |
2841 |
85.16 |
|
|
|
| 33 |
A" |
3125 |
2832 |
24.65 |
|
|
|
| 34 |
A" |
1616 |
1464 |
5.92 |
|
|
|
| 35 |
A" |
1599 |
1449 |
4.75 |
|
|
|
| 36 |
A" |
1431 |
1297 |
1.95 |
|
|
|
| 37 |
A" |
1414 |
1282 |
2.84 |
|
|
|
| 38 |
A" |
1381 |
1252 |
1.53 |
|
|
|
| 39 |
A" |
1304 |
1182 |
8.49 |
|
|
|
| 40 |
A" |
1278 |
1158 |
0.43 |
|
|
|
| 41 |
A" |
970 |
880 |
1.31 |
|
|
|
| 42 |
A" |
881 |
798 |
0.08 |
|
|
|
| 43 |
A" |
817 |
740 |
0.66 |
|
|
|
| 44 |
A" |
276 |
250 |
1.09 |
|
|
|
| 45 |
A" |
250 |
226 |
0.01 |
|
|
|
| 46 |
A" |
165 |
149 |
4.14 |
|
|
|
| 47 |
A" |
97 |
88 |
1.34 |
|
|
|
| 48 |
A" |
73 |
67 |
0.26 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 38390.1 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 34796.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.161 |
|
|
|
| 2 |
H |
0.043 |
|
|
|
| 3 |
H |
0.057 |
|
|
|
| 4 |
H |
0.057 |
|
|
|
| 5 |
C |
0.151 |
|
|
|
| 6 |
H |
0.024 |
|
|
|
| 7 |
H |
0.024 |
|
|
|
| 8 |
O |
-0.394 |
|
|
|
| 9 |
C |
0.136 |
|
|
|
| 10 |
H |
0.023 |
|
|
|
| 11 |
H |
0.023 |
|
|
|
| 12 |
C |
-0.011 |
|
|
|
| 13 |
H |
0.032 |
|
|
|
| 14 |
H |
0.032 |
|
|
|
| 15 |
C |
-0.196 |
|
|
|
| 16 |
H |
0.048 |
|
|
|
| 17 |
H |
0.048 |
|
|
|
| 18 |
H |
0.064 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.963 |
-0.499 |
0.000 |
1.085 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.163 |
1.848 |
0.000 |
| y |
1.848 |
-39.414 |
0.000 |
| z |
0.000 |
0.000 |
-39.484 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.286 |
1.848 |
0.000 |
| y |
1.848 |
-0.090 |
0.000 |
| z |
0.000 |
0.000 |
-0.195 |
|
| Polar |
|---|
| 3z2-r2 | -0.391 |
|---|
| x2-y2 | 0.251 |
|---|
| xy | 1.848 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.073 |
0.883 |
0.000 |
| y |
0.883 |
10.918 |
0.000 |
| z |
0.000 |
0.000 |
8.349 |
<r2> (average value of r
2) Å
2
| <r2> |
284.040 |
| (<r2>)1/2 |
16.853 |