Jump to
S1C2
S1C3
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -266.905468 |
| Energy at 298.15K | |
| HF Energy | -266.905468 |
| Nuclear repulsion energy | 196.740967 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3248 |
2944 |
36.69 |
|
|
|
| 2 |
A |
3173 |
2876 |
93.01 |
|
|
|
| 3 |
A |
3159 |
2863 |
65.78 |
|
|
|
| 4 |
A |
1697 |
1538 |
4.51 |
|
|
|
| 5 |
A |
1655 |
1500 |
0.98 |
|
|
|
| 6 |
A |
1521 |
1379 |
6.99 |
|
|
|
| 7 |
A |
1365 |
1238 |
10.70 |
|
|
|
| 8 |
A |
1342 |
1216 |
76.94 |
|
|
|
| 9 |
A |
1288 |
1167 |
216.32 |
|
|
|
| 10 |
A |
1256 |
1138 |
29.59 |
|
|
|
| 11 |
A |
1064 |
964 |
3.18 |
|
|
|
| 12 |
A |
1023 |
927 |
8.24 |
|
|
|
| 13 |
A |
803 |
728 |
0.58 |
|
|
|
| 14 |
A |
254 |
230 |
0.34 |
|
|
|
| 15 |
B |
3255 |
2951 |
53.76 |
|
|
|
| 16 |
B |
3209 |
2909 |
74.45 |
|
|
|
| 17 |
B |
3166 |
2870 |
87.73 |
|
|
|
| 18 |
B |
1645 |
1491 |
2.84 |
|
|
|
| 19 |
B |
1579 |
1431 |
13.62 |
|
|
|
| 20 |
B |
1480 |
1342 |
1.17 |
|
|
|
| 21 |
B |
1345 |
1219 |
9.42 |
|
|
|
| 22 |
B |
1265 |
1147 |
25.17 |
|
|
|
| 23 |
B |
1211 |
1098 |
57.15 |
|
|
|
| 24 |
B |
1060 |
961 |
70.95 |
|
|
|
| 25 |
B |
978 |
886 |
16.37 |
|
|
|
| 26 |
B |
707 |
641 |
5.46 |
|
|
|
| 27 |
B |
55i |
50i |
18.59 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21845.9 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 19801.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
1.199 |
| C2 |
-0.288 |
0.697 |
-0.939 |
| C3 |
0.288 |
-0.697 |
-0.939 |
| O4 |
0.000 |
1.122 |
0.370 |
| O5 |
0.000 |
-1.122 |
0.370 |
| H6 |
0.889 |
0.030 |
1.814 |
| H7 |
-0.889 |
-0.030 |
1.814 |
| H8 |
-1.360 |
0.679 |
-1.098 |
| H9 |
1.360 |
-0.679 |
-1.098 |
| H10 |
-0.173 |
-1.376 |
-1.638 |
| H11 |
0.173 |
1.376 |
-1.638 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
O5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 2.2671 | 2.2671 | 1.3953 | 1.3953 | 1.0810 | 1.0810 | 2.7543 | 2.7543 | 3.1581 | 3.1581 |
C2 | 2.2671 | | 1.5080 | 1.4062 | 2.2594 | 3.0672 | 2.9097 | 1.0837 | 2.1532 | 2.1907 | 1.0786 | C3 | 2.2671 | 1.5080 | | 2.2594 | 1.4062 | 2.9097 | 3.0672 | 2.1532 | 1.0837 | 1.0786 | 2.1907 | O4 | 1.3953 | 1.4062 | 2.2594 | | 2.2446 | 2.0169 | 2.0500 | 2.0494 | 2.6925 | 3.2101 | 2.0315 | O5 | 1.3953 | 2.2594 | 1.4062 | 2.2446 | | 2.0500 | 2.0169 | 2.6925 | 2.0494 | 2.0315 | 3.2101 | H6 | 1.0810 | 3.0672 | 2.9097 | 2.0169 | 2.0500 | | 1.7783 | 3.7357 | 3.0335 | 3.8757 | 3.7737 | H7 | 1.0810 | 2.9097 | 3.0672 | 2.0500 | 2.0169 | 1.7783 | | 3.0335 | 3.7357 | 3.7737 | 3.8757 | H8 | 2.7543 | 1.0837 | 2.1532 | 2.0494 | 2.6925 | 3.7357 | 3.0335 | | 3.0408 | 2.4346 | 1.7687 | H9 | 2.7543 | 2.1532 | 1.0837 | 2.6925 | 2.0494 | 3.0335 | 3.7357 | 3.0408 | | 1.7687 | 2.4346 | H10 | 3.1581 | 2.1907 | 1.0786 | 3.2101 | 2.0315 | 3.8757 | 3.7737 | 2.4346 | 1.7687 | | 2.7741 | H11 | 3.1581 | 1.0786 | 2.1907 | 2.0315 | 3.2101 | 3.7737 | 3.8757 | 1.7687 | 2.4346 | 2.7741 | |
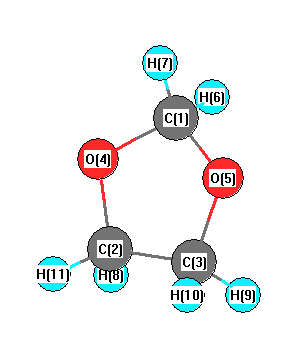 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O4 |
C2 |
108.040 |
|
C1 |
O5 |
C3 |
108.040 |
| C2 |
C3 |
O5 |
101.609 |
|
C2 |
C3 |
H9 |
111.311 |
| C2 |
C3 |
H10 |
114.745 |
|
C3 |
C2 |
O4 |
101.609 |
| C3 |
C2 |
H8 |
111.311 |
|
C3 |
C2 |
H11 |
114.745 |
| O4 |
C1 |
O5 |
107.086 |
|
O4 |
C1 |
H6 |
108.400 |
| O4 |
C1 |
H7 |
111.116 |
|
O4 |
C2 |
H8 |
110.115 |
| O4 |
C2 |
H11 |
108.967 |
|
O5 |
C1 |
H6 |
111.116 |
| O5 |
C1 |
H7 |
108.400 |
|
O5 |
C3 |
H9 |
110.115 |
| O5 |
C3 |
H10 |
108.967 |
|
H6 |
C1 |
H7 |
110.675 |
| H8 |
C2 |
H11 |
109.767 |
|
H9 |
C3 |
H10 |
109.767 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.358 |
|
|
|
| 2 |
C |
0.132 |
|
|
|
| 3 |
C |
0.132 |
|
|
|
| 4 |
O |
-0.428 |
|
|
|
| 5 |
O |
-0.428 |
|
|
|
| 6 |
H |
0.033 |
|
|
|
| 7 |
H |
0.033 |
|
|
|
| 8 |
H |
0.035 |
|
|
|
| 9 |
H |
0.035 |
|
|
|
| 10 |
H |
0.049 |
|
|
|
| 11 |
H |
0.049 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-1.251 |
1.251 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-29.146 |
-0.336 |
0.000 |
| y |
-0.336 |
-34.865 |
0.000 |
| z |
0.000 |
0.000 |
-25.931 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.252 |
-0.336 |
0.000 |
| y |
-0.336 |
-7.326 |
0.000 |
| z |
0.000 |
0.000 |
6.074 |
|
| Polar |
|---|
| 3z2-r2 | 12.149 |
|---|
| x2-y2 | 5.718 |
|---|
| xy | -0.336 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.280 |
-0.102 |
0.000 |
| y |
-0.102 |
5.353 |
0.000 |
| z |
0.000 |
0.000 |
6.502 |
<r2> (average value of r
2) Å
2
| <r2> |
91.818 |
| (<r2>)1/2 |
9.582 |
Jump to
S1C1
S1C3
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -266.905538 |
| Energy at 298.15K | -266.914015 |
| HF Energy | -266.905538 |
| Nuclear repulsion energy | 196.769140 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3262 |
2957 |
44.27 |
|
|
|
| 2 |
A |
3238 |
2935 |
49.20 |
|
|
|
| 3 |
A |
3234 |
2931 |
68.65 |
|
|
|
| 4 |
A |
3180 |
2883 |
58.93 |
|
|
|
| 5 |
A |
3155 |
2859 |
70.90 |
|
|
|
| 6 |
A |
3144 |
2850 |
112.43 |
|
|
|
| 7 |
A |
1693 |
1535 |
4.35 |
|
|
|
| 8 |
A |
1659 |
1504 |
1.30 |
|
|
|
| 9 |
A |
1645 |
1491 |
2.64 |
|
|
|
| 10 |
A |
1576 |
1429 |
14.46 |
|
|
|
| 11 |
A |
1520 |
1378 |
4.24 |
|
|
|
| 12 |
A |
1482 |
1344 |
1.65 |
|
|
|
| 13 |
A |
1380 |
1250 |
8.15 |
|
|
|
| 14 |
A |
1352 |
1226 |
20.77 |
|
|
|
| 15 |
A |
1325 |
1201 |
64.27 |
|
|
|
| 16 |
A |
1303 |
1181 |
132.52 |
|
|
|
| 17 |
A |
1263 |
1145 |
35.23 |
|
|
|
| 18 |
A |
1247 |
1131 |
98.07 |
|
|
|
| 19 |
A |
1206 |
1093 |
55.37 |
|
|
|
| 20 |
A |
1076 |
975 |
29.95 |
|
|
|
| 21 |
A |
1056 |
957 |
50.27 |
|
|
|
| 22 |
A |
1021 |
926 |
11.46 |
|
|
|
| 23 |
A |
964 |
874 |
14.33 |
|
|
|
| 24 |
A |
806 |
730 |
0.44 |
|
|
|
| 25 |
A |
719 |
652 |
5.74 |
|
|
|
| 26 |
A |
255 |
232 |
6.52 |
|
|
|
| 27 |
A |
65 |
59 |
11.71 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21912.7 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 19861.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.192 |
-0.033 |
0.073 |
| C2 |
-0.987 |
-0.694 |
-0.130 |
| C3 |
-0.887 |
0.783 |
0.188 |
| O4 |
0.335 |
-1.137 |
0.061 |
| O5 |
0.415 |
1.070 |
-0.246 |
| H6 |
1.619 |
0.069 |
1.066 |
| H7 |
1.976 |
-0.145 |
-0.660 |
| H8 |
-1.281 |
-0.855 |
-1.160 |
| H9 |
-0.973 |
0.963 |
1.256 |
| H10 |
-1.594 |
1.406 |
-0.337 |
| H11 |
-1.654 |
-1.241 |
0.520 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
O5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 2.2865 | 2.2368 | 1.3979 | 1.3864 | 1.0849 | 1.0790 | 2.8834 | 2.6606 | 3.1621 | 3.1241 |
C2 | 2.2865 | | 1.5136 | 1.4077 | 2.2556 | 2.9666 | 3.0602 | 1.0832 | 2.1597 | 2.1950 | 1.0798 | C3 | 2.2368 | 1.5136 | | 2.2796 | 1.4020 | 2.7496 | 3.1275 | 2.1576 | 1.0859 | 1.0781 | 2.1894 | O4 | 1.3979 | 1.4077 | 2.2796 | | 2.2294 | 2.0272 | 2.0488 | 2.0452 | 2.7474 | 3.2163 | 2.0438 | O5 | 1.3864 | 2.2556 | 1.4020 | 2.2294 | | 2.0425 | 2.0214 | 2.7232 | 2.0472 | 2.0381 | 3.1943 | H6 | 1.0849 | 2.9666 | 2.7496 | 2.0272 | 2.0425 | | 1.7747 | 3.7702 | 2.7484 | 3.7515 | 3.5669 | H7 | 1.0790 | 3.0602 | 3.1275 | 2.0488 | 2.0214 | 1.7747 | | 3.3714 | 3.6872 | 3.9057 | 3.9713 | H8 | 2.8834 | 1.0832 | 2.1576 | 2.0452 | 2.7232 | 3.7702 | 3.3714 | | 3.0389 | 2.4262 | 1.7630 | H9 | 2.6606 | 2.1597 | 1.0859 | 2.7474 | 2.0472 | 2.7484 | 3.6872 | 3.0389 | | 1.7654 | 2.4214 | H10 | 3.1621 | 2.1950 | 1.0781 | 3.2163 | 2.0381 | 3.7515 | 3.9057 | 2.4262 | 1.7654 | | 2.7825 | H11 | 3.1241 | 1.0798 | 2.1894 | 2.0438 | 3.1943 | 3.5669 | 3.9713 | 1.7630 | 2.4214 | 2.7825 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O4 |
C2 |
109.164 |
|
C1 |
O5 |
C3 |
106.680 |
| C2 |
C3 |
O5 |
101.291 |
|
C2 |
C3 |
H9 |
111.296 |
| C2 |
C3 |
H10 |
114.714 |
|
C3 |
C2 |
O4 |
102.521 |
| C3 |
C2 |
H8 |
111.292 |
|
C3 |
C2 |
H11 |
114.130 |
| O4 |
C1 |
O5 |
106.396 |
|
O4 |
C1 |
H6 |
108.816 |
| O4 |
C1 |
H7 |
110.960 |
|
O4 |
C2 |
H8 |
109.698 |
| O4 |
C2 |
H11 |
109.797 |
|
O5 |
C1 |
H6 |
110.891 |
| O5 |
C1 |
H7 |
109.525 |
|
O5 |
C3 |
H9 |
110.095 |
| O5 |
C3 |
H10 |
109.841 |
|
H6 |
C1 |
H7 |
110.193 |
| H8 |
C2 |
H11 |
109.192 |
|
H9 |
C3 |
H10 |
109.330 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.356 |
|
|
|
| 2 |
C |
0.153 |
|
|
|
| 3 |
C |
0.109 |
|
|
|
| 4 |
O |
-0.430 |
|
|
|
| 5 |
O |
-0.422 |
|
|
|
| 6 |
H |
0.023 |
|
|
|
| 7 |
H |
0.044 |
|
|
|
| 8 |
H |
0.034 |
|
|
|
| 9 |
H |
0.038 |
|
|
|
| 10 |
H |
0.057 |
|
|
|
| 11 |
H |
0.037 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.249 |
0.269 |
0.505 |
1.373 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.933 |
-0.470 |
0.263 |
| y |
-0.470 |
-34.612 |
0.896 |
| z |
0.263 |
0.896 |
-29.407 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.077 |
-0.470 |
0.263 |
| y |
-0.470 |
-6.942 |
0.896 |
| z |
0.263 |
0.896 |
0.865 |
|
| Polar |
|---|
| 3z2-r2 | 1.730 |
|---|
| x2-y2 | 8.680 |
|---|
| xy | -0.470 |
|---|
| xz | 0.263 |
|---|
| yz | 0.896 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.484 |
-0.036 |
0.044 |
| y |
-0.036 |
5.390 |
0.091 |
| z |
0.044 |
0.091 |
5.237 |
<r2> (average value of r
2) Å
2
| <r2> |
91.790 |
| (<r2>)1/2 |
9.581 |
Jump to
S1C1
S1C2
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -266.905467 |
| Energy at 298.15K | |
| HF Energy | -266.905467 |
| Nuclear repulsion energy | 196.761073 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3255 |
2950 |
53.37 |
|
|
|
| 2 |
A |
3248 |
2944 |
36.95 |
|
|
|
| 3 |
A |
3207 |
2907 |
74.90 |
|
|
|
| 4 |
A |
3172 |
2875 |
93.43 |
|
|
|
| 5 |
A |
3165 |
2869 |
88.24 |
|
|
|
| 6 |
A |
3157 |
2862 |
65.37 |
|
|
|
| 7 |
A |
1697 |
1538 |
4.51 |
|
|
|
| 8 |
A |
1655 |
1500 |
0.99 |
|
|
|
| 9 |
A |
1646 |
1492 |
2.85 |
|
|
|
| 10 |
A |
1579 |
1431 |
13.65 |
|
|
|
| 11 |
A |
1522 |
1379 |
6.99 |
|
|
|
| 12 |
A |
1481 |
1342 |
1.20 |
|
|
|
| 13 |
A |
1365 |
1238 |
10.77 |
|
|
|
| 14 |
A |
1345 |
1219 |
9.48 |
|
|
|
| 15 |
A |
1342 |
1217 |
78.52 |
|
|
|
| 16 |
A |
1289 |
1168 |
215.32 |
|
|
|
| 17 |
A |
1265 |
1147 |
25.48 |
|
|
|
| 18 |
A |
1256 |
1138 |
29.36 |
|
|
|
| 19 |
A |
1212 |
1099 |
56.41 |
|
|
|
| 20 |
A |
1065 |
965 |
3.26 |
|
|
|
| 21 |
A |
1061 |
962 |
71.40 |
|
|
|
| 22 |
A |
1022 |
927 |
7.84 |
|
|
|
| 23 |
A |
978 |
887 |
16.28 |
|
|
|
| 24 |
A |
803 |
728 |
0.57 |
|
|
|
| 25 |
A |
707 |
641 |
5.46 |
|
|
|
| 26 |
A |
255 |
231 |
0.34 |
|
|
|
| 27 |
A |
56i |
51i |
18.57 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21845.7 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 19800.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.198 |
-0.000 |
0.000 |
| C2 |
-0.938 |
0.730 |
0.187 |
| C3 |
-0.938 |
-0.730 |
-0.187 |
| O4 |
0.369 |
1.110 |
-0.171 |
| O5 |
0.369 |
-1.110 |
0.170 |
| H6 |
1.813 |
-0.105 |
-0.883 |
| H7 |
1.812 |
0.105 |
0.883 |
| H8 |
-1.085 |
0.863 |
1.253 |
| H9 |
-1.085 |
-0.863 |
-1.253 |
| H10 |
-1.642 |
-1.339 |
0.358 |
| H11 |
-1.642 |
1.339 |
-0.358 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
O5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 2.2658 | 2.2658 | 1.3960 | 1.3960 | 1.0808 | 1.0808 | 2.7436 | 2.7438 | 3.1603 | 3.1603 |
C2 | 2.2658 | | 1.5071 | 1.4072 | 2.2566 | 3.0676 | 2.9054 | 1.0837 | 2.1519 | 2.1921 | 1.0784 | C3 | 2.2658 | 1.5071 | | 2.2566 | 1.4072 | 2.9057 | 3.0675 | 2.1519 | 1.0837 | 1.0784 | 2.1921 | O4 | 1.3960 | 1.4072 | 2.2566 | | 2.2452 | 2.0169 | 2.0504 | 2.0494 | 2.6784 | 3.2120 | 2.0320 | O5 | 1.3960 | 2.2566 | 1.4072 | 2.2452 | | 2.0504 | 2.0169 | 2.6785 | 2.0493 | 2.0320 | 3.2120 | H6 | 1.0808 | 3.0676 | 2.9057 | 2.0169 | 2.0504 | | 1.7789 | 3.7275 | 3.0181 | 3.8724 | 3.7807 | H7 | 1.0808 | 2.9054 | 3.0675 | 2.0504 | 2.0169 | 1.7789 | | 3.0175 | 3.7276 | 3.7804 | 3.8722 | H8 | 2.7436 | 1.0837 | 2.1519 | 2.0494 | 2.6785 | 3.7275 | 3.0175 | | 3.0418 | 2.4409 | 1.7694 | H9 | 2.7438 | 2.1519 | 1.0837 | 2.6784 | 2.0493 | 3.0181 | 3.7276 | 3.0418 | | 1.7694 | 2.4409 | H10 | 3.1603 | 2.1921 | 1.0784 | 3.2120 | 2.0320 | 3.8724 | 3.7804 | 2.4409 | 1.7694 | | 2.7724 | H11 | 3.1603 | 1.0784 | 2.1921 | 2.0320 | 3.2120 | 3.7807 | 3.8722 | 1.7694 | 2.4409 | 2.7724 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O4 |
C2 |
107.855 |
|
C1 |
O5 |
C3 |
107.858 |
| C2 |
C3 |
O5 |
101.433 |
|
C2 |
C3 |
H9 |
111.267 |
| C2 |
C3 |
H10 |
114.946 |
|
C3 |
C2 |
O4 |
101.432 |
| C3 |
C2 |
H8 |
111.267 |
|
C3 |
C2 |
H11 |
114.947 |
| O4 |
C1 |
O5 |
107.056 |
|
O4 |
C1 |
H6 |
108.369 |
| O4 |
C1 |
H7 |
111.118 |
|
O4 |
C2 |
H8 |
110.040 |
| O4 |
C2 |
H11 |
108.956 |
|
O5 |
C1 |
H6 |
111.118 |
| O5 |
C1 |
H7 |
108.369 |
|
O5 |
C3 |
H9 |
110.040 |
| O5 |
C3 |
H10 |
108.956 |
|
H6 |
C1 |
H7 |
110.760 |
| H8 |
C2 |
H11 |
109.843 |
|
H9 |
C3 |
H10 |
109.843 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.358 |
|
|
|
| 2 |
C |
0.132 |
|
|
|
| 3 |
C |
0.132 |
|
|
|
| 4 |
O |
-0.428 |
|
|
|
| 5 |
O |
-0.428 |
|
|
|
| 6 |
H |
0.033 |
|
|
|
| 7 |
H |
0.033 |
|
|
|
| 8 |
H |
0.035 |
|
|
|
| 9 |
H |
0.035 |
|
|
|
| 10 |
H |
0.049 |
|
|
|
| 11 |
H |
0.049 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.249 |
0.000 |
0.000 |
1.249 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.940 |
-0.000 |
0.000 |
| y |
-0.000 |
-34.663 |
1.091 |
| z |
0.000 |
1.091 |
-29.344 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.064 |
-0.000 |
0.000 |
| y |
-0.000 |
-7.021 |
1.091 |
| z |
0.000 |
1.091 |
0.957 |
|
| Polar |
|---|
| 3z2-r2 | 1.914 |
|---|
| x2-y2 | 8.724 |
|---|
| xy | -0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 1.091 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.500 |
0.000 |
0.000 |
| y |
0.000 |
5.380 |
0.089 |
| z |
0.000 |
0.089 |
5.256 |
<r2> (average value of r
2) Å
2
| <r2> |
91.804 |
| (<r2>)1/2 |
9.581 |
