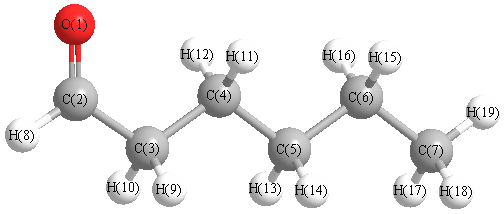
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3214 |
2913 |
61.93 |
|
|
|
| 2 |
A' |
3176 |
2878 |
37.42 |
|
|
|
| 3 |
A' |
3154 |
2858 |
31.66 |
|
|
|
| 4 |
A' |
3148 |
2853 |
52.46 |
|
|
|
| 5 |
A' |
3143 |
2849 |
47.97 |
|
|
|
| 6 |
A' |
3132 |
2838 |
8.62 |
|
|
|
| 7 |
A' |
3070 |
2783 |
121.99 |
|
|
|
| 8 |
A' |
1993 |
1806 |
226.31 |
|
|
|
| 9 |
A' |
1631 |
1478 |
8.03 |
|
|
|
| 10 |
A' |
1620 |
1468 |
1.29 |
|
|
|
| 11 |
A' |
1609 |
1458 |
0.60 |
|
|
|
| 12 |
A' |
1607 |
1456 |
0.51 |
|
|
|
| 13 |
A' |
1577 |
1430 |
10.85 |
|
|
|
| 14 |
A' |
1550 |
1405 |
4.75 |
|
|
|
| 15 |
A' |
1540 |
1396 |
4.53 |
|
|
|
| 16 |
A' |
1536 |
1393 |
14.91 |
|
|
|
| 17 |
A' |
1522 |
1380 |
4.78 |
|
|
|
| 18 |
A' |
1458 |
1322 |
9.09 |
|
|
|
| 19 |
A' |
1373 |
1245 |
1.05 |
|
|
|
| 20 |
A' |
1224 |
1109 |
6.24 |
|
|
|
| 21 |
A' |
1131 |
1025 |
2.30 |
|
|
|
| 22 |
A' |
1128 |
1022 |
4.41 |
|
|
|
| 23 |
A' |
1087 |
985 |
1.11 |
|
|
|
| 24 |
A' |
982 |
890 |
6.08 |
|
|
|
| 25 |
A' |
936 |
848 |
4.21 |
|
|
|
| 26 |
A' |
752 |
681 |
20.06 |
|
|
|
| 27 |
A' |
441 |
400 |
5.41 |
|
|
|
| 28 |
A' |
384 |
348 |
1.04 |
|
|
|
| 29 |
A' |
237 |
215 |
5.09 |
|
|
|
| 30 |
A' |
117 |
106 |
4.39 |
|
|
|
| 31 |
A" |
3216 |
2915 |
149.74 |
|
|
|
| 32 |
A" |
3208 |
2908 |
12.31 |
|
|
|
| 33 |
A" |
3174 |
2877 |
10.73 |
|
|
|
| 34 |
A" |
3160 |
2864 |
14.05 |
|
|
|
| 35 |
A" |
3149 |
2854 |
4.59 |
|
|
|
| 36 |
A" |
1615 |
1463 |
6.27 |
|
|
|
| 37 |
A" |
1442 |
1307 |
0.44 |
|
|
|
| 38 |
A" |
1438 |
1304 |
0.02 |
|
|
|
| 39 |
A" |
1400 |
1269 |
0.09 |
|
|
|
| 40 |
A" |
1336 |
1211 |
0.01 |
|
|
|
| 41 |
A" |
1276 |
1157 |
0.00 |
|
|
|
| 42 |
A" |
1082 |
981 |
0.09 |
|
|
|
| 43 |
A" |
979 |
887 |
0.55 |
|
|
|
| 44 |
A" |
856 |
776 |
0.05 |
|
|
|
| 45 |
A" |
786 |
712 |
1.77 |
|
|
|
| 46 |
A" |
728 |
660 |
1.58 |
|
|
|
| 47 |
A" |
263 |
239 |
0.00 |
|
|
|
| 48 |
A" |
186 |
168 |
2.57 |
|
|
|
| 49 |
A" |
142 |
129 |
0.57 |
|
|
|
| 50 |
A" |
104 |
94 |
2.98 |
|
|
|
| 51 |
A" |
58 |
53 |
2.72 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 39532.6 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 35832.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.399 |
|
|
|
| 2 |
C |
0.246 |
|
|
|
| 3 |
C |
-0.041 |
|
|
|
| 4 |
C |
-0.037 |
|
|
|
| 5 |
C |
-0.053 |
|
|
|
| 6 |
C |
-0.022 |
|
|
|
| 7 |
C |
-0.189 |
|
|
|
| 8 |
H |
0.031 |
|
|
|
| 9 |
H |
0.049 |
|
|
|
| 10 |
H |
0.049 |
|
|
|
| 11 |
H |
0.046 |
|
|
|
| 12 |
H |
0.046 |
|
|
|
| 13 |
H |
0.030 |
|
|
|
| 14 |
H |
0.030 |
|
|
|
| 15 |
H |
0.030 |
|
|
|
| 16 |
H |
0.030 |
|
|
|
| 17 |
H |
0.047 |
|
|
|
| 18 |
H |
0.047 |
|
|
|
| 19 |
H |
0.062 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.459 |
-2.466 |
0.000 |
2.866 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.514 |
-1.498 |
0.000 |
| y |
-1.498 |
-58.114 |
0.000 |
| z |
0.000 |
0.000 |
-43.950 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
10.517 |
-1.498 |
0.000 |
| y |
-1.498 |
-15.882 |
0.000 |
| z |
0.000 |
0.000 |
5.364 |
|
| Polar |
|---|
| 3z2-r2 | 10.729 |
|---|
| x2-y2 | 17.599 |
|---|
| xy | -1.498 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.033 |
-0.996 |
0.000 |
| y |
-0.996 |
11.813 |
0.000 |
| z |
0.000 |
0.000 |
8.722 |
<r2> (average value of r
2) Å
2
| <r2> |
391.312 |
| (<r2>)1/2 |
19.782 |