Jump to
S1C2
S2C1
S2C2
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -192.042538 |
| Energy at 298.15K | -192.048657 |
| HF Energy | -192.042538 |
| Nuclear repulsion energy | 120.504869 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3279 |
2972 |
13.28 |
73.78 |
0.52 |
0.69 |
| 2 |
A |
3216 |
2915 |
0.00 |
7.72 |
0.75 |
0.86 |
| 3 |
A |
3172 |
2875 |
12.11 |
276.31 |
0.01 |
0.02 |
| 4 |
A |
1981 |
1796 |
259.42 |
12.36 |
0.66 |
0.80 |
| 5 |
A |
1586 |
1438 |
26.43 |
12.82 |
0.69 |
0.82 |
| 6 |
A |
1586 |
1437 |
0.22 |
11.75 |
0.75 |
0.86 |
| 7 |
A |
1518 |
1376 |
9.23 |
1.63 |
0.67 |
0.81 |
| 8 |
A |
1178 |
1067 |
0.58 |
2.75 |
0.44 |
0.61 |
| 9 |
A |
966 |
875 |
0.00 |
1.15 |
0.75 |
0.86 |
| 10 |
A |
838 |
760 |
0.19 |
15.48 |
0.12 |
0.21 |
| 11 |
A |
402 |
364 |
1.84 |
0.31 |
0.47 |
0.64 |
| 12 |
A |
46 |
42 |
0.00 |
0.05 |
0.75 |
0.86 |
| 13 |
B |
3277 |
2970 |
18.76 |
50.83 |
0.75 |
0.86 |
| 14 |
B |
3226 |
2924 |
36.24 |
95.31 |
0.75 |
0.86 |
| 15 |
B |
3164 |
2868 |
2.59 |
0.21 |
0.75 |
0.86 |
| 16 |
B |
1608 |
1457 |
18.03 |
0.01 |
0.75 |
0.86 |
| 17 |
B |
1578 |
1430 |
0.16 |
0.67 |
0.75 |
0.86 |
| 18 |
B |
1527 |
1384 |
50.20 |
0.03 |
0.75 |
0.86 |
| 19 |
B |
1337 |
1211 |
71.47 |
1.76 |
0.75 |
0.86 |
| 20 |
B |
1229 |
1114 |
2.60 |
0.26 |
0.75 |
0.86 |
| 21 |
B |
967 |
877 |
5.39 |
2.56 |
0.75 |
0.86 |
| 22 |
B |
577 |
523 |
21.92 |
1.53 |
0.75 |
0.86 |
| 23 |
B |
533 |
483 |
2.25 |
0.96 |
0.75 |
0.86 |
| 24 |
B |
149 |
135 |
0.35 |
0.01 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 19469.1 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 17646.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.189 |
| O2 |
0.000 |
0.000 |
1.375 |
| C3 |
0.000 |
1.285 |
-0.606 |
| C4 |
0.000 |
-1.285 |
-0.606 |
| H5 |
-0.002 |
2.132 |
0.063 |
| H6 |
0.002 |
-2.132 |
0.063 |
| H7 |
0.875 |
1.326 |
-1.245 |
| H8 |
-0.872 |
1.324 |
-1.249 |
| H9 |
-0.875 |
-1.326 |
-1.245 |
| H10 |
0.872 |
-1.324 |
-1.249 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.1861 | 1.5109 | 1.5109 | 2.1357 | 2.1357 | 2.1403 | 2.1406 | 2.1403 | 2.1406 |
O2 | 1.1861 | | 2.3609 | 2.3609 | 2.5033 | 2.5033 | 3.0643 | 3.0660 | 3.0643 | 3.0660 | C3 | 1.5109 | 2.3609 | | 2.5705 | 1.0789 | 3.4821 | 1.0845 | 1.0845 | 2.8275 | 2.8257 | C4 | 1.5109 | 2.3609 | 2.5705 | | 3.4821 | 1.0789 | 2.8275 | 2.8257 | 1.0845 | 1.0845 | H5 | 2.1357 | 2.5033 | 1.0789 | 3.4821 | | 4.2641 | 1.7693 | 1.7694 | 3.7991 | 3.7991 | H6 | 2.1357 | 2.5033 | 3.4821 | 1.0789 | 4.2641 | | 3.7991 | 3.7991 | 1.7693 | 1.7694 | H7 | 2.1403 | 3.0643 | 1.0845 | 2.8275 | 1.7693 | 3.7991 | | 1.7464 | 3.1776 | 2.6507 | H8 | 2.1406 | 3.0660 | 1.0845 | 2.8257 | 1.7694 | 3.7991 | 1.7464 | | 2.6507 | 3.1711 | H9 | 2.1403 | 3.0643 | 2.8275 | 1.0845 | 3.7991 | 1.7693 | 3.1776 | 2.6507 | | 1.7464 | H10 | 2.1406 | 3.0660 | 2.8257 | 1.0845 | 3.7991 | 1.7694 | 2.6507 | 3.1711 | 1.7464 | |
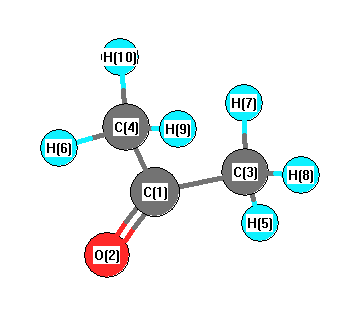 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
H5 |
109.989 |
|
C1 |
C3 |
H7 |
110.026 |
| C1 |
C3 |
H8 |
110.049 |
|
C1 |
C4 |
H6 |
109.989 |
| C1 |
C4 |
H9 |
110.026 |
|
C1 |
C4 |
H10 |
110.049 |
| O2 |
C1 |
C3 |
121.717 |
|
O2 |
C1 |
C4 |
121.717 |
| C3 |
C1 |
C4 |
116.567 |
|
H5 |
C3 |
H7 |
109.733 |
| H5 |
C3 |
H8 |
109.744 |
|
H6 |
C4 |
H9 |
109.733 |
| H6 |
C4 |
H10 |
109.744 |
|
H7 |
C3 |
H8 |
107.258 |
| H9 |
C4 |
H10 |
107.258 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.332 |
|
|
|
| 2 |
O |
-0.416 |
|
|
|
| 3 |
C |
-0.186 |
|
|
|
| 4 |
C |
-0.186 |
|
|
|
| 5 |
H |
0.102 |
|
|
|
| 6 |
H |
0.102 |
|
|
|
| 7 |
H |
0.063 |
|
|
|
| 8 |
H |
0.063 |
|
|
|
| 9 |
H |
0.063 |
|
|
|
| 10 |
H |
0.063 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-3.208 |
3.208 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.196 |
0.001 |
0.000 |
| y |
0.001 |
-23.588 |
0.000 |
| z |
0.000 |
0.000 |
-29.348 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.272 |
0.001 |
0.000 |
| y |
0.001 |
3.184 |
0.000 |
| z |
0.000 |
0.000 |
-5.456 |
|
| Polar |
|---|
| 3z2-r2 | -10.912 |
|---|
| x2-y2 | -0.608 |
|---|
| xy | 0.001 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.541 |
0.000 |
0.000 |
| y |
0.000 |
5.769 |
0.000 |
| z |
0.000 |
0.000 |
6.106 |
<r2> (average value of r
2) Å
2
| <r2> |
81.016 |
| (<r2>)1/2 |
9.001 |
Jump to
S1C1
S2C1
S2C2
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -192.042538 |
| Energy at 298.15K | -192.048656 |
| HF Energy | -192.042538 |
| Nuclear repulsion energy | 120.504299 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3279 |
2972 |
13.29 |
|
|
|
| 2 |
A |
3216 |
2915 |
0.00 |
|
|
|
| 3 |
A |
3172 |
2875 |
12.11 |
|
|
|
| 4 |
A |
1981 |
1796 |
259.41 |
|
|
|
| 5 |
A |
1586 |
1438 |
26.59 |
|
|
|
| 6 |
A |
1586 |
1437 |
0.05 |
|
|
|
| 7 |
A |
1518 |
1376 |
9.23 |
|
|
|
| 8 |
A |
1178 |
1067 |
0.57 |
|
|
|
| 9 |
A |
966 |
875 |
0.00 |
|
|
|
| 10 |
A |
838 |
760 |
0.19 |
|
|
|
| 11 |
A |
402 |
364 |
1.84 |
|
|
|
| 12 |
A |
46 |
41 |
0.00 |
|
|
|
| 13 |
B |
3277 |
2970 |
18.74 |
|
|
|
| 14 |
B |
3226 |
2924 |
36.25 |
|
|
|
| 15 |
B |
3164 |
2868 |
2.59 |
|
|
|
| 16 |
B |
1608 |
1457 |
18.03 |
|
|
|
| 17 |
B |
1578 |
1430 |
0.16 |
|
|
|
| 18 |
B |
1527 |
1384 |
50.18 |
|
|
|
| 19 |
B |
1336 |
1211 |
71.49 |
|
|
|
| 20 |
B |
1229 |
1114 |
2.60 |
|
|
|
| 21 |
B |
967 |
877 |
5.39 |
|
|
|
| 22 |
B |
577 |
523 |
21.92 |
|
|
|
| 23 |
B |
533 |
483 |
2.25 |
|
|
|
| 24 |
B |
149 |
135 |
0.35 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19468.9 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 17646.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.188 |
| O2 |
0.000 |
0.000 |
1.374 |
| C3 |
0.000 |
1.285 |
-0.606 |
| C4 |
0.000 |
-1.285 |
-0.606 |
| H5 |
0.144 |
2.127 |
0.054 |
| H6 |
-0.144 |
-2.127 |
0.054 |
| H7 |
0.776 |
1.268 |
-1.362 |
| H8 |
-0.950 |
1.389 |
-1.119 |
| H9 |
-0.776 |
-1.268 |
-1.362 |
| H10 |
0.950 |
-1.389 |
-1.119 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.1860 | 1.5106 | 1.5106 | 2.1362 | 2.1362 | 2.1478 | 2.1310 | 2.1478 | 2.1310 |
O2 | 1.1860 | | 2.3603 | 2.3603 | 2.5077 | 2.5077 | 3.1140 | 3.0082 | 3.1140 | 3.0082 | C3 | 1.5106 | 2.3603 | | 2.5704 | 1.0790 | 3.4784 | 1.0838 | 1.0849 | 2.7737 | 2.8840 | C4 | 1.5106 | 2.3603 | 2.5704 | | 3.4784 | 1.0790 | 2.7737 | 2.8840 | 1.0838 | 1.0849 | H5 | 2.1362 | 2.5077 | 1.0790 | 3.4784 | | 4.2639 | 1.7725 | 1.7658 | 3.7918 | 3.7932 | H6 | 2.1362 | 2.5077 | 3.4784 | 1.0790 | 4.2639 | | 3.7918 | 3.7932 | 1.7725 | 1.7658 | H7 | 2.1478 | 3.1140 | 1.0838 | 2.7737 | 1.7725 | 3.7918 | | 1.7470 | 2.9733 | 2.6737 | H8 | 2.1310 | 3.0082 | 1.0849 | 2.8840 | 1.7658 | 3.7932 | 1.7470 | | 2.6737 | 3.3653 | H9 | 2.1478 | 3.1140 | 2.7737 | 1.0838 | 3.7918 | 1.7725 | 2.9733 | 2.6737 | | 1.7470 | H10 | 2.1310 | 3.0082 | 2.8840 | 1.0849 | 3.7932 | 1.7658 | 2.6737 | 3.3653 | 1.7470 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
H5 |
110.042 |
|
C1 |
C3 |
H7 |
110.687 |
| C1 |
C3 |
H8 |
109.281 |
|
C1 |
C4 |
H6 |
110.042 |
| C1 |
C4 |
H9 |
110.687 |
|
C1 |
C4 |
H10 |
109.281 |
| O2 |
C1 |
C3 |
121.699 |
|
O2 |
C1 |
C4 |
121.699 |
| C3 |
C1 |
C4 |
116.601 |
|
H5 |
C3 |
H7 |
110.075 |
| H5 |
C3 |
H8 |
109.376 |
|
H6 |
C4 |
H9 |
110.075 |
| H6 |
C4 |
H10 |
109.376 |
|
H7 |
C3 |
H8 |
107.327 |
| H9 |
C4 |
H10 |
107.327 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.332 |
|
|
|
| 2 |
O |
-0.416 |
|
|
|
| 3 |
C |
-0.186 |
|
|
|
| 4 |
C |
-0.186 |
|
|
|
| 5 |
H |
0.102 |
|
|
|
| 6 |
H |
0.102 |
|
|
|
| 7 |
H |
0.063 |
|
|
|
| 8 |
H |
0.063 |
|
|
|
| 9 |
H |
0.063 |
|
|
|
| 10 |
H |
0.063 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-3.208 |
3.208 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.196 |
-0.000 |
0.000 |
| y |
-0.000 |
-23.587 |
0.000 |
| z |
0.000 |
0.000 |
-29.348 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.272 |
-0.000 |
0.000 |
| y |
-0.000 |
3.185 |
0.000 |
| z |
0.000 |
0.000 |
-5.456 |
|
| Polar |
|---|
| 3z2-r2 | -10.913 |
|---|
| x2-y2 | -0.609 |
|---|
| xy | -0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.541 |
-0.000 |
0.000 |
| y |
-0.000 |
5.770 |
0.000 |
| z |
0.000 |
0.000 |
6.106 |
<r2> (average value of r
2) Å
2
| <r2> |
81.017 |
| (<r2>)1/2 |
9.001 |
Jump to
S1C1
S1C2
S2C2
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -191.957031 |
| Energy at 298.15K | -191.963185 |
| HF Energy | -191.957031 |
| Nuclear repulsion energy | 117.741041 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3246 |
2942 |
36.67 |
|
|
|
| 2 |
A' |
3210 |
2909 |
32.37 |
|
|
|
| 3 |
A' |
3129 |
2837 |
48.96 |
|
|
|
| 4 |
A' |
1615 |
1464 |
4.46 |
|
|
|
| 5 |
A' |
1600 |
1450 |
9.32 |
|
|
|
| 6 |
A' |
1535 |
1391 |
2.98 |
|
|
|
| 7 |
A' |
1401 |
1270 |
49.48 |
|
|
|
| 8 |
A' |
1202 |
1090 |
0.01 |
|
|
|
| 9 |
A' |
1057 |
958 |
10.71 |
|
|
|
| 10 |
A' |
842 |
763 |
6.20 |
|
|
|
| 11 |
A' |
419 |
380 |
0.45 |
|
|
|
| 12 |
A' |
347 |
315 |
0.55 |
|
|
|
| 13 |
A' |
201 |
182 |
0.34 |
|
|
|
| 14 |
A" |
3243 |
2939 |
11.91 |
|
|
|
| 15 |
A" |
3208 |
2907 |
14.37 |
|
|
|
| 16 |
A" |
3125 |
2832 |
24.55 |
|
|
|
| 17 |
A" |
1595 |
1445 |
0.07 |
|
|
|
| 18 |
A" |
1586 |
1438 |
3.63 |
|
|
|
| 19 |
A" |
1520 |
1377 |
2.31 |
|
|
|
| 20 |
A" |
1288 |
1168 |
7.94 |
|
|
|
| 21 |
A" |
1048 |
950 |
4.03 |
|
|
|
| 22 |
A" |
1028 |
932 |
0.00 |
|
|
|
| 23 |
A" |
413 |
374 |
8.24 |
|
|
|
| 24 |
A" |
182 |
165 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19018.2 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 17238.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.182 |
0.258 |
0.000 |
| O2 |
-0.544 |
1.266 |
0.000 |
| C3 |
0.182 |
-0.581 |
1.295 |
| C4 |
0.182 |
-0.581 |
-1.295 |
| H5 |
-0.826 |
-0.662 |
1.691 |
| H6 |
-0.826 |
-0.662 |
-1.691 |
| H7 |
0.563 |
-1.572 |
1.125 |
| H8 |
0.803 |
-0.117 |
2.055 |
| H9 |
0.563 |
-1.572 |
-1.125 |
| H10 |
0.803 |
-0.117 |
-2.055 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.2417 | 1.5437 | 1.5437 | 2.1733 | 2.1733 | 2.1821 | 2.1794 | 2.1821 | 2.1794 |
O2 | 1.2417 | | 2.3701 | 2.3701 | 2.5804 | 2.5804 | 3.2473 | 2.8198 | 3.2473 | 2.8198 | C3 | 1.5437 | 2.3701 | | 2.5905 | 1.0856 | 3.1529 | 1.0747 | 1.0855 | 2.6431 | 3.4386 | C4 | 1.5437 | 2.3701 | 2.5905 | | 3.1529 | 1.0856 | 2.6431 | 3.4386 | 1.0747 | 1.0855 | H5 | 2.1733 | 2.5804 | 1.0856 | 3.1529 | | 3.3825 | 1.7538 | 1.7559 | 3.2695 | 4.1211 | H6 | 2.1733 | 2.5804 | 3.1529 | 1.0856 | 3.3825 | | 3.2695 | 4.1211 | 1.7538 | 1.7559 | H7 | 2.1821 | 3.2473 | 1.0747 | 2.6431 | 1.7538 | 3.2695 | | 1.7425 | 2.2509 | 3.5052 | H8 | 2.1794 | 2.8198 | 1.0855 | 3.4386 | 1.7559 | 4.1211 | 1.7425 | | 3.5052 | 4.1094 | H9 | 2.1821 | 3.2473 | 2.6431 | 1.0747 | 3.2695 | 1.7538 | 2.2509 | 3.5052 | | 1.7425 | H10 | 2.1794 | 2.8198 | 3.4386 | 1.0855 | 4.1211 | 1.7559 | 3.5052 | 4.1094 | 1.7425 | |
More geometry information
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.161 |
|
|
|
| 2 |
O |
-0.231 |
|
|
|
| 3 |
C |
-0.160 |
|
|
|
| 4 |
C |
-0.160 |
|
|
|
| 5 |
H |
0.057 |
|
|
|
| 6 |
H |
0.057 |
|
|
|
| 7 |
H |
0.059 |
|
|
|
| 8 |
H |
0.079 |
|
|
|
| 9 |
H |
0.059 |
|
|
|
| 10 |
H |
0.079 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.549 |
-1.652 |
0.000 |
1.741 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.329 |
0.482 |
0.000 |
| y |
0.482 |
-27.191 |
0.000 |
| z |
0.000 |
0.000 |
-23.365 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.050 |
0.482 |
0.000 |
| y |
0.482 |
-2.344 |
0.000 |
| z |
0.000 |
0.000 |
3.394 |
|
| Polar |
|---|
| 3z2-r2 | 6.789 |
|---|
| x2-y2 | 0.863 |
|---|
| xy | 0.482 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.082 |
-0.280 |
0.000 |
| y |
-0.280 |
5.527 |
0.000 |
| z |
0.000 |
0.000 |
5.993 |
<r2> (average value of r
2) Å
2
| <r2> |
82.398 |
| (<r2>)1/2 |
9.077 |
Jump to
S1C1
S1C2
S2C1
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -191.957031 |
| Energy at 298.15K | -191.963185 |
| HF Energy | -191.957031 |
| Nuclear repulsion energy | 117.741041 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
Geometric Data calculated at HF/Def2TZVPP
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
