Jump to
S1C2
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -748.324603 |
| Energy at 298.15K | |
| HF Energy | -748.324603 |
| Nuclear repulsion energy | 570.408081 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
2039 |
1849 |
0.00 |
|
|
|
| 2 |
Ag |
1559 |
1413 |
0.00 |
|
|
|
| 3 |
Ag |
1464 |
1327 |
0.00 |
|
|
|
| 4 |
Ag |
1298 |
1177 |
0.00 |
|
|
|
| 5 |
Ag |
784 |
711 |
0.00 |
|
|
|
| 6 |
Ag |
645 |
585 |
0.00 |
|
|
|
| 7 |
Ag |
409 |
371 |
0.00 |
|
|
|
| 8 |
Ag |
386 |
350 |
0.00 |
|
|
|
| 9 |
Ag |
239 |
216 |
0.00 |
|
|
|
| 10 |
Au |
691 |
627 |
6.55 |
|
|
|
| 11 |
Au |
388 |
351 |
5.04 |
|
|
|
| 12 |
Au |
124 |
112 |
0.10 |
|
|
|
| 13 |
Au |
17i |
15i |
0.16 |
|
|
|
| 14 |
Bg |
762 |
691 |
0.00 |
|
|
|
| 15 |
Bg |
537 |
486 |
0.00 |
|
|
|
| 16 |
Bg |
204 |
185 |
0.00 |
|
|
|
| 17 |
Bu |
1961 |
1777 |
537.87 |
|
|
|
| 18 |
Bu |
1497 |
1357 |
468.42 |
|
|
|
| 19 |
Bu |
1334 |
1209 |
447.43 |
|
|
|
| 20 |
Bu |
1057 |
958 |
357.55 |
|
|
|
| 21 |
Bu |
689 |
624 |
7.87 |
|
|
|
| 22 |
Bu |
549 |
498 |
3.57 |
|
|
|
| 23 |
Bu |
318 |
288 |
6.97 |
|
|
|
| 24 |
Bu |
146 |
133 |
0.83 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 9530.7 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 8638.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.154 |
1.838 |
0.000 |
| C2 |
0.463 |
0.565 |
0.000 |
| C3 |
-0.463 |
-0.565 |
0.000 |
| C4 |
-0.154 |
-1.838 |
0.000 |
| F5 |
1.039 |
2.775 |
0.000 |
| F6 |
-1.039 |
2.322 |
0.000 |
| F7 |
1.743 |
0.245 |
0.000 |
| F8 |
-1.743 |
-0.245 |
0.000 |
| F9 |
1.039 |
-2.322 |
0.000 |
| F10 |
-1.039 |
-2.775 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
F5 |
F6 |
F7 |
F8 |
F9 |
F10 |
| C1 | | 1.3101 | 2.4801 | 3.6881 | 1.2890 | 1.2874 | 2.2505 | 2.8169 | 4.2530 | 4.7641 |
C2 | 1.3101 | | 1.4604 | 2.4801 | 2.2840 | 2.3120 | 1.3197 | 2.3502 | 2.9437 | 3.6615 | C3 | 2.4801 | 1.4604 | | 1.3101 | 3.6615 | 2.9437 | 2.3502 | 1.3197 | 2.3120 | 2.2840 | C4 | 3.6881 | 2.4801 | 1.3101 | | 4.7641 | 4.2530 | 2.8169 | 2.2505 | 1.2874 | 1.2890 | F5 | 1.2890 | 2.2840 | 3.6615 | 4.7641 | | 2.1264 | 2.6265 | 4.1058 | 5.0970 | 5.9257 | F6 | 1.2874 | 2.3120 | 2.9437 | 4.2530 | 2.1264 | | 3.4725 | 2.6618 | 5.0880 | 5.0970 | F7 | 2.2505 | 1.3197 | 2.3502 | 2.8169 | 2.6265 | 3.4725 | | 3.5210 | 2.6618 | 4.1058 | F8 | 2.8169 | 2.3502 | 1.3197 | 2.2505 | 4.1058 | 2.6618 | 3.5210 | | 3.4725 | 2.6265 | F9 | 4.2530 | 2.9437 | 2.3120 | 1.2874 | 5.0970 | 5.0880 | 2.6618 | 3.4725 | | 2.1264 | F10 | 4.7641 | 3.6615 | 2.2840 | 1.2890 | 5.9257 | 5.0970 | 4.1058 | 2.6265 | 2.1264 | |
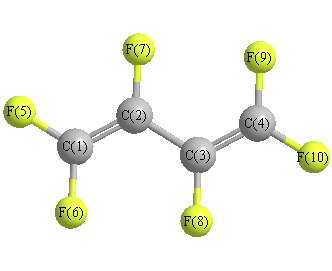 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
126.983 |
|
C1 |
C2 |
F7 |
117.689 |
| C2 |
C1 |
F5 |
122.982 |
|
C2 |
C1 |
F6 |
125.771 |
| C2 |
C3 |
C4 |
126.983 |
|
C2 |
C3 |
F8 |
115.329 |
| C3 |
C2 |
F7 |
115.329 |
|
C3 |
C4 |
F9 |
125.771 |
| C3 |
C4 |
F10 |
122.982 |
|
C4 |
C3 |
F8 |
117.689 |
| F5 |
C1 |
F6 |
111.247 |
|
F9 |
C4 |
F10 |
111.247 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.532 |
|
|
|
| 2 |
C |
0.133 |
|
|
|
| 3 |
C |
0.133 |
|
|
|
| 4 |
C |
0.532 |
|
|
|
| 5 |
F |
-0.212 |
|
|
|
| 6 |
F |
-0.208 |
|
|
|
| 7 |
F |
-0.244 |
|
|
|
| 8 |
F |
-0.244 |
|
|
|
| 9 |
F |
-0.208 |
|
|
|
| 10 |
F |
-0.212 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-56.441 |
-0.324 |
0.000 |
| y |
-0.324 |
-53.135 |
0.000 |
| z |
0.000 |
0.000 |
-48.938 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.405 |
-0.324 |
0.000 |
| y |
-0.324 |
-0.445 |
0.000 |
| z |
0.000 |
0.000 |
5.850 |
|
| Polar |
|---|
| 3z2-r2 | 11.700 |
|---|
| x2-y2 | -3.307 |
|---|
| xy | -0.324 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.114 |
-0.281 |
0.000 |
| y |
-0.281 |
11.349 |
0.000 |
| z |
0.000 |
0.000 |
4.382 |
<r2> (average value of r
2) Å
2
| <r2> |
410.092 |
| (<r2>)1/2 |
20.251 |
Jump to
S1C1
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -748.327791 |
| Energy at 298.15K | -748.329536 |
| HF Energy | -748.327791 |
| Nuclear repulsion energy | 572.839751 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
2024 |
1834 |
92.87 |
|
|
|
| 2 |
A |
1528 |
1385 |
0.47 |
|
|
|
| 3 |
A |
1476 |
1338 |
240.69 |
|
|
|
| 4 |
A |
1259 |
1141 |
344.18 |
|
|
|
| 5 |
A |
793 |
719 |
0.00 |
|
|
|
| 6 |
A |
770 |
698 |
1.83 |
|
|
|
| 7 |
A |
606 |
549 |
0.23 |
|
|
|
| 8 |
A |
527 |
478 |
0.56 |
|
|
|
| 9 |
A |
411 |
372 |
1.03 |
|
|
|
| 10 |
A |
286 |
260 |
0.21 |
|
|
|
| 11 |
A |
206 |
187 |
0.44 |
|
|
|
| 12 |
A |
105 |
95 |
0.32 |
|
|
|
| 13 |
A |
48 |
44 |
0.00 |
|
|
|
| 14 |
B |
1997 |
1810 |
333.79 |
|
|
|
| 15 |
B |
1494 |
1354 |
313.15 |
|
|
|
| 16 |
B |
1321 |
1198 |
177.48 |
|
|
|
| 17 |
B |
1081 |
980 |
236.07 |
|
|
|
| 18 |
B |
745 |
676 |
5.68 |
|
|
|
| 19 |
B |
693 |
628 |
9.94 |
|
|
|
| 20 |
B |
626 |
568 |
3.50 |
|
|
|
| 21 |
B |
475 |
431 |
4.60 |
|
|
|
| 22 |
B |
326 |
295 |
5.57 |
|
|
|
| 23 |
B |
229 |
208 |
3.82 |
|
|
|
| 24 |
B |
120 |
109 |
0.31 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 9573.5 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 8677.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.181 |
1.579 |
-0.356 |
| C2 |
0.163 |
0.710 |
0.558 |
| C3 |
-0.163 |
-0.710 |
0.558 |
| C4 |
0.181 |
-1.579 |
-0.356 |
| F5 |
0.163 |
2.819 |
-0.341 |
| F6 |
-0.872 |
1.297 |
-1.405 |
| F7 |
0.843 |
1.132 |
1.611 |
| F8 |
-0.843 |
-1.132 |
1.611 |
| F9 |
0.872 |
-1.297 |
-1.405 |
| F10 |
-0.163 |
-2.819 |
-0.341 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
F5 |
F6 |
F7 |
F8 |
F9 |
F10 |
| C1 | | 1.3066 | 2.4649 | 3.1786 | 1.2867 | 1.2880 | 2.2618 | 3.4141 | 3.2378 | 4.3979 |
C2 | 1.3066 | | 1.4578 | 2.4649 | 2.2918 | 2.2956 | 1.3225 | 2.3487 | 2.8959 | 3.6563 | C3 | 2.4649 | 1.4578 | | 1.3066 | 3.6563 | 2.8959 | 2.3487 | 1.3225 | 2.2956 | 2.2918 | C4 | 3.1786 | 2.4649 | 1.3066 | | 4.3979 | 3.2378 | 3.4141 | 2.2618 | 1.2880 | 1.2867 | F5 | 1.2867 | 2.2918 | 3.6563 | 4.3979 | | 2.1261 | 2.6676 | 4.5199 | 4.3104 | 5.6471 | F6 | 1.2880 | 2.2956 | 2.8959 | 3.2378 | 2.1261 | | 3.4737 | 3.8728 | 3.1267 | 4.3104 | F7 | 2.2618 | 1.3225 | 2.3487 | 3.4141 | 2.6676 | 3.4737 | | 2.8231 | 3.8728 | 4.5199 | F8 | 3.4141 | 2.3487 | 1.3225 | 2.2618 | 4.5199 | 3.8728 | 2.8231 | | 3.4737 | 2.6676 | F9 | 3.2378 | 2.8959 | 2.2956 | 1.2880 | 4.3104 | 3.1267 | 3.8728 | 3.4737 | | 2.1261 | F10 | 4.3979 | 3.6563 | 2.2918 | 1.2867 | 5.6471 | 4.3104 | 4.5199 | 2.6676 | 2.1261 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
126.080 |
|
C1 |
C2 |
F7 |
118.691 |
| C2 |
C1 |
F5 |
124.194 |
|
C2 |
C1 |
F6 |
124.445 |
| C2 |
C3 |
C4 |
126.080 |
|
C2 |
C3 |
F8 |
115.206 |
| C3 |
C2 |
F7 |
115.206 |
|
C3 |
C4 |
F9 |
124.445 |
| C3 |
C4 |
F10 |
124.194 |
|
C4 |
C3 |
F8 |
118.691 |
| F5 |
C1 |
F6 |
111.331 |
|
F9 |
C4 |
F10 |
111.331 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.497 |
|
|
|
| 2 |
C |
0.178 |
|
|
|
| 3 |
C |
0.178 |
|
|
|
| 4 |
C |
0.497 |
|
|
|
| 5 |
F |
-0.210 |
|
|
|
| 6 |
F |
-0.216 |
|
|
|
| 7 |
F |
-0.248 |
|
|
|
| 8 |
F |
-0.248 |
|
|
|
| 9 |
F |
-0.216 |
|
|
|
| 10 |
F |
-0.210 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.996 |
0.996 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-51.074 |
-1.827 |
0.000 |
| y |
-1.827 |
-53.279 |
0.000 |
| z |
0.000 |
0.000 |
-54.597 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.864 |
-1.827 |
0.000 |
| y |
-1.827 |
-0.444 |
0.000 |
| z |
0.000 |
0.000 |
-2.420 |
|
| Polar |
|---|
| 3z2-r2 | -4.840 |
|---|
| x2-y2 | 2.205 |
|---|
| xy | -1.827 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.011 |
-0.248 |
0.000 |
| y |
-0.248 |
8.634 |
0.000 |
| z |
0.000 |
0.000 |
6.809 |
<r2> (average value of r
2) Å
2
| <r2> |
381.431 |
| (<r2>)1/2 |
19.530 |
