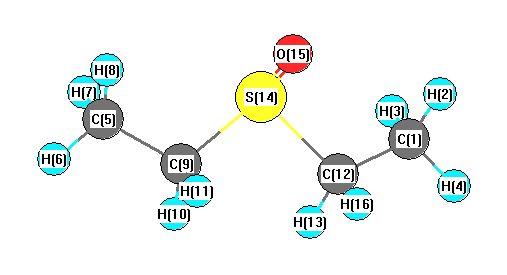
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3263 |
2958 |
21.57 |
|
|
|
| 2 |
A |
3257 |
2952 |
18.92 |
|
|
|
| 3 |
A |
3238 |
2935 |
4.45 |
|
|
|
| 4 |
A |
3234 |
2932 |
68.56 |
|
|
|
| 5 |
A |
3229 |
2927 |
11.84 |
|
|
|
| 6 |
A |
3223 |
2921 |
6.37 |
|
|
|
| 7 |
A |
3202 |
2902 |
2.73 |
|
|
|
| 8 |
A |
3192 |
2893 |
13.93 |
|
|
|
| 9 |
A |
3175 |
2878 |
30.07 |
|
|
|
| 10 |
A |
3171 |
2874 |
21.74 |
|
|
|
| 11 |
A |
1625 |
1473 |
2.44 |
|
|
|
| 12 |
A |
1620 |
1469 |
5.88 |
|
|
|
| 13 |
A |
1617 |
1466 |
2.45 |
|
|
|
| 14 |
A |
1615 |
1464 |
9.32 |
|
|
|
| 15 |
A |
1595 |
1446 |
7.02 |
|
|
|
| 16 |
A |
1582 |
1434 |
4.45 |
|
|
|
| 17 |
A |
1539 |
1395 |
2.82 |
|
|
|
| 18 |
A |
1533 |
1390 |
1.96 |
|
|
|
| 19 |
A |
1446 |
1311 |
0.59 |
|
|
|
| 20 |
A |
1420 |
1287 |
6.43 |
|
|
|
| 21 |
A |
1378 |
1249 |
1.33 |
|
|
|
| 22 |
A |
1362 |
1234 |
0.40 |
|
|
|
| 23 |
A |
1197 |
1085 |
147.70 |
|
|
|
| 24 |
A |
1179 |
1069 |
22.41 |
|
|
|
| 25 |
A |
1147 |
1039 |
11.08 |
|
|
|
| 26 |
A |
1137 |
1031 |
17.42 |
|
|
|
| 27 |
A |
1105 |
1002 |
1.63 |
|
|
|
| 28 |
A |
1049 |
951 |
1.54 |
|
|
|
| 29 |
A |
1038 |
941 |
3.39 |
|
|
|
| 30 |
A |
851 |
772 |
8.07 |
|
|
|
| 31 |
A |
815 |
739 |
8.44 |
|
|
|
| 32 |
A |
761 |
690 |
13.11 |
|
|
|
| 33 |
A |
672 |
609 |
2.08 |
|
|
|
| 34 |
A |
508 |
461 |
9.15 |
|
|
|
| 35 |
A |
410 |
372 |
5.04 |
|
|
|
| 36 |
A |
343 |
311 |
6.12 |
|
|
|
| 37 |
A |
290 |
263 |
1.46 |
|
|
|
| 38 |
A |
229 |
207 |
3.82 |
|
|
|
| 39 |
A |
224 |
203 |
6.21 |
|
|
|
| 40 |
A |
220 |
199 |
1.57 |
|
|
|
| 41 |
A |
126 |
114 |
2.79 |
|
|
|
| 42 |
A |
70 |
64 |
0.08 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31944.5 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 28954.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.209 |
|
|
|
| 2 |
H |
0.071 |
|
|
|
| 3 |
H |
0.045 |
|
|
|
| 4 |
H |
0.103 |
|
|
|
| 5 |
C |
-0.215 |
|
|
|
| 6 |
H |
0.071 |
|
|
|
| 7 |
H |
0.075 |
|
|
|
| 8 |
H |
0.058 |
|
|
|
| 9 |
C |
-0.115 |
|
|
|
| 10 |
H |
0.089 |
|
|
|
| 11 |
H |
0.071 |
|
|
|
| 12 |
C |
-0.121 |
|
|
|
| 13 |
H |
0.059 |
|
|
|
| 14 |
S |
0.628 |
|
|
|
| 15 |
O |
-0.700 |
|
|
|
| 16 |
H |
0.091 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.061 |
3.836 |
-0.918 |
4.450 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-43.307 |
4.549 |
-2.909 |
| y |
4.549 |
-51.006 |
1.279 |
| z |
-2.909 |
1.279 |
-45.785 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.088 |
4.549 |
-2.909 |
| y |
4.549 |
-6.459 |
1.279 |
| z |
-2.909 |
1.279 |
1.371 |
|
| Polar |
|---|
| 3z2-r2 | 2.743 |
|---|
| x2-y2 | 7.698 |
|---|
| xy | 4.549 |
|---|
| xz | -2.909 |
|---|
| yz | 1.279 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.007 |
0.068 |
-0.239 |
| y |
0.068 |
9.957 |
-0.205 |
| z |
-0.239 |
-0.205 |
8.849 |
<r2> (average value of r
2) Å
2
| <r2> |
221.203 |
| (<r2>)1/2 |
14.873 |