Jump to
S1C2
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -193.198933 |
| Energy at 298.15K | -193.208068 |
| HF Energy | -193.198933 |
| Nuclear repulsion energy | 131.326387 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
4184 |
3792 |
57.09 |
|
|
|
| 2 |
A' |
3220 |
2919 |
55.28 |
|
|
|
| 3 |
A' |
3169 |
2873 |
27.82 |
|
|
|
| 4 |
A' |
3155 |
2859 |
48.80 |
|
|
|
| 5 |
A' |
3125 |
2833 |
57.69 |
|
|
|
| 6 |
A' |
1651 |
1497 |
3.74 |
|
|
|
| 7 |
A' |
1628 |
1476 |
5.53 |
|
|
|
| 8 |
A' |
1611 |
1460 |
0.89 |
|
|
|
| 9 |
A' |
1590 |
1441 |
4.03 |
|
|
|
| 10 |
A' |
1539 |
1395 |
0.58 |
|
|
|
| 11 |
A' |
1469 |
1331 |
8.90 |
|
|
|
| 12 |
A' |
1355 |
1229 |
75.33 |
|
|
|
| 13 |
A' |
1186 |
1075 |
29.68 |
|
|
|
| 14 |
A' |
1164 |
1055 |
85.84 |
|
|
|
| 15 |
A' |
1098 |
995 |
3.68 |
|
|
|
| 16 |
A' |
946 |
857 |
4.39 |
|
|
|
| 17 |
A' |
488 |
443 |
10.80 |
|
|
|
| 18 |
A' |
296 |
268 |
4.19 |
|
|
|
| 19 |
A" |
3219 |
2918 |
115.22 |
|
|
|
| 20 |
A" |
3191 |
2892 |
0.07 |
|
|
|
| 21 |
A" |
3145 |
2851 |
45.58 |
|
|
|
| 22 |
A" |
1615 |
1464 |
6.03 |
|
|
|
| 23 |
A" |
1430 |
1297 |
0.01 |
|
|
|
| 24 |
A" |
1377 |
1248 |
0.39 |
|
|
|
| 25 |
A" |
1290 |
1169 |
2.45 |
|
|
|
| 26 |
A" |
968 |
878 |
1.97 |
|
|
|
| 27 |
A" |
816 |
740 |
0.28 |
|
|
|
| 28 |
A" |
282 |
256 |
120.27 |
|
|
|
| 29 |
A" |
246 |
223 |
6.94 |
|
|
|
| 30 |
A" |
138 |
125 |
6.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25295.7 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 22928.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.442 |
1.227 |
0.000 |
| C2 |
0.000 |
0.739 |
0.000 |
| C3 |
0.102 |
-0.771 |
0.000 |
| O4 |
1.459 |
-1.130 |
0.000 |
| H5 |
-1.484 |
2.308 |
0.000 |
| H6 |
-1.977 |
0.877 |
0.876 |
| H7 |
-1.977 |
0.877 |
-0.876 |
| H8 |
0.523 |
1.120 |
0.870 |
| H9 |
0.523 |
1.120 |
-0.870 |
| H10 |
-0.391 |
-1.182 |
0.877 |
| H11 |
-0.391 |
-1.182 |
-0.877 |
| H12 |
1.542 |
-2.064 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.5221 | 2.5251 | 3.7381 | 1.0824 | 1.0844 | 1.0844 | 2.1518 | 2.1518 | 2.7705 | 2.7705 | 4.4428 |
C2 | 1.5221 | | 1.5142 | 2.3717 | 2.1597 | 2.1671 | 2.1671 | 1.0840 | 1.0840 | 2.1479 | 2.1479 | 3.2000 | C3 | 2.5251 | 1.5142 | | 1.4041 | 3.4641 | 2.7943 | 2.7943 | 2.1238 | 2.1238 | 1.0867 | 1.0867 | 1.9357 | O4 | 3.7381 | 2.3717 | 1.4041 | | 4.5263 | 4.0753 | 4.0753 | 2.5875 | 2.5875 | 2.0484 | 2.0484 | 0.9378 | H5 | 1.0824 | 2.1597 | 3.4641 | 4.5263 | | 1.7489 | 1.7489 | 2.4897 | 2.4897 | 3.7611 | 3.7611 | 5.3179 | H6 | 1.0844 | 2.1671 | 2.7943 | 4.0753 | 1.7489 | | 1.7517 | 2.5125 | 3.0593 | 2.5993 | 3.1352 | 4.6700 | H7 | 1.0844 | 2.1671 | 2.7943 | 4.0753 | 1.7489 | 1.7517 | | 3.0593 | 2.5125 | 3.1352 | 2.5993 | 4.6700 | H8 | 2.1518 | 1.0840 | 2.1238 | 2.5875 | 2.4897 | 2.5125 | 3.0593 | | 1.7395 | 2.4766 | 3.0307 | 3.4545 | H9 | 2.1518 | 1.0840 | 2.1238 | 2.5875 | 2.4897 | 3.0593 | 2.5125 | 1.7395 | | 3.0307 | 2.4766 | 3.4545 | H10 | 2.7705 | 2.1479 | 1.0867 | 2.0484 | 3.7611 | 2.5993 | 3.1352 | 2.4766 | 3.0307 | | 1.7545 | 2.2992 | H11 | 2.7705 | 2.1479 | 1.0867 | 2.0484 | 3.7611 | 3.1352 | 2.5993 | 3.0307 | 2.4766 | 1.7545 | | 2.2992 | H12 | 4.4428 | 3.2000 | 1.9357 | 0.9378 | 5.3179 | 4.6700 | 4.6700 | 3.4545 | 3.4545 | 2.2992 | 2.2992 | |
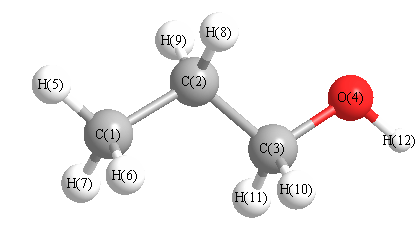 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
112.532 |
|
C1 |
C2 |
H8 |
110.182 |
| C1 |
C2 |
H9 |
110.182 |
|
C2 |
C1 |
H5 |
110.902 |
| C2 |
C1 |
H6 |
111.378 |
|
C2 |
C1 |
H7 |
111.378 |
| C2 |
C3 |
O4 |
108.663 |
|
C2 |
C3 |
H10 |
110.264 |
| C2 |
C3 |
H11 |
110.264 |
|
C3 |
C2 |
H8 |
108.519 |
| C3 |
C2 |
H9 |
108.519 |
|
C3 |
O4 |
H12 |
109.893 |
| O4 |
C3 |
H10 |
109.996 |
|
O4 |
C3 |
H11 |
109.996 |
| H5 |
C1 |
H6 |
107.635 |
|
H5 |
C1 |
H7 |
107.635 |
| H6 |
C1 |
H7 |
107.732 |
|
H8 |
C2 |
H9 |
106.715 |
| H10 |
C3 |
H11 |
107.655 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.192 |
|
|
|
| 2 |
C |
-0.009 |
|
|
|
| 3 |
C |
0.140 |
|
|
|
| 4 |
O |
-0.414 |
|
|
|
| 5 |
H |
0.063 |
|
|
|
| 6 |
H |
0.047 |
|
|
|
| 7 |
H |
0.047 |
|
|
|
| 8 |
H |
0.032 |
|
|
|
| 9 |
H |
0.032 |
|
|
|
| 10 |
H |
0.028 |
|
|
|
| 11 |
H |
0.028 |
|
|
|
| 12 |
H |
0.195 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.191 |
-0.984 |
0.000 |
1.545 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-29.849 |
-0.911 |
0.000 |
| y |
-0.911 |
-21.959 |
0.000 |
| z |
0.000 |
0.000 |
-26.608 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.565 |
-0.911 |
0.000 |
| y |
-0.911 |
6.270 |
0.000 |
| z |
0.000 |
0.000 |
-0.704 |
|
| Polar |
|---|
| 3z2-r2 | -1.409 |
|---|
| x2-y2 | -7.890 |
|---|
| xy | -0.911 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.223 |
-0.508 |
0.000 |
| y |
-0.508 |
6.265 |
0.000 |
| z |
0.000 |
0.000 |
5.397 |
<r2> (average value of r
2) Å
2
| <r2> |
106.349 |
| (<r2>)1/2 |
10.313 |
Jump to
S1C1
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -193.198735 |
| Energy at 298.15K | |
| HF Energy | -193.198735 |
| Nuclear repulsion energy | 133.398804 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
4187 |
3795 |
58.96 |
89.13 |
0.26 |
0.41 |
| 2 |
A |
3241 |
2938 |
43.42 |
34.83 |
0.70 |
0.82 |
| 3 |
A |
3212 |
2911 |
83.97 |
57.06 |
0.48 |
0.65 |
| 4 |
A |
3189 |
2890 |
31.37 |
82.91 |
0.68 |
0.81 |
| 5 |
A |
3166 |
2870 |
63.48 |
207.42 |
0.04 |
0.08 |
| 6 |
A |
3157 |
2861 |
73.82 |
120.13 |
0.16 |
0.27 |
| 7 |
A |
3147 |
2852 |
10.44 |
73.14 |
0.68 |
0.81 |
| 8 |
A |
3126 |
2833 |
61.87 |
93.97 |
0.15 |
0.27 |
| 9 |
A |
1649 |
1495 |
2.41 |
4.12 |
0.73 |
0.85 |
| 10 |
A |
1625 |
1473 |
7.82 |
2.05 |
0.72 |
0.83 |
| 11 |
A |
1611 |
1460 |
6.25 |
8.27 |
0.74 |
0.85 |
| 12 |
A |
1600 |
1450 |
1.13 |
10.27 |
0.72 |
0.84 |
| 13 |
A |
1584 |
1435 |
5.64 |
1.64 |
0.02 |
0.04 |
| 14 |
A |
1539 |
1395 |
2.48 |
0.48 |
0.62 |
0.76 |
| 15 |
A |
1500 |
1360 |
1.86 |
0.84 |
0.75 |
0.86 |
| 16 |
A |
1430 |
1296 |
24.60 |
7.76 |
0.70 |
0.82 |
| 17 |
A |
1382 |
1253 |
2.59 |
4.19 |
0.73 |
0.85 |
| 18 |
A |
1348 |
1222 |
59.70 |
0.82 |
0.65 |
0.79 |
| 19 |
A |
1262 |
1144 |
10.36 |
1.15 |
0.10 |
0.18 |
| 20 |
A |
1211 |
1098 |
13.90 |
4.54 |
0.61 |
0.76 |
| 21 |
A |
1149 |
1041 |
62.08 |
2.75 |
0.57 |
0.72 |
| 22 |
A |
1064 |
964 |
24.27 |
5.33 |
0.75 |
0.86 |
| 23 |
A |
997 |
903 |
2.32 |
0.61 |
0.72 |
0.83 |
| 24 |
A |
911 |
825 |
1.48 |
13.27 |
0.11 |
0.20 |
| 25 |
A |
833 |
755 |
0.82 |
0.73 |
0.28 |
0.44 |
| 26 |
A |
511 |
463 |
9.02 |
0.22 |
0.57 |
0.73 |
| 27 |
A |
347 |
315 |
7.38 |
0.27 |
0.23 |
0.38 |
| 28 |
A |
266 |
241 |
118.67 |
1.92 |
0.74 |
0.85 |
| 29 |
A |
241 |
219 |
3.92 |
0.05 |
0.48 |
0.65 |
| 30 |
A |
150 |
136 |
7.32 |
0.12 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 25315.3 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 22945.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.550 |
-0.508 |
0.125 |
| C2 |
-0.629 |
0.635 |
-0.287 |
| C3 |
0.768 |
0.530 |
0.293 |
| O4 |
1.390 |
-0.623 |
-0.213 |
| H5 |
-2.535 |
-0.385 |
-0.310 |
| H6 |
-1.667 |
-0.546 |
1.203 |
| H7 |
-1.154 |
-1.461 |
-0.201 |
| H8 |
-0.550 |
0.675 |
-1.368 |
| H9 |
-1.053 |
1.583 |
0.034 |
| H10 |
0.715 |
0.486 |
1.379 |
| H11 |
1.341 |
1.414 |
0.026 |
| H12 |
2.247 |
-0.720 |
0.156 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.5249 | 2.5453 | 2.9609 | 1.0838 | 1.0853 | 1.0816 | 2.1512 | 2.1514 | 2.7732 | 3.4729 | 3.8026 |
C2 | 1.5249 | | 1.5163 | 2.3801 | 2.1616 | 2.1658 | 2.1630 | 1.0849 | 1.0860 | 2.1455 | 2.1412 | 3.2099 | C3 | 2.5453 | 1.5163 | | 1.4039 | 3.4797 | 2.8131 | 2.8113 | 2.1253 | 2.1194 | 1.0881 | 1.0869 | 1.9407 | O4 | 2.9609 | 2.3801 | 1.4039 | | 3.9327 | 3.3693 | 2.6784 | 2.6041 | 3.3003 | 2.0537 | 2.0517 | 0.9383 | H5 | 1.0838 | 2.1616 | 3.4797 | 3.9327 | | 1.7519 | 1.7537 | 2.4861 | 2.4871 | 3.7646 | 4.2859 | 4.8159 | H6 | 1.0853 | 2.1658 | 2.8131 | 3.3693 | 1.7519 | | 1.7528 | 3.0573 | 2.5049 | 2.6015 | 3.7778 | 4.0551 | H7 | 1.0816 | 2.1630 | 2.8113 | 2.6784 | 1.7537 | 1.7528 | | 2.5079 | 3.0546 | 3.1275 | 3.8137 | 3.4994 | H8 | 2.1512 | 1.0849 | 2.1253 | 2.6041 | 2.4861 | 3.0573 | 2.5079 | | 1.7439 | 3.0300 | 2.4626 | 3.4771 | H9 | 2.1514 | 1.0860 | 2.1194 | 3.3003 | 2.4871 | 2.5049 | 3.0546 | 1.7439 | | 2.4776 | 2.3996 | 4.0254 | H10 | 2.7732 | 2.1455 | 1.0881 | 2.0537 | 3.7646 | 2.6015 | 3.1275 | 3.0300 | 2.4776 | | 1.7561 | 2.3010 | H11 | 3.4729 | 2.1412 | 1.0869 | 2.0517 | 4.2859 | 3.7778 | 3.8137 | 2.4626 | 2.3996 | 1.7561 | | 2.3217 | H12 | 3.8026 | 3.2099 | 1.9407 | 0.9383 | 4.8159 | 4.0551 | 3.4994 | 3.4771 | 4.0254 | 2.3010 | 2.3217 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
113.636 |
|
C1 |
C2 |
H8 |
109.888 |
| C1 |
C2 |
H9 |
109.834 |
|
C2 |
C1 |
H5 |
110.784 |
| C2 |
C1 |
H6 |
111.026 |
|
C2 |
C1 |
H7 |
111.025 |
| C2 |
C3 |
O4 |
109.117 |
|
C2 |
C3 |
H10 |
109.836 |
| C2 |
C3 |
H11 |
109.571 |
|
C3 |
C2 |
H8 |
108.435 |
| C3 |
C2 |
H9 |
107.916 |
|
C3 |
O4 |
H12 |
110.323 |
| O4 |
C3 |
H10 |
110.354 |
|
O4 |
C3 |
H11 |
110.265 |
| H5 |
C1 |
H6 |
107.736 |
|
H5 |
C1 |
H7 |
108.164 |
| H6 |
C1 |
H7 |
107.974 |
|
H8 |
C2 |
H9 |
106.890 |
| H10 |
C3 |
H11 |
107.681 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.180 |
|
|
|
| 2 |
C |
-0.021 |
|
|
|
| 3 |
C |
0.132 |
|
|
|
| 4 |
O |
-0.417 |
|
|
|
| 5 |
H |
0.056 |
|
|
|
| 6 |
H |
0.038 |
|
|
|
| 7 |
H |
0.074 |
|
|
|
| 8 |
H |
0.033 |
|
|
|
| 9 |
H |
0.022 |
|
|
|
| 10 |
H |
0.028 |
|
|
|
| 11 |
H |
0.040 |
|
|
|
| 12 |
H |
0.194 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.709 |
0.986 |
1.003 |
1.575 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.952 |
-0.138 |
2.005 |
| y |
-0.138 |
-27.317 |
-0.576 |
| z |
2.005 |
-0.576 |
-26.878 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.145 |
-0.138 |
2.005 |
| y |
-0.138 |
-2.402 |
-0.576 |
| z |
2.005 |
-0.576 |
-1.743 |
|
| Polar |
|---|
| 3z2-r2 | -3.486 |
|---|
| x2-y2 | 4.365 |
|---|
| xy | -0.138 |
|---|
| xz | 2.005 |
|---|
| yz | -0.576 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.467 |
0.076 |
0.084 |
| y |
0.076 |
5.851 |
0.001 |
| z |
0.084 |
0.001 |
5.543 |
<r2> (average value of r
2) Å
2
| <r2> |
95.042 |
| (<r2>)1/2 |
9.749 |
