Jump to
S1C2
Vibrational Frequencies calculated at QCISD/Def2TZVPP
Geometric Data calculated at QCISD/Def2TZVPP
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -194.012297 |
| Energy at 298.15K | |
| HF Energy | -193.197742 |
| Nuclear repulsion energy | 132.985602 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3904 |
3904 |
31.80 |
|
|
|
| 2 |
A |
3146 |
3146 |
23.73 |
|
|
|
| 3 |
A |
3122 |
3122 |
47.42 |
|
|
|
| 4 |
A |
3098 |
3098 |
17.62 |
|
|
|
| 5 |
A |
3065 |
3065 |
57.88 |
|
|
|
| 6 |
A |
3054 |
3054 |
36.83 |
|
|
|
| 7 |
A |
3049 |
3049 |
9.65 |
|
|
|
| 8 |
A |
3017 |
3017 |
53.17 |
|
|
|
| 9 |
A |
1543 |
1543 |
2.31 |
|
|
|
| 10 |
A |
1528 |
1528 |
6.60 |
|
|
|
| 11 |
A |
1514 |
1514 |
6.32 |
|
|
|
| 12 |
A |
1495 |
1495 |
2.21 |
|
|
|
| 13 |
A |
1480 |
1480 |
3.58 |
|
|
|
| 14 |
A |
1434 |
1434 |
3.08 |
|
|
|
| 15 |
A |
1397 |
1397 |
0.97 |
|
|
|
| 16 |
A |
1342 |
1342 |
20.45 |
|
|
|
| 17 |
A |
1291 |
1291 |
0.90 |
|
|
|
| 18 |
A |
1267 |
1267 |
38.40 |
|
|
|
| 19 |
A |
1179 |
1179 |
6.20 |
|
|
|
| 20 |
A |
1142 |
1142 |
16.90 |
|
|
|
| 21 |
A |
1093 |
1093 |
35.17 |
|
|
|
| 22 |
A |
1003 |
1003 |
37.52 |
|
|
|
| 23 |
A |
935 |
935 |
1.95 |
|
|
|
| 24 |
A |
883 |
883 |
1.25 |
|
|
|
| 25 |
A |
780 |
780 |
0.73 |
|
|
|
| 26 |
A |
479 |
479 |
7.92 |
|
|
|
| 27 |
A |
327 |
327 |
6.73 |
|
|
|
| 28 |
A |
250 |
250 |
97.32 |
|
|
|
| 29 |
A |
227 |
227 |
6.95 |
|
|
|
| 30 |
A |
138 |
138 |
8.93 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24091.4 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 24091.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/Def2TZVPP
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.532 |
-0.517 |
0.128 |
| C2 |
-0.633 |
0.643 |
-0.291 |
| C3 |
0.762 |
0.544 |
0.295 |
| O4 |
1.378 |
-0.633 |
-0.218 |
| H5 |
-2.527 |
-0.420 |
-0.306 |
| H6 |
-1.642 |
-0.548 |
1.214 |
| H7 |
-1.109 |
-1.467 |
-0.194 |
| H8 |
-0.548 |
0.677 |
-1.379 |
| H9 |
-1.070 |
1.592 |
0.028 |
| H10 |
0.701 |
0.499 |
1.388 |
| H11 |
1.345 |
1.430 |
0.024 |
| H12 |
2.244 |
-0.717 |
0.179 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.5259 | 2.5330 | 2.9324 | 1.0902 | 1.0920 | 1.0887 | 2.1605 | 2.1616 | 2.7580 | 3.4748 | 3.7813 |
C2 | 1.5259 | | 1.5169 | 2.3824 | 2.1712 | 2.1682 | 2.1647 | 1.0920 | 1.0927 | 2.1498 | 2.1519 | 3.2167 | C3 | 2.5330 | 1.5169 | | 1.4232 | 3.4797 | 2.7959 | 2.7896 | 2.1302 | 2.1278 | 1.0958 | 1.0942 | 1.9487 | O4 | 2.9324 | 2.3824 | 1.4232 | | 3.9114 | 3.3433 | 2.6227 | 2.6025 | 3.3168 | 2.0780 | 2.0766 | 0.9565 | H5 | 1.0902 | 2.1712 | 3.4797 | 3.9114 | | 1.7641 | 1.7667 | 2.5041 | 2.5064 | 3.7598 | 4.3031 | 4.8045 | H6 | 1.0920 | 2.1682 | 2.7959 | 3.3433 | 1.7641 | | 1.7642 | 3.0700 | 2.5127 | 2.5724 | 3.7745 | 4.0249 | H7 | 1.0887 | 2.1647 | 2.7896 | 2.6227 | 1.7667 | 1.7642 | | 2.5137 | 3.0675 | 3.1054 | 3.8021 | 3.4554 | H8 | 2.1605 | 1.0920 | 2.1302 | 2.6025 | 2.5041 | 3.0700 | 2.5137 | | 1.7576 | 3.0415 | 2.4731 | 3.4881 | H9 | 2.1616 | 1.0927 | 2.1278 | 3.3168 | 2.5064 | 2.5127 | 3.0675 | 1.7576 | | 2.4864 | 2.4199 | 4.0417 | H10 | 2.7580 | 2.1498 | 1.0958 | 2.0780 | 3.7598 | 2.5724 | 3.1054 | 3.0415 | 2.4864 | | 1.7723 | 2.3065 | H11 | 3.4748 | 2.1519 | 1.0942 | 2.0766 | 4.3031 | 3.7745 | 3.8021 | 2.4731 | 2.4199 | 1.7723 | | 2.3325 | H12 | 3.7813 | 3.2167 | 1.9487 | 0.9565 | 4.8045 | 4.0249 | 3.4554 | 3.4881 | 4.0417 | 2.3065 | 2.3325 | |
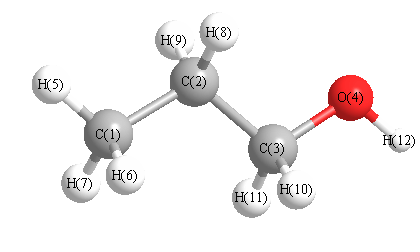 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
112.705 |
|
C1 |
C2 |
H8 |
110.134 |
| C1 |
C2 |
H9 |
110.175 |
|
C2 |
C1 |
H5 |
111.093 |
| C2 |
C1 |
H6 |
110.737 |
|
C2 |
C1 |
H7 |
110.664 |
| C2 |
C3 |
O4 |
108.209 |
|
C2 |
C3 |
H10 |
109.687 |
| C2 |
C3 |
H11 |
109.941 |
|
C3 |
C2 |
H8 |
108.372 |
| C3 |
C2 |
H9 |
108.145 |
|
C3 |
O4 |
H12 |
108.345 |
| O4 |
C3 |
H10 |
110.486 |
|
O4 |
C3 |
H11 |
110.475 |
| H5 |
C1 |
H6 |
107.880 |
|
H5 |
C1 |
H7 |
108.354 |
| H6 |
C1 |
H7 |
107.998 |
|
H8 |
C2 |
H9 |
107.124 |
| H10 |
C3 |
H11 |
108.042 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
