Jump to
S1C2
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -232.247621 |
| Energy at 298.15K | -232.259050 |
| HF Energy | -232.247621 |
| Nuclear repulsion energy | 186.065563 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
4184 |
3793 |
58.20 |
|
|
|
| 2 |
A' |
3214 |
2913 |
62.89 |
|
|
|
| 3 |
A' |
3163 |
2867 |
56.53 |
|
|
|
| 4 |
A' |
3154 |
2859 |
21.37 |
|
|
|
| 5 |
A' |
3144 |
2849 |
62.64 |
|
|
|
| 6 |
A' |
3126 |
2833 |
37.87 |
|
|
|
| 7 |
A' |
1652 |
1498 |
2.97 |
|
|
|
| 8 |
A' |
1628 |
1476 |
4.87 |
|
|
|
| 9 |
A' |
1617 |
1465 |
1.27 |
|
|
|
| 10 |
A' |
1609 |
1458 |
0.38 |
|
|
|
| 11 |
A' |
1591 |
1442 |
7.89 |
|
|
|
| 12 |
A' |
1539 |
1395 |
1.33 |
|
|
|
| 13 |
A' |
1519 |
1377 |
3.37 |
|
|
|
| 14 |
A' |
1427 |
1293 |
20.19 |
|
|
|
| 15 |
A' |
1340 |
1215 |
66.89 |
|
|
|
| 16 |
A' |
1200 |
1088 |
2.94 |
|
|
|
| 17 |
A' |
1171 |
1062 |
104.61 |
|
|
|
| 18 |
A' |
1122 |
1017 |
1.52 |
|
|
|
| 19 |
A' |
1060 |
961 |
0.53 |
|
|
|
| 20 |
A' |
966 |
876 |
10.87 |
|
|
|
| 21 |
A' |
472 |
428 |
13.36 |
|
|
|
| 22 |
A' |
420 |
380 |
0.08 |
|
|
|
| 23 |
A' |
201 |
182 |
2.49 |
|
|
|
| 24 |
A" |
3216 |
2915 |
142.23 |
|
|
|
| 25 |
A" |
3199 |
2900 |
17.42 |
|
|
|
| 26 |
A" |
3160 |
2864 |
6.70 |
|
|
|
| 27 |
A" |
3146 |
2852 |
38.42 |
|
|
|
| 28 |
A" |
1614 |
1463 |
6.24 |
|
|
|
| 29 |
A" |
1432 |
1298 |
0.00 |
|
|
|
| 30 |
A" |
1429 |
1295 |
0.30 |
|
|
|
| 31 |
A" |
1358 |
1231 |
1.02 |
|
|
|
| 32 |
A" |
1293 |
1172 |
2.62 |
|
|
|
| 33 |
A" |
1026 |
930 |
0.19 |
|
|
|
| 34 |
A" |
872 |
791 |
1.05 |
|
|
|
| 35 |
A" |
794 |
720 |
1.10 |
|
|
|
| 36 |
A" |
281 |
255 |
120.44 |
|
|
|
| 37 |
A" |
264 |
240 |
4.63 |
|
|
|
| 38 |
A" |
128 |
116 |
2.06 |
|
|
|
| 39 |
A" |
115 |
105 |
5.42 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31923.7 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 28935.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.350 |
-0.340 |
0.000 |
| C2 |
0.000 |
0.344 |
0.000 |
| C3 |
-1.160 |
-0.644 |
0.000 |
| C4 |
-2.519 |
0.044 |
0.000 |
| O5 |
2.347 |
0.652 |
0.000 |
| H6 |
1.455 |
-0.970 |
0.878 |
| H7 |
1.455 |
-0.970 |
-0.878 |
| H8 |
-0.061 |
0.988 |
0.870 |
| H9 |
-0.061 |
0.988 |
-0.870 |
| H10 |
-1.086 |
-1.291 |
0.869 |
| H11 |
-1.086 |
-1.291 |
-0.869 |
| H12 |
-3.324 |
-0.681 |
0.000 |
| H13 |
-2.638 |
0.672 |
0.875 |
| H14 |
-2.638 |
0.672 |
-0.875 |
| H15 |
3.191 |
0.245 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
O5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
| C1 | | 1.5129 | 2.5286 | 3.8879 | 1.4058 | 1.0859 | 1.0859 | 2.1238 | 2.1238 | 2.7561 | 2.7561 | 4.6866 | 4.2066 | 4.2066 | 1.9321 |
C2 | 1.5129 | | 1.5236 | 2.5369 | 2.3667 | 2.1477 | 2.1477 | 1.0848 | 1.0848 | 2.1465 | 2.1465 | 3.4785 | 2.7991 | 2.7991 | 3.1929 | C3 | 2.5286 | 1.5236 | | 1.5227 | 3.7386 | 2.7779 | 2.7779 | 2.1518 | 2.1518 | 1.0862 | 1.0862 | 2.1642 | 2.1639 | 2.1639 | 4.4416 | C4 | 3.8879 | 2.5369 | 1.5227 | | 4.9034 | 4.1941 | 4.1941 | 2.7736 | 2.7736 | 2.1423 | 2.1423 | 1.0832 | 1.0841 | 1.0841 | 5.7140 | O5 | 1.4058 | 2.3667 | 3.7386 | 4.9034 | | 2.0484 | 2.0484 | 2.5818 | 2.5818 | 4.0392 | 4.0392 | 5.8253 | 5.0612 | 5.0612 | 0.9376 | H6 | 1.0859 | 2.1477 | 2.7779 | 4.1941 | 2.0484 | | 1.7559 | 2.4763 | 3.0312 | 2.5614 | 3.1005 | 4.8677 | 4.4103 | 4.7460 | 2.2941 | H7 | 1.0859 | 2.1477 | 2.7779 | 4.1941 | 2.0484 | 1.7559 | | 3.0312 | 2.4763 | 3.1005 | 2.5614 | 4.8677 | 4.7460 | 4.4103 | 2.2941 | H8 | 2.1238 | 1.0848 | 2.1518 | 2.7736 | 2.5818 | 2.4763 | 3.0312 | | 1.7405 | 2.4994 | 3.0451 | 3.7674 | 2.5968 | 3.1290 | 3.4476 | H9 | 2.1238 | 1.0848 | 2.1518 | 2.7736 | 2.5818 | 3.0312 | 2.4763 | 1.7405 | | 3.0451 | 2.4994 | 3.7674 | 3.1290 | 2.5968 | 3.4476 | H10 | 2.7561 | 2.1465 | 1.0862 | 2.1423 | 4.0392 | 2.5614 | 3.1005 | 2.4994 | 3.0451 | | 1.7384 | 2.4771 | 2.5025 | 3.0505 | 4.6276 | H11 | 2.7561 | 2.1465 | 1.0862 | 2.1423 | 4.0392 | 3.1005 | 2.5614 | 3.0451 | 2.4994 | 1.7384 | | 2.4771 | 3.0505 | 2.5025 | 4.6276 | H12 | 4.6866 | 3.4785 | 2.1642 | 1.0832 | 5.8253 | 4.8677 | 4.8677 | 3.7674 | 3.7674 | 2.4771 | 2.4771 | | 1.7513 | 1.7513 | 6.5811 | H13 | 4.2066 | 2.7991 | 2.1639 | 1.0841 | 5.0612 | 4.4103 | 4.7460 | 2.5968 | 3.1290 | 2.5025 | 3.0505 | 1.7513 | | 1.7505 | 5.9105 | H14 | 4.2066 | 2.7991 | 2.1639 | 1.0841 | 5.0612 | 4.7460 | 4.4103 | 3.1290 | 2.5968 | 3.0505 | 2.5025 | 1.7513 | 1.7505 | | 5.9105 | H15 | 1.9321 | 3.1929 | 4.4416 | 5.7140 | 0.9376 | 2.2941 | 2.2941 | 3.4476 | 3.4476 | 4.6276 | 4.6276 | 6.5811 | 5.9105 | 5.9105 | |
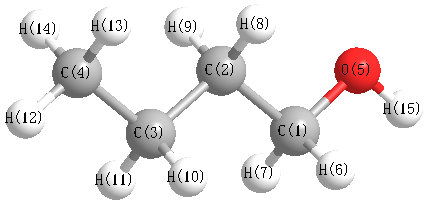 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
112.762 |
|
C1 |
C2 |
H8 |
108.567 |
| C1 |
C2 |
H9 |
108.567 |
|
C1 |
O5 |
H15 |
109.451 |
| C2 |
C1 |
O5 |
108.309 |
|
C2 |
C1 |
H6 |
110.389 |
| C2 |
C1 |
H7 |
110.389 |
|
C2 |
C3 |
C4 |
112.764 |
| C2 |
C3 |
H10 |
109.527 |
|
C2 |
C3 |
H11 |
109.527 |
| C3 |
C2 |
H8 |
110.027 |
|
C3 |
C2 |
H9 |
110.027 |
| C3 |
C4 |
H12 |
111.180 |
|
C3 |
C4 |
H13 |
111.095 |
| C3 |
C4 |
H14 |
111.095 |
|
C4 |
C3 |
H10 |
109.258 |
| C4 |
C3 |
H11 |
109.258 |
|
O5 |
C1 |
H6 |
109.930 |
| O5 |
C1 |
H7 |
109.930 |
|
H6 |
C1 |
H7 |
107.895 |
| H8 |
C2 |
H9 |
106.691 |
|
H10 |
C3 |
H11 |
106.299 |
| H12 |
C4 |
H13 |
107.815 |
|
H12 |
C4 |
H14 |
107.815 |
| H13 |
C4 |
H14 |
107.680 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.129 |
|
|
|
| 2 |
C |
-0.024 |
|
|
|
| 3 |
C |
-0.045 |
|
|
|
| 4 |
C |
-0.182 |
|
|
|
| 5 |
O |
-0.410 |
|
|
|
| 6 |
H |
0.029 |
|
|
|
| 7 |
H |
0.029 |
|
|
|
| 8 |
H |
0.034 |
|
|
|
| 9 |
H |
0.034 |
|
|
|
| 10 |
H |
0.027 |
|
|
|
| 11 |
H |
0.027 |
|
|
|
| 12 |
H |
0.060 |
|
|
|
| 13 |
H |
0.049 |
|
|
|
| 14 |
H |
0.049 |
|
|
|
| 15 |
H |
0.193 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.232 |
-1.594 |
0.000 |
1.611 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-29.957 |
-3.621 |
0.000 |
| y |
-3.621 |
-35.216 |
0.000 |
| z |
0.000 |
0.000 |
-33.150 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.226 |
-3.621 |
0.000 |
| y |
-3.621 |
-3.663 |
0.000 |
| z |
0.000 |
0.000 |
-0.563 |
|
| Polar |
|---|
| 3z2-r2 | -1.127 |
|---|
| x2-y2 | 5.259 |
|---|
| xy | -3.621 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.890 |
0.058 |
0.000 |
| y |
0.058 |
7.224 |
0.000 |
| z |
0.000 |
0.000 |
6.788 |
<r2> (average value of r
2) Å
2
| <r2> |
185.856 |
| (<r2>)1/2 |
13.633 |
Jump to
S1C1
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -232.247488 |
| Energy at 298.15K | -232.258995 |
| HF Energy | -232.247488 |
| Nuclear repulsion energy | 188.815009 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
4186 |
3794 |
60.43 |
|
|
|
| 2 |
A |
3221 |
2920 |
102.20 |
|
|
|
| 3 |
A |
3212 |
2912 |
70.45 |
|
|
|
| 4 |
A |
3193 |
2894 |
1.95 |
|
|
|
| 5 |
A |
3183 |
2885 |
52.45 |
|
|
|
| 6 |
A |
3161 |
2865 |
90.42 |
|
|
|
| 7 |
A |
3152 |
2857 |
40.40 |
|
|
|
| 8 |
A |
3144 |
2849 |
36.84 |
|
|
|
| 9 |
A |
3143 |
2849 |
13.41 |
|
|
|
| 10 |
A |
3126 |
2833 |
53.66 |
|
|
|
| 11 |
A |
1648 |
1494 |
3.50 |
|
|
|
| 12 |
A |
1628 |
1476 |
4.58 |
|
|
|
| 13 |
A |
1615 |
1463 |
6.27 |
|
|
|
| 14 |
A |
1611 |
1460 |
1.08 |
|
|
|
| 15 |
A |
1600 |
1450 |
3.95 |
|
|
|
| 16 |
A |
1584 |
1435 |
8.11 |
|
|
|
| 17 |
A |
1540 |
1396 |
0.49 |
|
|
|
| 18 |
A |
1533 |
1389 |
1.61 |
|
|
|
| 19 |
A |
1451 |
1315 |
2.34 |
|
|
|
| 20 |
A |
1441 |
1306 |
8.81 |
|
|
|
| 21 |
A |
1408 |
1276 |
30.42 |
|
|
|
| 22 |
A |
1368 |
1240 |
10.16 |
|
|
|
| 23 |
A |
1338 |
1213 |
39.47 |
|
|
|
| 24 |
A |
1257 |
1139 |
9.74 |
|
|
|
| 25 |
A |
1214 |
1101 |
9.49 |
|
|
|
| 26 |
A |
1170 |
1061 |
72.68 |
|
|
|
| 27 |
A |
1107 |
1003 |
4.67 |
|
|
|
| 28 |
A |
1055 |
956 |
4.16 |
|
|
|
| 29 |
A |
1028 |
931 |
1.23 |
|
|
|
| 30 |
A |
920 |
834 |
7.76 |
|
|
|
| 31 |
A |
889 |
806 |
1.16 |
|
|
|
| 32 |
A |
798 |
723 |
0.73 |
|
|
|
| 33 |
A |
543 |
492 |
10.56 |
|
|
|
| 34 |
A |
366 |
332 |
2.65 |
|
|
|
| 35 |
A |
297 |
269 |
49.63 |
|
|
|
| 36 |
A |
262 |
238 |
8.15 |
|
|
|
| 37 |
A |
248 |
225 |
69.68 |
|
|
|
| 38 |
A |
152 |
138 |
6.65 |
|
|
|
| 39 |
A |
93 |
84 |
0.59 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31940.7 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 28951.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.435 |
0.508 |
0.234 |
| C2 |
0.000 |
0.688 |
-0.223 |
| C3 |
-0.965 |
-0.348 |
0.345 |
| C4 |
-2.398 |
-0.140 |
-0.131 |
| O5 |
1.930 |
-0.711 |
-0.258 |
| H6 |
2.041 |
1.332 |
-0.134 |
| H7 |
1.483 |
0.524 |
1.321 |
| H8 |
-0.323 |
1.684 |
0.069 |
| H9 |
-0.021 |
0.656 |
-1.308 |
| H10 |
-0.936 |
-0.305 |
1.430 |
| H11 |
-0.629 |
-1.339 |
0.066 |
| H12 |
-3.063 |
-0.885 |
0.289 |
| H13 |
-2.771 |
0.837 |
0.159 |
| H14 |
-2.464 |
-0.209 |
-1.212 |
| H15 |
2.809 |
-0.850 |
0.037 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
O5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
| C1 | | 1.5168 | 2.5508 | 3.9050 | 1.4047 | 1.0868 | 1.0879 | 2.1218 | 2.1267 | 2.7779 | 2.7746 | 4.7095 | 4.2204 | 4.2206 | 1.9417 |
C2 | 1.5168 | | 1.5260 | 2.5392 | 2.3838 | 2.1411 | 2.1469 | 1.0868 | 1.0857 | 2.1442 | 2.1420 | 3.4818 | 2.8021 | 2.8033 | 3.2131 | C3 | 2.5508 | 1.5260 | | 1.5246 | 2.9790 | 3.4765 | 2.7762 | 2.1491 | 2.1515 | 1.0869 | 1.0826 | 2.1664 | 2.1686 | 2.1656 | 3.8200 | C4 | 3.9050 | 2.5392 | 1.5246 | | 4.3674 | 4.6764 | 4.1969 | 2.7700 | 2.7687 | 2.1454 | 2.1467 | 1.0838 | 1.0851 | 1.0847 | 5.2586 | O5 | 1.4047 | 2.3838 | 2.9790 | 4.3674 | | 2.0500 | 2.0540 | 3.3043 | 2.6035 | 3.3512 | 2.6540 | 5.0255 | 4.9672 | 4.5243 | 0.9382 | H6 | 1.0868 | 2.1411 | 3.4765 | 4.6764 | 2.0500 | | 1.7552 | 2.3981 | 2.4674 | 3.7402 | 3.7811 | 5.5805 | 4.8464 | 4.8818 | 2.3196 | H7 | 1.0879 | 2.1469 | 2.7762 | 4.1969 | 2.0540 | 1.7552 | | 2.4849 | 3.0319 | 2.5599 | 3.0830 | 4.8702 | 4.4215 | 4.7471 | 2.3011 | H8 | 2.1218 | 1.0868 | 2.1491 | 2.7700 | 3.3043 | 2.3981 | 2.4849 | | 1.7448 | 2.4872 | 3.0382 | 3.7629 | 2.5926 | 3.1323 | 4.0290 | H9 | 2.1267 | 1.0857 | 2.1515 | 2.7687 | 2.6035 | 2.4674 | 3.0319 | 1.7448 | | 3.0426 | 2.4965 | 3.7649 | 3.1222 | 2.5933 | 3.4770 | H10 | 2.7779 | 2.1442 | 1.0869 | 2.1454 | 3.3512 | 3.7402 | 2.5599 | 2.4872 | 3.0426 | | 1.7394 | 2.4824 | 2.5074 | 3.0535 | 4.0336 | H11 | 2.7746 | 2.1420 | 1.0826 | 2.1467 | 2.6540 | 3.7811 | 3.0830 | 3.0382 | 2.4965 | 1.7394 | | 2.4862 | 3.0552 | 2.5053 | 3.4728 | H12 | 4.7095 | 3.4818 | 2.1664 | 1.0838 | 5.0255 | 5.5805 | 4.8702 | 3.7629 | 3.7649 | 2.4824 | 2.4862 | | 1.7518 | 1.7512 | 5.8779 | H13 | 4.2204 | 2.8021 | 2.1686 | 1.0851 | 4.9672 | 4.8464 | 4.4215 | 2.5926 | 3.1222 | 2.5074 | 3.0552 | 1.7518 | | 1.7519 | 5.8316 | H14 | 4.2206 | 2.8033 | 2.1656 | 1.0847 | 4.5243 | 4.8818 | 4.7471 | 3.1323 | 2.5933 | 3.0535 | 2.5053 | 1.7512 | 1.7519 | | 5.4573 | H15 | 1.9417 | 3.2131 | 3.8200 | 5.2586 | 0.9382 | 2.3196 | 2.3011 | 4.0290 | 3.4770 | 4.0336 | 3.4728 | 5.8779 | 5.8316 | 5.4573 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
113.923 |
|
C1 |
C2 |
H8 |
108.024 |
| C1 |
C2 |
H9 |
108.469 |
|
C1 |
O5 |
H15 |
110.361 |
| C2 |
C1 |
O5 |
109.303 |
|
C2 |
C1 |
H6 |
109.540 |
| C2 |
C1 |
H7 |
109.930 |
|
C2 |
C3 |
C4 |
112.682 |
| C2 |
C3 |
H10 |
109.139 |
|
C2 |
C3 |
H11 |
109.212 |
| C3 |
C2 |
H8 |
109.529 |
|
C3 |
C2 |
H9 |
109.784 |
| C3 |
C4 |
H12 |
111.186 |
|
C3 |
C4 |
H13 |
111.284 |
| C3 |
C4 |
H14 |
111.064 |
|
C4 |
C3 |
H10 |
109.334 |
| C4 |
C3 |
H11 |
109.686 |
|
O5 |
C1 |
H6 |
110.085 |
| O5 |
C1 |
H7 |
110.335 |
|
H6 |
C1 |
H7 |
107.628 |
| H8 |
C2 |
H9 |
106.860 |
|
H10 |
C3 |
H11 |
106.602 |
| H12 |
C4 |
H13 |
107.740 |
|
H12 |
C4 |
H14 |
107.712 |
| H13 |
C4 |
H14 |
107.683 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.117 |
|
|
|
| 2 |
C |
-0.041 |
|
|
|
| 3 |
C |
-0.027 |
|
|
|
| 4 |
C |
-0.184 |
|
|
|
| 5 |
O |
-0.415 |
|
|
|
| 6 |
H |
0.043 |
|
|
|
| 7 |
H |
0.030 |
|
|
|
| 8 |
H |
0.026 |
|
|
|
| 9 |
H |
0.037 |
|
|
|
| 10 |
H |
0.017 |
|
|
|
| 11 |
H |
0.052 |
|
|
|
| 12 |
H |
0.059 |
|
|
|
| 13 |
H |
0.043 |
|
|
|
| 14 |
H |
0.049 |
|
|
|
| 15 |
H |
0.192 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.907 |
0.902 |
0.912 |
1.571 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-28.841 |
0.478 |
1.851 |
| y |
0.478 |
-33.903 |
-0.533 |
| z |
1.851 |
-0.533 |
-33.719 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.970 |
0.478 |
1.851 |
| y |
0.478 |
-2.623 |
-0.533 |
| z |
1.851 |
-0.533 |
-2.347 |
|
| Polar |
|---|
| 3z2-r2 | -4.694 |
|---|
| x2-y2 | 5.062 |
|---|
| xy | 0.478 |
|---|
| xz | 1.851 |
|---|
| yz | -0.533 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.688 |
0.077 |
0.124 |
| y |
0.077 |
7.243 |
-0.019 |
| z |
0.124 |
-0.019 |
6.963 |
<r2> (average value of r
2) Å
2
| <r2> |
168.819 |
| (<r2>)1/2 |
12.993 |
