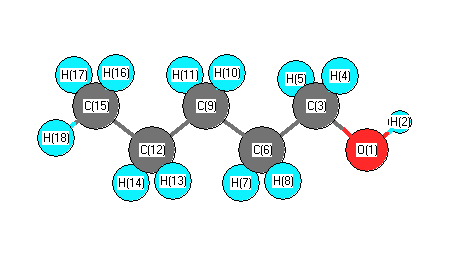
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
4184 |
3793 |
59.13 |
|
|
|
| 2 |
A' |
3213 |
2912 |
64.13 |
|
|
|
| 3 |
A' |
3164 |
2868 |
63.58 |
|
|
|
| 4 |
A' |
3153 |
2858 |
39.59 |
|
|
|
| 5 |
A' |
3146 |
2852 |
44.39 |
|
|
|
| 6 |
A' |
3137 |
2843 |
46.47 |
|
|
|
| 7 |
A' |
3125 |
2832 |
23.94 |
|
|
|
| 8 |
A' |
1652 |
1498 |
3.24 |
|
|
|
| 9 |
A' |
1630 |
1478 |
6.25 |
|
|
|
| 10 |
A' |
1620 |
1468 |
0.01 |
|
|
|
| 11 |
A' |
1611 |
1460 |
1.02 |
|
|
|
| 12 |
A' |
1607 |
1457 |
0.68 |
|
|
|
| 13 |
A' |
1592 |
1443 |
7.68 |
|
|
|
| 14 |
A' |
1541 |
1397 |
1.01 |
|
|
|
| 15 |
A' |
1535 |
1391 |
2.19 |
|
|
|
| 16 |
A' |
1480 |
1342 |
8.90 |
|
|
|
| 17 |
A' |
1403 |
1272 |
25.16 |
|
|
|
| 18 |
A' |
1330 |
1206 |
53.41 |
|
|
|
| 19 |
A' |
1209 |
1096 |
1.89 |
|
|
|
| 20 |
A' |
1174 |
1064 |
110.48 |
|
|
|
| 21 |
A' |
1130 |
1024 |
0.33 |
|
|
|
| 22 |
A' |
1093 |
991 |
0.41 |
|
|
|
| 23 |
A' |
1069 |
969 |
10.27 |
|
|
|
| 24 |
A' |
956 |
867 |
2.10 |
|
|
|
| 25 |
A' |
532 |
482 |
12.39 |
|
|
|
| 26 |
A' |
387 |
351 |
0.11 |
|
|
|
| 27 |
A' |
337 |
306 |
4.50 |
|
|
|
| 28 |
A' |
146 |
132 |
1.58 |
|
|
|
| 29 |
A" |
3213 |
2912 |
186.33 |
|
|
|
| 30 |
A" |
3203 |
2904 |
14.13 |
|
|
|
| 31 |
A" |
3173 |
2876 |
5.54 |
|
|
|
| 32 |
A" |
3150 |
2855 |
15.35 |
|
|
|
| 33 |
A" |
3146 |
2851 |
28.15 |
|
|
|
| 34 |
A" |
1614 |
1463 |
6.20 |
|
|
|
| 35 |
A" |
1443 |
1308 |
0.48 |
|
|
|
| 36 |
A" |
1432 |
1298 |
0.35 |
|
|
|
| 37 |
A" |
1409 |
1277 |
0.26 |
|
|
|
| 38 |
A" |
1344 |
1218 |
0.77 |
|
|
|
| 39 |
A" |
1294 |
1173 |
2.08 |
|
|
|
| 40 |
A" |
1062 |
962 |
0.84 |
|
|
|
| 41 |
A" |
927 |
840 |
0.10 |
|
|
|
| 42 |
A" |
827 |
749 |
0.37 |
|
|
|
| 43 |
A" |
785 |
712 |
1.88 |
|
|
|
| 44 |
A" |
281 |
254 |
121.33 |
|
|
|
| 45 |
A" |
262 |
238 |
2.80 |
|
|
|
| 46 |
A" |
165 |
149 |
1.99 |
|
|
|
| 47 |
A" |
106 |
96 |
4.25 |
|
|
|
| 48 |
A" |
84 |
76 |
2.30 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 38536.9 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 34929.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.409 |
|
|
|
| 2 |
H |
0.193 |
|
|
|
| 3 |
C |
0.123 |
|
|
|
| 4 |
H |
0.029 |
|
|
|
| 5 |
H |
0.029 |
|
|
|
| 6 |
C |
-0.013 |
|
|
|
| 7 |
H |
0.035 |
|
|
|
| 8 |
H |
0.035 |
|
|
|
| 9 |
C |
-0.086 |
|
|
|
| 10 |
H |
0.030 |
|
|
|
| 11 |
H |
0.030 |
|
|
|
| 12 |
C |
-0.022 |
|
|
|
| 13 |
H |
0.029 |
|
|
|
| 14 |
H |
0.029 |
|
|
|
| 15 |
C |
-0.185 |
|
|
|
| 16 |
H |
0.047 |
|
|
|
| 17 |
H |
0.047 |
|
|
|
| 18 |
H |
0.061 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.278 |
0.822 |
0.000 |
1.520 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.076 |
-3.859 |
0.000 |
| y |
-3.859 |
-44.185 |
0.000 |
| z |
0.000 |
0.000 |
-39.705 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.870 |
-3.859 |
0.000 |
| y |
-3.859 |
-6.795 |
0.000 |
| z |
0.000 |
0.000 |
-0.074 |
|
| Polar |
|---|
| 3z2-r2 | -0.149 |
|---|
| x2-y2 | 9.110 |
|---|
| xy | -3.859 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.333 |
-1.063 |
0.000 |
| y |
-1.063 |
10.453 |
0.000 |
| z |
0.000 |
0.000 |
8.165 |
<r2> (average value of r
2) Å
2
| <r2> |
300.175 |
| (<r2>)1/2 |
17.326 |