Jump to
S1C2
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -232.255107 |
| Energy at 298.15K | -232.266655 |
| HF Energy | -232.255107 |
| Nuclear repulsion energy | 200.687187 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
4169 |
3779 |
35.77 |
|
|
|
| 2 |
A' |
3239 |
2936 |
83.92 |
|
|
|
| 3 |
A' |
3228 |
2926 |
2.82 |
|
|
|
| 4 |
A' |
3215 |
2914 |
78.49 |
|
|
|
| 5 |
A' |
3172 |
2875 |
6.68 |
|
|
|
| 6 |
A' |
3157 |
2862 |
50.53 |
|
|
|
| 7 |
A' |
1634 |
1481 |
6.70 |
|
|
|
| 8 |
A' |
1623 |
1471 |
3.09 |
|
|
|
| 9 |
A' |
1603 |
1453 |
0.02 |
|
|
|
| 10 |
A' |
1553 |
1408 |
4.68 |
|
|
|
| 11 |
A' |
1535 |
1391 |
25.13 |
|
|
|
| 12 |
A' |
1469 |
1332 |
42.18 |
|
|
|
| 13 |
A' |
1360 |
1233 |
67.25 |
|
|
|
| 14 |
A' |
1254 |
1136 |
55.11 |
|
|
|
| 15 |
A' |
1115 |
1011 |
16.49 |
|
|
|
| 16 |
A' |
1028 |
932 |
42.73 |
|
|
|
| 17 |
A' |
994 |
901 |
0.10 |
|
|
|
| 18 |
A' |
794 |
720 |
2.36 |
|
|
|
| 19 |
A' |
495 |
448 |
9.40 |
|
|
|
| 20 |
A' |
446 |
404 |
0.42 |
|
|
|
| 21 |
A' |
362 |
329 |
1.30 |
|
|
|
| 22 |
A' |
292 |
265 |
0.07 |
|
|
|
| 23 |
A" |
3240 |
2936 |
60.62 |
|
|
|
| 24 |
A" |
3230 |
2927 |
5.00 |
|
|
|
| 25 |
A" |
3206 |
2906 |
5.28 |
|
|
|
| 26 |
A" |
3151 |
2856 |
20.22 |
|
|
|
| 27 |
A" |
1615 |
1464 |
2.35 |
|
|
|
| 28 |
A" |
1601 |
1451 |
0.06 |
|
|
|
| 29 |
A" |
1591 |
1442 |
0.04 |
|
|
|
| 30 |
A" |
1527 |
1385 |
14.67 |
|
|
|
| 31 |
A" |
1364 |
1236 |
22.03 |
|
|
|
| 32 |
A" |
1135 |
1029 |
2.34 |
|
|
|
| 33 |
A" |
1037 |
940 |
0.00 |
|
|
|
| 34 |
A" |
982 |
890 |
0.11 |
|
|
|
| 35 |
A" |
502 |
455 |
11.14 |
|
|
|
| 36 |
A" |
365 |
331 |
8.34 |
|
|
|
| 37 |
A" |
309 |
280 |
85.59 |
|
|
|
| 38 |
A" |
279 |
253 |
26.37 |
|
|
|
| 39 |
A" |
218 |
198 |
3.13 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31543.3 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 28590.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.000 |
0.021 |
0.000 |
| O2 |
-0.485 |
1.350 |
0.000 |
| C3 |
1.517 |
0.141 |
0.000 |
| C4 |
-0.485 |
-0.699 |
1.256 |
| C5 |
-0.485 |
-0.699 |
-1.256 |
| H6 |
-1.424 |
1.353 |
0.000 |
| H7 |
1.983 |
-0.837 |
0.000 |
| H8 |
1.850 |
0.682 |
-0.877 |
| H9 |
1.850 |
0.682 |
0.877 |
| H10 |
-0.101 |
-1.711 |
1.300 |
| H11 |
-0.101 |
-1.711 |
-1.300 |
| H12 |
-0.160 |
-0.168 |
2.142 |
| H13 |
-1.569 |
-0.754 |
1.274 |
| H14 |
-0.160 |
-0.168 |
-2.142 |
| H15 |
-1.569 |
-0.754 |
-1.274 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
| C1 | | 1.4145 | 1.5224 | 1.5266 | 1.5266 | 1.9502 | 2.1606 | 2.1515 | 2.1515 | 2.1681 | 2.1681 | 2.1559 | 2.1644 | 2.1559 | 2.1644 |
O2 | 1.4145 | | 2.3387 | 2.4029 | 2.4029 | 0.9394 | 3.2972 | 2.5819 | 2.5819 | 3.3476 | 3.3476 | 2.6449 | 2.6879 | 2.6449 | 2.6879 | C3 | 1.5224 | 2.3387 | | 2.5085 | 2.5085 | 3.1816 | 1.0834 | 1.0825 | 1.0825 | 2.7824 | 2.7824 | 2.7377 | 3.4570 | 2.7377 | 3.4570 | C4 | 1.5266 | 2.4029 | 2.5085 | | 2.5121 | 2.5829 | 2.7723 | 3.4507 | 2.7390 | 1.0836 | 2.7759 | 1.0826 | 1.0857 | 3.4544 | 2.7529 | C5 | 1.5266 | 2.4029 | 2.5085 | 2.5121 | | 2.5829 | 2.7723 | 2.7390 | 3.4507 | 2.7759 | 1.0836 | 3.4544 | 2.7529 | 1.0826 | 1.0857 | H6 | 1.9502 | 0.9394 | 3.1816 | 2.5829 | 2.5829 | | 4.0504 | 3.4555 | 3.4555 | 3.5821 | 3.5821 | 2.9155 | 2.4669 | 2.9155 | 2.4669 | H7 | 2.1606 | 3.2972 | 1.0834 | 2.7723 | 2.7723 | 4.0504 | | 1.7590 | 1.7590 | 2.6071 | 2.6071 | 3.1024 | 3.7742 | 3.1024 | 3.7742 | H8 | 2.1515 | 2.5819 | 1.0825 | 3.4507 | 2.7390 | 3.4555 | 1.7590 | | 1.7537 | 3.7780 | 3.1167 | 3.7246 | 4.2869 | 2.5221 | 3.7297 | H9 | 2.1515 | 2.5819 | 1.0825 | 2.7390 | 3.4507 | 3.4555 | 1.7590 | 1.7537 | | 3.1167 | 3.7780 | 2.5221 | 3.7297 | 3.7246 | 4.2869 | H10 | 2.1681 | 3.3476 | 2.7824 | 1.0836 | 2.7759 | 3.5821 | 2.6071 | 3.7780 | 3.1167 | | 2.5999 | 1.7590 | 1.7524 | 3.7724 | 3.1135 | H11 | 2.1681 | 3.3476 | 2.7824 | 2.7759 | 1.0836 | 3.5821 | 2.6071 | 3.1167 | 3.7780 | 2.5999 | | 3.7724 | 3.1135 | 1.7590 | 1.7524 | H12 | 2.1559 | 2.6449 | 2.7377 | 1.0826 | 3.4544 | 2.9155 | 3.1024 | 3.7246 | 2.5221 | 1.7590 | 3.7724 | | 1.7560 | 4.2835 | 3.7411 | H13 | 2.1644 | 2.6879 | 3.4570 | 1.0857 | 2.7529 | 2.4669 | 3.7742 | 4.2869 | 3.7297 | 1.7524 | 3.1135 | 1.7560 | | 3.7411 | 2.5475 | H14 | 2.1559 | 2.6449 | 2.7377 | 3.4544 | 1.0826 | 2.9155 | 3.1024 | 2.5221 | 3.7246 | 3.7724 | 1.7590 | 4.2835 | 3.7411 | | 1.7560 | H15 | 2.1644 | 2.6879 | 3.4570 | 2.7529 | 1.0857 | 2.4669 | 3.7742 | 3.7297 | 4.2869 | 3.1135 | 1.7524 | 3.7411 | 2.5475 | 1.7560 | |
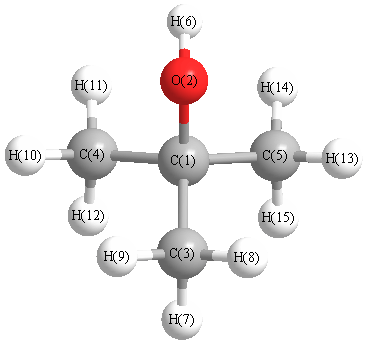 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C5 |
H2 |
33.698 |
|
C1 |
C5 |
H3 |
34.586 |
| C1 |
C5 |
H4 |
34.637 |
|
C1 |
C9 |
H6 |
30.843 |
| C1 |
C9 |
H7 |
66.196 |
|
C1 |
C9 |
H8 |
65.950 |
| C1 |
C13 |
H10 |
66.250 |
|
C1 |
C13 |
H11 |
44.103 |
| C1 |
C13 |
H12 |
65.764 |
|
C1 |
O15 |
H14 |
65.764 |
| H2 |
C5 |
H3 |
56.823 |
|
H2 |
C5 |
H4 |
58.485 |
| H3 |
C5 |
H4 |
59.952 |
|
C5 |
C1 |
C9 |
138.867 |
| C5 |
C1 |
C13 |
94.888 |
|
C5 |
C1 |
O15 |
27.972 |
| H6 |
C9 |
H7 |
96.476 |
|
H6 |
C9 |
H8 |
75.301 |
| H7 |
C9 |
H8 |
60.101 |
|
C9 |
C1 |
C13 |
119.574 |
| C9 |
C1 |
O15 |
166.707 |
|
H10 |
C13 |
H11 |
56.611 |
| H10 |
C13 |
H12 |
60.181 |
|
H11 |
C13 |
H12 |
97.636 |
| C13 |
C1 |
O15 |
72.099 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.421 |
|
|
|
| 2 |
O |
-0.446 |
|
|
|
| 3 |
C |
-0.197 |
|
|
|
| 4 |
C |
-0.238 |
|
|
|
| 5 |
C |
-0.238 |
|
|
|
| 6 |
H |
0.189 |
|
|
|
| 7 |
H |
0.042 |
|
|
|
| 8 |
H |
0.066 |
|
|
|
| 9 |
H |
0.066 |
|
|
|
| 10 |
H |
0.048 |
|
|
|
| 11 |
H |
0.048 |
|
|
|
| 12 |
H |
0.073 |
|
|
|
| 13 |
H |
0.047 |
|
|
|
| 14 |
H |
0.073 |
|
|
|
| 15 |
H |
0.047 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.833 |
-1.370 |
0.000 |
1.603 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-30.473 |
-1.428 |
0.000 |
| y |
-1.428 |
-36.094 |
0.000 |
| z |
0.000 |
0.000 |
-33.686 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.417 |
-1.428 |
0.000 |
| y |
-1.428 |
-4.015 |
0.000 |
| z |
0.000 |
0.000 |
-0.402 |
|
| Polar |
|---|
| 3z2-r2 | -0.805 |
|---|
| x2-y2 | 5.622 |
|---|
| xy | -1.428 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.821 |
0.073 |
0.000 |
| y |
0.073 |
7.320 |
0.000 |
| z |
0.000 |
0.000 |
7.628 |
<r2> (average value of r
2) Å
2
| <r2> |
124.570 |
| (<r2>)1/2 |
11.161 |
Jump to
S1C1
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -232.255107 |
| Energy at 298.15K | -232.266670 |
| HF Energy | -232.255107 |
| Nuclear repulsion energy | 200.697921 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
4165 |
3775 |
35.86 |
|
|
|
| 2 |
A |
3239 |
2936 |
60.38 |
|
|
|
| 3 |
A |
3239 |
2936 |
83.12 |
|
|
|
| 4 |
A |
3229 |
2927 |
5.66 |
|
|
|
| 5 |
A |
3228 |
2926 |
3.50 |
|
|
|
| 6 |
A |
3215 |
2914 |
78.84 |
|
|
|
| 7 |
A |
3205 |
2905 |
4.93 |
|
|
|
| 8 |
A |
3171 |
2874 |
6.72 |
|
|
|
| 9 |
A |
3157 |
2861 |
50.25 |
|
|
|
| 10 |
A |
3150 |
2855 |
20.24 |
|
|
|
| 11 |
A |
1634 |
1481 |
6.69 |
|
|
|
| 12 |
A |
1623 |
1471 |
3.09 |
|
|
|
| 13 |
A |
1615 |
1464 |
2.34 |
|
|
|
| 14 |
A |
1603 |
1453 |
0.02 |
|
|
|
| 15 |
A |
1601 |
1451 |
0.06 |
|
|
|
| 16 |
A |
1591 |
1442 |
0.04 |
|
|
|
| 17 |
A |
1553 |
1408 |
4.66 |
|
|
|
| 18 |
A |
1535 |
1391 |
25.25 |
|
|
|
| 19 |
A |
1528 |
1385 |
14.74 |
|
|
|
| 20 |
A |
1469 |
1332 |
42.33 |
|
|
|
| 21 |
A |
1365 |
1237 |
21.95 |
|
|
|
| 22 |
A |
1361 |
1233 |
67.15 |
|
|
|
| 23 |
A |
1254 |
1137 |
55.14 |
|
|
|
| 24 |
A |
1135 |
1029 |
2.33 |
|
|
|
| 25 |
A |
1116 |
1011 |
16.34 |
|
|
|
| 26 |
A |
1037 |
940 |
0.00 |
|
|
|
| 27 |
A |
1029 |
932 |
42.69 |
|
|
|
| 28 |
A |
994 |
901 |
0.09 |
|
|
|
| 29 |
A |
983 |
891 |
0.11 |
|
|
|
| 30 |
A |
794 |
720 |
2.36 |
|
|
|
| 31 |
A |
502 |
455 |
11.29 |
|
|
|
| 32 |
A |
495 |
449 |
9.38 |
|
|
|
| 33 |
A |
446 |
405 |
0.42 |
|
|
|
| 34 |
A |
365 |
331 |
9.39 |
|
|
|
| 35 |
A |
363 |
329 |
1.30 |
|
|
|
| 36 |
A |
313 |
284 |
90.56 |
|
|
|
| 37 |
A |
293 |
266 |
0.07 |
|
|
|
| 38 |
A |
281 |
255 |
20.57 |
|
|
|
| 39 |
A |
220 |
199 |
2.76 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31547.3 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 28594.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.007 |
-0.000 |
0.013 |
| H2 |
1.722 |
1.274 |
-0.271 |
| H3 |
0.207 |
2.143 |
-0.107 |
| H4 |
0.589 |
1.308 |
-1.606 |
| C5 |
0.667 |
1.258 |
-0.527 |
| H6 |
-1.972 |
0.875 |
0.122 |
| H7 |
-1.971 |
-0.876 |
0.122 |
| H8 |
-1.669 |
-0.000 |
-1.371 |
| C9 |
-1.498 |
-0.000 |
-0.302 |
| H10 |
0.207 |
-2.143 |
-0.107 |
| H11 |
1.722 |
-1.274 |
-0.272 |
| H12 |
0.589 |
-1.308 |
-1.606 |
| C13 |
0.667 |
-1.258 |
-0.527 |
| H14 |
0.978 |
0.000 |
1.700 |
| O15 |
0.079 |
-0.000 |
1.431 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
H4 |
C5 |
H6 |
H7 |
H8 |
C9 |
H10 |
H11 |
H12 |
C13 |
H14 |
O15 |
| C1 | | 2.1663 | 2.1574 | 2.1651 | 1.5261 | 2.1534 | 2.1534 | 2.1632 | 1.5237 | 2.1574 | 2.1663 | 2.1651 | 1.5261 | 1.9538 | 1.4209 |
H2 | 2.1663 | | 1.7542 | 1.7503 | 1.0854 | 3.7353 | 4.2912 | 3.7857 | 3.4626 | 3.7416 | 2.5480 | 3.1195 | 2.7551 | 2.4622 | 2.6874 | H3 | 2.1574 | 1.7542 | | 1.7575 | 1.0820 | 2.5308 | 3.7299 | 3.1165 | 2.7456 | 4.2868 | 3.7416 | 3.7823 | 3.4582 | 2.9078 | 2.6415 | H4 | 2.1651 | 1.7503 | 1.7575 | | 1.0829 | 3.1191 | 3.7830 | 2.6206 | 2.7871 | 3.7823 | 3.1194 | 2.6167 | 2.7853 | 3.5765 | 3.3460 | C5 | 1.5261 | 1.0854 | 1.0820 | 1.0829 | | 2.7436 | 3.4547 | 2.7846 | 2.5139 | 3.4582 | 2.7550 | 2.7853 | 2.5166 | 2.5766 | 2.4007 | H6 | 2.1534 | 3.7353 | 2.5308 | 3.1191 | 2.7436 | | 1.7512 | 1.7571 | 1.0819 | 3.7299 | 4.2912 | 3.7830 | 3.4547 | 3.4582 | 2.5854 | H7 | 2.1534 | 4.2912 | 3.7299 | 3.7830 | 3.4547 | 1.7512 | | 1.7571 | 1.0819 | 2.5308 | 3.7353 | 3.1190 | 2.7436 | 3.4582 | 2.5855 | H8 | 2.1632 | 3.7857 | 3.1165 | 2.6206 | 2.7846 | 1.7571 | 1.7571 | | 1.0831 | 3.1166 | 3.7857 | 2.6206 | 2.7847 | 4.0552 | 3.3031 | C9 | 1.5237 | 3.4626 | 2.7456 | 2.7871 | 2.5139 | 1.0819 | 1.0819 | 1.0831 | | 2.7457 | 3.4626 | 2.7870 | 2.5139 | 3.1843 | 2.3430 | H10 | 2.1574 | 3.7416 | 4.2868 | 3.7823 | 3.4582 | 3.7299 | 2.5308 | 3.1166 | 2.7457 | | 1.7542 | 1.7575 | 1.0820 | 2.9078 | 2.6415 | H11 | 2.1663 | 2.5480 | 3.7416 | 3.1194 | 2.7550 | 4.2912 | 3.7353 | 3.7857 | 3.4626 | 1.7542 | | 1.7503 | 1.0854 | 2.4622 | 2.6874 | H12 | 2.1651 | 3.1195 | 3.7823 | 2.6167 | 2.7853 | 3.7830 | 3.1190 | 2.6206 | 2.7870 | 1.7575 | 1.7503 | | 1.0829 | 3.5765 | 3.3460 | C13 | 1.5261 | 2.7551 | 3.4582 | 2.7853 | 2.5166 | 3.4547 | 2.7436 | 2.7847 | 2.5139 | 1.0820 | 1.0854 | 1.0829 | | 2.5766 | 2.4007 | H14 | 1.9538 | 2.4622 | 2.9078 | 3.5765 | 2.5766 | 3.4582 | 3.4582 | 4.0552 | 3.1843 | 2.9078 | 2.4622 | 3.5765 | 2.5766 | | 0.9390 | O15 | 1.4209 | 2.6874 | 2.6415 | 3.3460 | 2.4007 | 2.5854 | 2.5855 | 3.3031 | 2.3430 | 2.6415 | 2.6874 | 3.3460 | 2.4007 | 0.9390 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C5 |
H2 |
110.974 |
|
C1 |
C5 |
H3 |
110.464 |
| C1 |
C5 |
H4 |
111.029 |
|
C1 |
C9 |
H6 |
110.322 |
| C1 |
C9 |
H7 |
110.322 |
|
C1 |
C9 |
H8 |
111.035 |
| C1 |
C13 |
H10 |
110.464 |
|
C1 |
C13 |
H11 |
110.974 |
| C1 |
C13 |
H12 |
111.029 |
|
C1 |
O15 |
H14 |
110.094 |
| H2 |
C5 |
H3 |
108.063 |
|
H2 |
C5 |
H4 |
107.653 |
| H3 |
C5 |
H4 |
108.545 |
|
C5 |
C1 |
C9 |
111.038 |
| C5 |
C1 |
C13 |
111.085 |
|
C5 |
C1 |
O15 |
109.049 |
| H6 |
C9 |
H7 |
108.064 |
|
H6 |
C9 |
H8 |
108.507 |
| H7 |
C9 |
H8 |
108.507 |
|
C9 |
C1 |
C13 |
111.038 |
| C9 |
C1 |
O15 |
105.391 |
|
H10 |
C13 |
H11 |
108.063 |
| H10 |
C13 |
H12 |
108.545 |
|
H11 |
C13 |
H12 |
107.653 |
| C13 |
C1 |
O15 |
109.049 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.421 |
|
|
|
| 2 |
H |
0.047 |
|
|
|
| 3 |
H |
0.073 |
|
|
|
| 4 |
H |
0.048 |
|
|
|
| 5 |
C |
-0.238 |
|
|
|
| 6 |
H |
0.066 |
|
|
|
| 7 |
H |
0.066 |
|
|
|
| 8 |
H |
0.043 |
|
|
|
| 9 |
C |
-0.197 |
|
|
|
| 10 |
H |
0.073 |
|
|
|
| 11 |
H |
0.047 |
|
|
|
| 12 |
H |
0.048 |
|
|
|
| 13 |
C |
-0.238 |
|
|
|
| 14 |
H |
0.189 |
|
|
|
| 15 |
O |
-0.446 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.211 |
0.000 |
-1.050 |
1.603 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.823 |
0.000 |
2.798 |
| y |
0.000 |
-33.692 |
0.000 |
| z |
2.798 |
0.000 |
-34.741 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.393 |
0.000 |
2.798 |
| y |
0.000 |
-0.410 |
0.000 |
| z |
2.798 |
0.000 |
-1.983 |
|
| Polar |
|---|
| 3z2-r2 | -3.966 |
|---|
| x2-y2 | 1.868 |
|---|
| xy | 0.000 |
|---|
| xz | 2.798 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.817 |
0.000 |
0.087 |
| y |
0.000 |
7.627 |
0.000 |
| z |
0.087 |
0.000 |
7.324 |
<r2> (average value of r
2) Å
2
| <r2> |
124.553 |
| (<r2>)1/2 |
11.160 |
