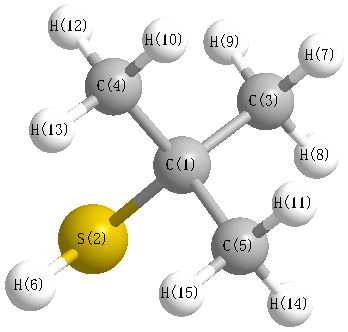
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3242 |
2938 |
39.84 |
|
|
|
| 2 |
A' |
3225 |
2923 |
54.97 |
|
|
|
| 3 |
A' |
3215 |
2914 |
40.90 |
|
|
|
| 4 |
A' |
3167 |
2871 |
36.42 |
|
|
|
| 5 |
A' |
3157 |
2861 |
40.09 |
|
|
|
| 6 |
A' |
2851 |
2584 |
6.04 |
|
|
|
| 7 |
A' |
1633 |
1480 |
6.19 |
|
|
|
| 8 |
A' |
1619 |
1467 |
8.91 |
|
|
|
| 9 |
A' |
1606 |
1455 |
0.07 |
|
|
|
| 10 |
A' |
1557 |
1412 |
0.36 |
|
|
|
| 11 |
A' |
1529 |
1386 |
6.27 |
|
|
|
| 12 |
A' |
1359 |
1232 |
0.28 |
|
|
|
| 13 |
A' |
1309 |
1186 |
42.93 |
|
|
|
| 14 |
A' |
1151 |
1044 |
1.82 |
|
|
|
| 15 |
A' |
1016 |
921 |
1.08 |
|
|
|
| 16 |
A' |
948 |
859 |
3.49 |
|
|
|
| 17 |
A' |
881 |
799 |
2.84 |
|
|
|
| 18 |
A' |
632 |
573 |
5.99 |
|
|
|
| 19 |
A' |
421 |
381 |
0.89 |
|
|
|
| 20 |
A' |
392 |
355 |
0.32 |
|
|
|
| 21 |
A' |
318 |
288 |
0.36 |
|
|
|
| 22 |
A' |
301 |
273 |
0.57 |
|
|
|
| 23 |
A" |
3240 |
2937 |
45.32 |
|
|
|
| 24 |
A" |
3232 |
2930 |
4.77 |
|
|
|
| 25 |
A" |
3209 |
2908 |
2.43 |
|
|
|
| 26 |
A" |
3152 |
2857 |
20.65 |
|
|
|
| 27 |
A" |
1620 |
1468 |
7.74 |
|
|
|
| 28 |
A" |
1603 |
1453 |
0.18 |
|
|
|
| 29 |
A" |
1596 |
1447 |
0.00 |
|
|
|
| 30 |
A" |
1529 |
1386 |
6.87 |
|
|
|
| 31 |
A" |
1342 |
1216 |
2.43 |
|
|
|
| 32 |
A" |
1136 |
1030 |
0.05 |
|
|
|
| 33 |
A" |
1050 |
951 |
0.01 |
|
|
|
| 34 |
A" |
1005 |
911 |
0.10 |
|
|
|
| 35 |
A" |
428 |
388 |
0.36 |
|
|
|
| 36 |
A" |
323 |
293 |
1.81 |
|
|
|
| 37 |
A" |
297 |
270 |
0.11 |
|
|
|
| 38 |
A" |
264 |
239 |
6.96 |
|
|
|
| 39 |
A" |
207 |
188 |
10.21 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30379.1 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 27535.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.255 |
|
|
|
| 2 |
S |
-0.192 |
|
|
|
| 3 |
C |
-0.237 |
|
|
|
| 4 |
C |
-0.234 |
|
|
|
| 5 |
C |
-0.234 |
|
|
|
| 6 |
H |
0.054 |
|
|
|
| 7 |
H |
0.056 |
|
|
|
| 8 |
H |
0.080 |
|
|
|
| 9 |
H |
0.080 |
|
|
|
| 10 |
H |
0.054 |
|
|
|
| 11 |
H |
0.054 |
|
|
|
| 12 |
H |
0.075 |
|
|
|
| 13 |
H |
0.058 |
|
|
|
| 14 |
H |
0.075 |
|
|
|
| 15 |
H |
0.058 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.634 |
-0.768 |
0.000 |
1.806 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.901 |
2.247 |
0.000 |
| y |
2.247 |
-39.570 |
0.000 |
| z |
0.000 |
0.000 |
-42.495 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.868 |
2.247 |
0.000 |
| y |
2.247 |
3.128 |
0.000 |
| z |
0.000 |
0.000 |
-1.260 |
|
| Polar |
|---|
| 3z2-r2 | -2.519 |
|---|
| x2-y2 | -3.330 |
|---|
| xy | 2.247 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.757 |
0.110 |
0.000 |
| y |
0.110 |
9.353 |
0.000 |
| z |
0.000 |
0.000 |
8.894 |
<r2> (average value of r
2) Å
2
| <r2> |
164.498 |
| (<r2>)1/2 |
12.826 |