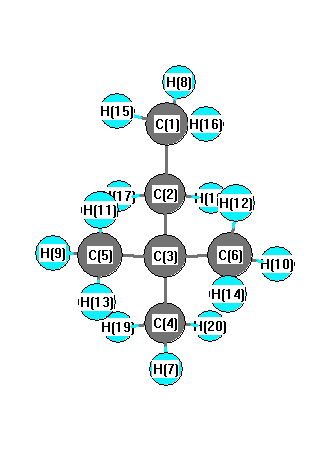
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3229 |
2926 |
91.62 |
|
|
|
| 2 |
A' |
3215 |
2914 |
174.42 |
|
|
|
| 3 |
A' |
3212 |
2912 |
3.41 |
|
|
|
| 4 |
A' |
3203 |
2903 |
2.36 |
|
|
|
| 5 |
A' |
3169 |
2872 |
7.64 |
|
|
|
| 6 |
A' |
3161 |
2865 |
37.69 |
|
|
|
| 7 |
A' |
3149 |
2854 |
45.13 |
|
|
|
| 8 |
A' |
3141 |
2847 |
25.10 |
|
|
|
| 9 |
A' |
1638 |
1485 |
10.87 |
|
|
|
| 10 |
A' |
1629 |
1476 |
6.48 |
|
|
|
| 11 |
A' |
1626 |
1474 |
4.14 |
|
|
|
| 12 |
A' |
1609 |
1458 |
0.01 |
|
|
|
| 13 |
A' |
1601 |
1452 |
0.24 |
|
|
|
| 14 |
A' |
1555 |
1410 |
4.23 |
|
|
|
| 15 |
A' |
1539 |
1395 |
3.26 |
|
|
|
| 16 |
A' |
1526 |
1383 |
5.79 |
|
|
|
| 17 |
A' |
1496 |
1356 |
0.01 |
|
|
|
| 18 |
A' |
1386 |
1256 |
3.53 |
|
|
|
| 19 |
A' |
1341 |
1215 |
6.52 |
|
|
|
| 20 |
A' |
1184 |
1073 |
1.42 |
|
|
|
| 21 |
A' |
1094 |
992 |
0.37 |
|
|
|
| 22 |
A' |
1089 |
987 |
2.50 |
|
|
|
| 23 |
A' |
1000 |
906 |
0.31 |
|
|
|
| 24 |
A' |
933 |
845 |
0.09 |
|
|
|
| 25 |
A' |
748 |
678 |
0.12 |
|
|
|
| 26 |
A' |
516 |
468 |
0.11 |
|
|
|
| 27 |
A' |
435 |
394 |
0.06 |
|
|
|
| 28 |
A' |
379 |
344 |
0.01 |
|
|
|
| 29 |
A' |
291 |
264 |
0.03 |
|
|
|
| 30 |
A' |
270 |
245 |
0.02 |
|
|
|
| 31 |
A" |
3233 |
2931 |
44.66 |
|
|
|
| 32 |
A" |
3217 |
2916 |
72.25 |
|
|
|
| 33 |
A" |
3208 |
2908 |
32.17 |
|
|
|
| 34 |
A" |
3203 |
2903 |
0.19 |
|
|
|
| 35 |
A" |
3167 |
2870 |
16.79 |
|
|
|
| 36 |
A" |
3152 |
2857 |
45.88 |
|
|
|
| 37 |
A" |
1631 |
1478 |
0.13 |
|
|
|
| 38 |
A" |
1620 |
1468 |
9.45 |
|
|
|
| 39 |
A" |
1605 |
1455 |
0.20 |
|
|
|
| 40 |
A" |
1600 |
1451 |
0.14 |
|
|
|
| 41 |
A" |
1526 |
1384 |
7.36 |
|
|
|
| 42 |
A" |
1444 |
1309 |
0.73 |
|
|
|
| 43 |
A" |
1353 |
1226 |
2.16 |
|
|
|
| 44 |
A" |
1197 |
1085 |
0.03 |
|
|
|
| 45 |
A" |
1079 |
978 |
0.43 |
|
|
|
| 46 |
A" |
1036 |
939 |
0.01 |
|
|
|
| 47 |
A" |
1001 |
907 |
0.00 |
|
|
|
| 48 |
A" |
841 |
763 |
1.50 |
|
|
|
| 49 |
A" |
447 |
405 |
0.03 |
|
|
|
| 50 |
A" |
356 |
322 |
0.00 |
|
|
|
| 51 |
A" |
307 |
278 |
0.00 |
|
|
|
| 52 |
A" |
267 |
242 |
0.01 |
|
|
|
| 53 |
A" |
215 |
195 |
0.00 |
|
|
|
| 54 |
A" |
86 |
78 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 43575.5 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 39496.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.210 |
|
|
|
| 2 |
C |
-0.112 |
|
|
|
| 3 |
C |
0.325 |
|
|
|
| 4 |
C |
-0.247 |
|
|
|
| 5 |
C |
-0.275 |
|
|
|
| 6 |
C |
-0.275 |
|
|
|
| 7 |
H |
0.057 |
|
|
|
| 8 |
H |
0.065 |
|
|
|
| 9 |
H |
0.061 |
|
|
|
| 10 |
H |
0.061 |
|
|
|
| 11 |
H |
0.060 |
|
|
|
| 12 |
H |
0.060 |
|
|
|
| 13 |
H |
0.061 |
|
|
|
| 14 |
H |
0.061 |
|
|
|
| 15 |
H |
0.052 |
|
|
|
| 16 |
H |
0.052 |
|
|
|
| 17 |
H |
0.043 |
|
|
|
| 18 |
H |
0.043 |
|
|
|
| 19 |
H |
0.058 |
|
|
|
| 20 |
H |
0.058 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.052 |
0.013 |
0.000 |
0.054 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.296 |
0.282 |
0.000 |
| y |
0.282 |
-42.802 |
0.000 |
| z |
0.000 |
0.000 |
-42.275 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.242 |
0.282 |
0.000 |
| y |
0.282 |
-0.517 |
0.000 |
| z |
0.000 |
0.000 |
0.275 |
|
| Polar |
|---|
| 3z2-r2 | 0.549 |
|---|
| x2-y2 | 0.506 |
|---|
| xy | 0.282 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.765 |
-0.430 |
0.000 |
| y |
-0.430 |
10.358 |
0.000 |
| z |
0.000 |
0.000 |
10.066 |
<r2> (average value of r
2) Å
2
| <r2> |
196.316 |
| (<r2>)1/2 |
14.011 |