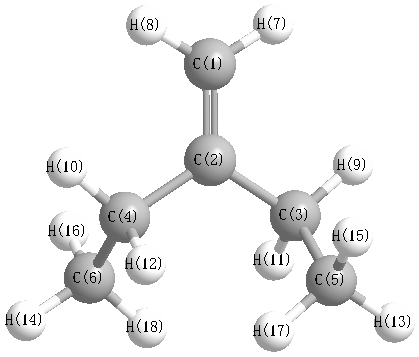
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3277 |
2970 |
13.52 |
|
|
|
| 2 |
A |
3233 |
2930 |
65.05 |
|
|
|
| 3 |
A |
3219 |
2917 |
20.64 |
|
|
|
| 4 |
A |
3199 |
2899 |
2.35 |
|
|
|
| 5 |
A |
3167 |
2871 |
22.56 |
|
|
|
| 6 |
A |
3158 |
2862 |
7.85 |
|
|
|
| 7 |
A |
1839 |
1667 |
30.44 |
|
|
|
| 8 |
A |
1634 |
1481 |
0.21 |
|
|
|
| 9 |
A |
1617 |
1465 |
3.96 |
|
|
|
| 10 |
A |
1607 |
1456 |
7.42 |
|
|
|
| 11 |
A |
1576 |
1428 |
0.95 |
|
|
|
| 12 |
A |
1533 |
1389 |
0.54 |
|
|
|
| 13 |
A |
1467 |
1329 |
7.02 |
|
|
|
| 14 |
A |
1390 |
1260 |
0.20 |
|
|
|
| 15 |
A |
1216 |
1102 |
1.61 |
|
|
|
| 16 |
A |
1160 |
1051 |
0.23 |
|
|
|
| 17 |
A |
1063 |
964 |
0.01 |
|
|
|
| 18 |
A |
868 |
787 |
2.01 |
|
|
|
| 19 |
A |
816 |
740 |
0.02 |
|
|
|
| 20 |
A |
764 |
693 |
0.06 |
|
|
|
| 21 |
A |
480 |
435 |
0.60 |
|
|
|
| 22 |
A |
284 |
257 |
0.14 |
|
|
|
| 23 |
A |
243 |
220 |
0.01 |
|
|
|
| 24 |
A |
49 |
45 |
0.01 |
|
|
|
| 25 |
B |
3351 |
3037 |
24.31 |
|
|
|
| 26 |
B |
3226 |
2924 |
80.17 |
|
|
|
| 27 |
B |
3220 |
2918 |
79.28 |
|
|
|
| 28 |
B |
3201 |
2901 |
12.38 |
|
|
|
| 29 |
B |
3164 |
2868 |
75.71 |
|
|
|
| 30 |
B |
3161 |
2865 |
11.61 |
|
|
|
| 31 |
B |
1623 |
1472 |
14.14 |
|
|
|
| 32 |
B |
1618 |
1467 |
0.65 |
|
|
|
| 33 |
B |
1607 |
1457 |
3.21 |
|
|
|
| 34 |
B |
1533 |
1390 |
4.51 |
|
|
|
| 35 |
B |
1485 |
1346 |
1.49 |
|
|
|
| 36 |
B |
1427 |
1293 |
1.47 |
|
|
|
| 37 |
B |
1369 |
1241 |
1.83 |
|
|
|
| 38 |
B |
1206 |
1093 |
3.01 |
|
|
|
| 39 |
B |
1124 |
1019 |
1.10 |
|
|
|
| 40 |
B |
1098 |
995 |
0.47 |
|
|
|
| 41 |
B |
1049 |
951 |
51.52 |
|
|
|
| 42 |
B |
963 |
873 |
2.83 |
|
|
|
| 43 |
B |
848 |
768 |
1.27 |
|
|
|
| 44 |
B |
648 |
588 |
9.84 |
|
|
|
| 45 |
B |
466 |
423 |
1.94 |
|
|
|
| 46 |
B |
304 |
276 |
0.32 |
|
|
|
| 47 |
B |
221 |
200 |
0.53 |
|
|
|
| 48 |
B |
113 |
103 |
0.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 38440.1 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 34842.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.155 |
|
|
|
| 2 |
C |
-0.032 |
|
|
|
| 3 |
C |
-0.038 |
|
|
|
| 4 |
C |
-0.038 |
|
|
|
| 5 |
C |
-0.189 |
|
|
|
| 6 |
C |
-0.189 |
|
|
|
| 7 |
H |
0.053 |
|
|
|
| 8 |
H |
0.053 |
|
|
|
| 9 |
H |
0.055 |
|
|
|
| 10 |
H |
0.055 |
|
|
|
| 11 |
H |
0.043 |
|
|
|
| 12 |
H |
0.043 |
|
|
|
| 13 |
H |
0.062 |
|
|
|
| 14 |
H |
0.062 |
|
|
|
| 15 |
H |
0.060 |
|
|
|
| 16 |
H |
0.060 |
|
|
|
| 17 |
H |
0.048 |
|
|
|
| 18 |
H |
0.048 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.557 |
0.557 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.284 |
-0.012 |
0.000 |
| y |
-0.012 |
-38.835 |
0.000 |
| z |
0.000 |
0.000 |
-40.454 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.639 |
-0.012 |
0.000 |
| y |
-0.012 |
2.534 |
0.000 |
| z |
0.000 |
0.000 |
0.106 |
|
| Polar |
|---|
| 3z2-r2 | 0.212 |
|---|
| x2-y2 | -3.449 |
|---|
| xy | -0.012 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.938 |
0.593 |
0.000 |
| y |
0.593 |
10.211 |
0.000 |
| z |
0.000 |
0.000 |
12.219 |
<r2> (average value of r
2) Å
2
| <r2> |
203.685 |
| (<r2>)1/2 |
14.272 |