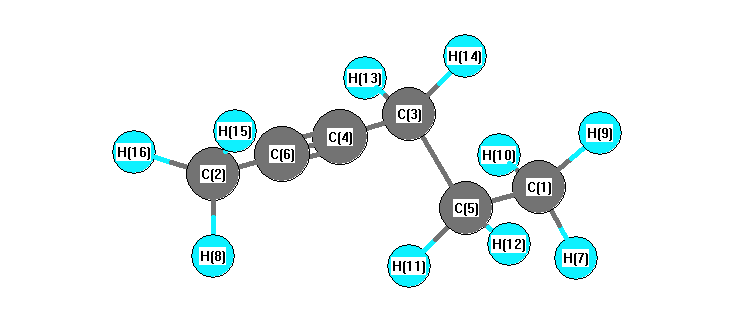
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3234 |
2932 |
16.82 |
|
|
|
| 2 |
A' |
3218 |
2917 |
57.65 |
|
|
|
| 3 |
A' |
3175 |
2878 |
38.32 |
|
|
|
| 4 |
A' |
3173 |
2876 |
46.54 |
|
|
|
| 5 |
A' |
3166 |
2870 |
12.88 |
|
|
|
| 6 |
A' |
3154 |
2858 |
30.32 |
|
|
|
| 7 |
A' |
2532 |
2295 |
0.02 |
|
|
|
| 8 |
A' |
1627 |
1475 |
5.18 |
|
|
|
| 9 |
A' |
1610 |
1459 |
0.58 |
|
|
|
| 10 |
A' |
1602 |
1452 |
2.17 |
|
|
|
| 11 |
A' |
1600 |
1450 |
7.06 |
|
|
|
| 12 |
A' |
1539 |
1395 |
1.03 |
|
|
|
| 13 |
A' |
1538 |
1394 |
4.97 |
|
|
|
| 14 |
A' |
1517 |
1375 |
2.84 |
|
|
|
| 15 |
A' |
1421 |
1288 |
13.99 |
|
|
|
| 16 |
A' |
1208 |
1095 |
2.31 |
|
|
|
| 17 |
A' |
1198 |
1086 |
0.02 |
|
|
|
| 18 |
A' |
1153 |
1045 |
0.00 |
|
|
|
| 19 |
A' |
1101 |
998 |
1.68 |
|
|
|
| 20 |
A' |
955 |
866 |
2.32 |
|
|
|
| 21 |
A' |
805 |
730 |
0.22 |
|
|
|
| 22 |
A' |
562 |
509 |
0.96 |
|
|
|
| 23 |
A' |
371 |
336 |
1.18 |
|
|
|
| 24 |
A' |
307 |
279 |
7.26 |
|
|
|
| 25 |
A' |
111 |
100 |
2.14 |
|
|
|
| 26 |
A" |
3235 |
2932 |
16.64 |
|
|
|
| 27 |
A" |
3223 |
2921 |
100.23 |
|
|
|
| 28 |
A" |
3198 |
2899 |
0.18 |
|
|
|
| 29 |
A" |
3183 |
2885 |
1.22 |
|
|
|
| 30 |
A" |
1614 |
1463 |
6.26 |
|
|
|
| 31 |
A" |
1600 |
1450 |
6.76 |
|
|
|
| 32 |
A" |
1432 |
1298 |
0.07 |
|
|
|
| 33 |
A" |
1367 |
1239 |
0.01 |
|
|
|
| 34 |
A" |
1223 |
1109 |
0.11 |
|
|
|
| 35 |
A" |
1153 |
1045 |
0.00 |
|
|
|
| 36 |
A" |
943 |
855 |
0.45 |
|
|
|
| 37 |
A" |
797 |
722 |
1.30 |
|
|
|
| 38 |
A" |
478 |
433 |
0.00 |
|
|
|
| 39 |
A" |
256 |
232 |
0.35 |
|
|
|
| 40 |
A" |
236 |
214 |
10.62 |
|
|
|
| 41 |
A" |
88 |
80 |
0.38 |
|
|
|
| 42 |
A" |
15i |
14i |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32943.1 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 29859.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.203 |
|
|
|
| 2 |
C |
-0.274 |
|
|
|
| 3 |
C |
-0.296 |
|
|
|
| 4 |
C |
0.014 |
|
|
|
| 5 |
C |
0.038 |
|
|
|
| 6 |
C |
0.062 |
|
|
|
| 7 |
H |
0.063 |
|
|
|
| 8 |
H |
0.091 |
|
|
|
| 9 |
H |
0.050 |
|
|
|
| 10 |
H |
0.050 |
|
|
|
| 11 |
H |
0.037 |
|
|
|
| 12 |
H |
0.037 |
|
|
|
| 13 |
H |
0.074 |
|
|
|
| 14 |
H |
0.074 |
|
|
|
| 15 |
H |
0.092 |
|
|
|
| 16 |
H |
0.092 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.122 |
-0.110 |
0.000 |
0.164 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.936 |
-3.038 |
0.000 |
| y |
-3.038 |
-35.430 |
0.000 |
| z |
0.000 |
0.000 |
-38.946 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.252 |
-3.038 |
0.000 |
| y |
-3.038 |
2.511 |
0.000 |
| z |
0.000 |
0.000 |
-2.763 |
|
| Polar |
|---|
| 3z2-r2 | -5.526 |
|---|
| x2-y2 | -1.506 |
|---|
| xy | -3.038 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.934 |
-2.593 |
0.000 |
| y |
-2.593 |
12.434 |
0.000 |
| z |
0.000 |
0.000 |
7.910 |
<r2> (average value of r
2) Å
2
| <r2> |
292.333 |
| (<r2>)1/2 |
17.098 |