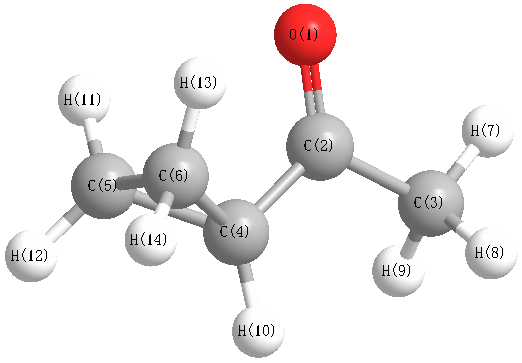
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3379 |
3063 |
11.33 |
|
|
|
| 2 |
A' |
3313 |
3003 |
11.74 |
|
|
|
| 3 |
A' |
3284 |
2977 |
12.58 |
|
|
|
| 4 |
A' |
3279 |
2972 |
20.12 |
|
|
|
| 5 |
A' |
3166 |
2869 |
5.89 |
|
|
|
| 6 |
A' |
1960 |
1777 |
212.58 |
|
|
|
| 7 |
A' |
1629 |
1477 |
11.59 |
|
|
|
| 8 |
A' |
1582 |
1434 |
13.82 |
|
|
|
| 9 |
A' |
1545 |
1401 |
88.64 |
|
|
|
| 10 |
A' |
1510 |
1369 |
5.95 |
|
|
|
| 11 |
A' |
1308 |
1185 |
44.42 |
|
|
|
| 12 |
A' |
1283 |
1163 |
55.11 |
|
|
|
| 13 |
A' |
1202 |
1090 |
13.17 |
|
|
|
| 14 |
A' |
1172 |
1062 |
8.82 |
|
|
|
| 15 |
A' |
1050 |
952 |
50.79 |
|
|
|
| 16 |
A' |
972 |
881 |
27.35 |
|
|
|
| 17 |
A' |
865 |
784 |
0.69 |
|
|
|
| 18 |
A' |
796 |
721 |
0.34 |
|
|
|
| 19 |
A' |
638 |
578 |
17.30 |
|
|
|
| 20 |
A' |
396 |
359 |
4.83 |
|
|
|
| 21 |
A' |
253 |
230 |
4.94 |
|
|
|
| 22 |
A" |
3364 |
3049 |
0.22 |
|
|
|
| 23 |
A" |
3278 |
2972 |
20.80 |
|
|
|
| 24 |
A" |
3219 |
2917 |
16.30 |
|
|
|
| 25 |
A" |
1596 |
1446 |
8.47 |
|
|
|
| 26 |
A" |
1583 |
1435 |
2.67 |
|
|
|
| 27 |
A" |
1312 |
1189 |
0.32 |
|
|
|
| 28 |
A" |
1237 |
1121 |
0.53 |
|
|
|
| 29 |
A" |
1215 |
1101 |
2.96 |
|
|
|
| 30 |
A" |
1158 |
1050 |
2.59 |
|
|
|
| 31 |
A" |
965 |
874 |
0.89 |
|
|
|
| 32 |
A" |
893 |
810 |
7.77 |
|
|
|
| 33 |
A" |
655 |
594 |
3.84 |
|
|
|
| 34 |
A" |
279 |
253 |
0.89 |
|
|
|
| 35 |
A" |
152 |
138 |
0.02 |
|
|
|
| 36 |
A" |
62 |
56 |
2.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 27774.0 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 25174.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.453 |
|
|
|
| 2 |
C |
0.320 |
|
|
|
| 3 |
C |
-0.197 |
|
|
|
| 4 |
C |
-0.028 |
|
|
|
| 5 |
C |
-0.102 |
|
|
|
| 6 |
C |
-0.102 |
|
|
|
| 7 |
H |
0.103 |
|
|
|
| 8 |
H |
0.071 |
|
|
|
| 9 |
H |
0.071 |
|
|
|
| 10 |
H |
0.056 |
|
|
|
| 11 |
H |
0.076 |
|
|
|
| 12 |
H |
0.055 |
|
|
|
| 13 |
H |
0.076 |
|
|
|
| 14 |
H |
0.055 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.081 |
-0.018 |
0.000 |
3.081 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.509 |
-1.258 |
0.000 |
| y |
-1.258 |
-34.177 |
0.000 |
| z |
0.000 |
0.000 |
-36.171 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.335 |
-1.258 |
0.000 |
| y |
-1.258 |
4.163 |
0.000 |
| z |
0.000 |
0.000 |
1.173 |
|
| Polar |
|---|
| 3z2-r2 | 2.345 |
|---|
| x2-y2 | -6.332 |
|---|
| xy | -1.258 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.033 |
0.059 |
0.000 |
| y |
0.059 |
9.494 |
0.000 |
| z |
0.000 |
0.000 |
7.294 |
<r2> (average value of r
2) Å
2
| <r2> |
167.408 |
| (<r2>)1/2 |
12.939 |