Jump to
S1C2
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -234.266123 |
| Energy at 298.15K | |
| HF Energy | -234.266123 |
| Nuclear repulsion energy | 229.825438 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3296 |
2987 |
45.73 |
|
|
|
| 2 |
A' |
3268 |
2962 |
12.86 |
|
|
|
| 3 |
A' |
3243 |
2939 |
17.92 |
|
|
|
| 4 |
A' |
3214 |
2913 |
61.42 |
|
|
|
| 5 |
A' |
3154 |
2859 |
42.87 |
|
|
|
| 6 |
A' |
3153 |
2858 |
46.13 |
|
|
|
| 7 |
A' |
3149 |
2854 |
44.26 |
|
|
|
| 8 |
A' |
3143 |
2849 |
4.45 |
|
|
|
| 9 |
A' |
1862 |
1687 |
5.31 |
|
|
|
| 10 |
A' |
1627 |
1474 |
3.72 |
|
|
|
| 11 |
A' |
1611 |
1460 |
0.46 |
|
|
|
| 12 |
A' |
1609 |
1458 |
5.08 |
|
|
|
| 13 |
A' |
1601 |
1451 |
2.36 |
|
|
|
| 14 |
A' |
1562 |
1415 |
3.02 |
|
|
|
| 15 |
A' |
1540 |
1396 |
3.00 |
|
|
|
| 16 |
A' |
1531 |
1388 |
0.24 |
|
|
|
| 17 |
A' |
1505 |
1364 |
1.22 |
|
|
|
| 18 |
A' |
1413 |
1281 |
15.93 |
|
|
|
| 19 |
A' |
1374 |
1245 |
1.03 |
|
|
|
| 20 |
A' |
1247 |
1130 |
0.29 |
|
|
|
| 21 |
A' |
1166 |
1057 |
0.62 |
|
|
|
| 22 |
A' |
1122 |
1017 |
2.43 |
|
|
|
| 23 |
A' |
1056 |
957 |
4.46 |
|
|
|
| 24 |
A' |
969 |
878 |
2.91 |
|
|
|
| 25 |
A' |
965 |
875 |
3.47 |
|
|
|
| 26 |
A' |
636 |
577 |
4.08 |
|
|
|
| 27 |
A' |
420 |
381 |
0.10 |
|
|
|
| 28 |
A' |
308 |
279 |
0.07 |
|
|
|
| 29 |
A' |
152 |
138 |
0.03 |
|
|
|
| 30 |
A" |
3211 |
2910 |
100.64 |
|
|
|
| 31 |
A" |
3192 |
2893 |
29.78 |
|
|
|
| 32 |
A" |
3181 |
2883 |
24.09 |
|
|
|
| 33 |
A" |
3157 |
2862 |
0.04 |
|
|
|
| 34 |
A" |
1614 |
1463 |
6.21 |
|
|
|
| 35 |
A" |
1604 |
1454 |
4.22 |
|
|
|
| 36 |
A" |
1436 |
1302 |
0.44 |
|
|
|
| 37 |
A" |
1367 |
1239 |
0.11 |
|
|
|
| 38 |
A" |
1216 |
1102 |
1.09 |
|
|
|
| 39 |
A" |
1168 |
1059 |
4.34 |
|
|
|
| 40 |
A" |
1106 |
1003 |
0.92 |
|
|
|
| 41 |
A" |
954 |
865 |
4.35 |
|
|
|
| 42 |
A" |
810 |
734 |
2.69 |
|
|
|
| 43 |
A" |
729 |
660 |
37.09 |
|
|
|
| 44 |
A" |
449 |
407 |
0.07 |
|
|
|
| 45 |
A" |
258 |
234 |
0.02 |
|
|
|
| 46 |
A" |
143 |
129 |
0.41 |
|
|
|
| 47 |
A" |
116 |
106 |
0.00 |
|
|
|
| 48 |
A" |
50i |
45i |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 38276.9 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 34694.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
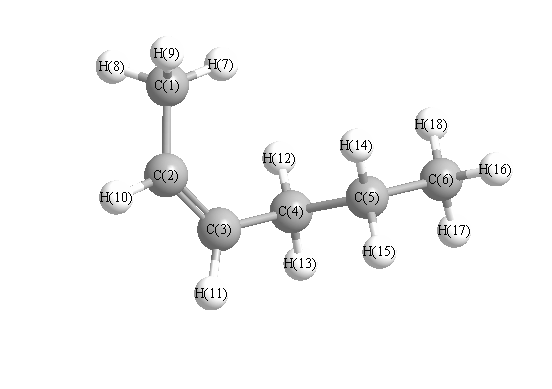
Geometric Data calculated at HF/Def2TZVPP
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.860 |
-2.062 |
0.000 |
| C2 |
0.362 |
-2.069 |
0.000 |
| C3 |
-0.427 |
-1.019 |
0.000 |
| C4 |
0.000 |
0.428 |
0.000 |
| C5 |
-1.183 |
1.391 |
0.000 |
| C6 |
-0.757 |
2.852 |
0.000 |
| H7 |
2.270 |
-1.062 |
0.000 |
| H8 |
2.246 |
-2.581 |
0.872 |
| H9 |
2.246 |
-2.581 |
-0.872 |
| H10 |
-0.101 |
-3.040 |
0.000 |
| H11 |
-1.487 |
-1.199 |
0.000 |
| H12 |
0.623 |
0.631 |
-0.867 |
| H13 |
0.623 |
0.631 |
0.867 |
| H14 |
-1.806 |
1.195 |
0.868 |
| H15 |
-1.806 |
1.195 |
-0.868 |
| H16 |
-1.617 |
3.510 |
0.000 |
| H17 |
-0.162 |
3.087 |
0.876 |
| H18 |
-0.162 |
3.087 |
-0.876 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 1.4983 | 2.5136 | 3.1080 | 4.6029 | 5.5670 | 1.0804 | 1.0854 | 1.0854 | 2.1917 | 3.4570 | 3.0872 | 3.0872 | 4.9802 | 4.9802 | 6.5677 | 5.5998 | 5.5998 |
C2 | 1.4983 | | 1.3137 | 2.5236 | 3.7900 | 5.0467 | 2.1577 | 2.1378 | 2.1378 | 1.0755 | 2.0439 | 2.8480 | 2.8480 | 4.0140 | 4.0140 | 5.9200 | 5.2559 | 5.2559 | C3 | 2.5136 | 1.3137 | | 1.5087 | 2.5261 | 3.8846 | 2.6973 | 3.2161 | 3.2161 | 2.0473 | 1.0758 | 2.1392 | 2.1392 | 2.7493 | 2.7493 | 4.6827 | 4.2061 | 4.2061 | C4 | 3.1080 | 2.5236 | 1.5087 | | 1.5258 | 2.5391 | 2.7155 | 3.8546 | 3.8546 | 3.4697 | 2.2047 | 1.0866 | 1.0866 | 2.1457 | 2.1457 | 3.4804 | 2.8036 | 2.8036 | C5 | 4.6029 | 3.7900 | 2.5261 | 1.5258 | | 1.5213 | 4.2361 | 5.3196 | 5.3196 | 4.5617 | 2.6084 | 2.1432 | 2.1432 | 1.0863 | 1.0863 | 2.1626 | 2.1644 | 2.1644 | C6 | 5.5670 | 5.0467 | 3.8846 | 2.5391 | 1.5213 | | 4.9478 | 6.2683 | 6.2683 | 5.9282 | 4.1163 | 2.7547 | 2.7547 | 2.1441 | 2.1441 | 1.0830 | 1.0847 | 1.0847 | H7 | 1.0804 | 2.1577 | 2.6973 | 2.7155 | 4.2361 | 4.9478 | | 1.7514 | 1.7514 | 3.0878 | 3.7601 | 2.5157 | 2.5157 | 4.7395 | 4.7395 | 6.0011 | 4.8877 | 4.8877 | H8 | 1.0854 | 2.1378 | 3.2161 | 3.8546 | 5.3196 | 6.2683 | 1.7514 | | 1.7431 | 2.5452 | 4.0751 | 3.9962 | 3.5984 | 5.5387 | 5.8056 | 7.2651 | 6.1576 | 6.4007 | H9 | 1.0854 | 2.1378 | 3.2161 | 3.8546 | 5.3196 | 6.2683 | 1.7514 | 1.7431 | | 2.5452 | 4.0751 | 3.5984 | 3.9962 | 5.8056 | 5.5387 | 7.2651 | 6.4007 | 6.1576 | H10 | 2.1917 | 1.0755 | 2.0473 | 3.4697 | 4.5617 | 5.9282 | 3.0878 | 2.5452 | 2.5452 | | 2.3046 | 3.8409 | 3.8409 | 4.6477 | 4.6477 | 6.7233 | 6.1892 | 6.1892 | H11 | 3.4570 | 2.0439 | 1.0758 | 2.2047 | 2.6084 | 4.1163 | 3.7601 | 4.0751 | 4.0751 | 2.3046 | | 2.9251 | 2.9251 | 2.5672 | 2.5672 | 4.7111 | 4.5709 | 4.5709 | H12 | 3.0872 | 2.8480 | 2.1392 | 1.0866 | 2.1432 | 2.7547 | 2.5157 | 3.9962 | 3.5984 | 3.8409 | 2.9251 | | 1.7332 | 3.0381 | 2.4939 | 3.7496 | 3.1115 | 2.5780 | H13 | 3.0872 | 2.8480 | 2.1392 | 1.0866 | 2.1432 | 2.7547 | 2.5157 | 3.5984 | 3.9962 | 3.8409 | 2.9251 | 1.7332 | | 2.4939 | 3.0381 | 3.7496 | 2.5780 | 3.1115 | H14 | 4.9802 | 4.0140 | 2.7493 | 2.1457 | 1.0863 | 2.1441 | 4.7395 | 5.5387 | 5.8056 | 4.6477 | 2.5672 | 3.0381 | 2.4939 | | 1.7370 | 2.4793 | 2.5060 | 3.0531 | H15 | 4.9802 | 4.0140 | 2.7493 | 2.1457 | 1.0863 | 2.1441 | 4.7395 | 5.8056 | 5.5387 | 4.6477 | 2.5672 | 2.4939 | 3.0381 | 1.7370 | | 2.4793 | 3.0531 | 2.5060 | H16 | 6.5677 | 5.9200 | 4.6827 | 3.4804 | 2.1626 | 1.0830 | 6.0011 | 7.2651 | 7.2651 | 6.7233 | 4.7111 | 3.7496 | 3.7496 | 2.4793 | 2.4793 | | 1.7506 | 1.7506 | H17 | 5.5998 | 5.2559 | 4.2061 | 2.8036 | 2.1644 | 1.0847 | 4.8877 | 6.1576 | 6.4007 | 6.1892 | 4.5709 | 3.1115 | 2.5780 | 2.5060 | 3.0531 | 1.7506 | | 1.7512 | H18 | 5.5998 | 5.2559 | 4.2061 | 2.8036 | 2.1644 | 1.0847 | 4.8877 | 6.4007 | 6.1576 | 6.1892 | 4.5709 | 2.5780 | 3.1115 | 3.0531 | 2.5060 | 1.7506 | 1.7512 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
126.612 |
|
C1 |
C2 |
H10 |
115.790 |
| C2 |
C1 |
H7 |
112.587 |
|
C2 |
C1 |
H8 |
110.660 |
| C2 |
C1 |
H9 |
110.660 |
|
C2 |
C3 |
C4 |
126.657 |
| C2 |
C3 |
H11 |
117.253 |
|
C3 |
C2 |
H10 |
117.598 |
| C3 |
C4 |
C5 |
112.708 |
|
C3 |
C4 |
H12 |
109.960 |
| C3 |
C4 |
H13 |
109.960 |
|
C4 |
C3 |
H11 |
116.090 |
| C4 |
C5 |
C6 |
112.875 |
|
C4 |
C5 |
H14 |
109.303 |
| C4 |
C5 |
H15 |
109.303 |
|
C5 |
C4 |
H12 |
109.094 |
| C5 |
C4 |
H13 |
109.094 |
|
C5 |
C6 |
H16 |
111.167 |
| C5 |
C6 |
H17 |
111.204 |
|
C5 |
C6 |
H18 |
111.204 |
| C6 |
C5 |
H14 |
109.488 |
|
C6 |
C5 |
H15 |
109.488 |
| H7 |
C1 |
H8 |
107.936 |
|
H7 |
C1 |
H9 |
107.936 |
| H8 |
C1 |
H9 |
106.836 |
|
H12 |
C4 |
H13 |
105.783 |
| H14 |
C5 |
H16 |
93.539 |
|
H16 |
C6 |
H17 |
107.721 |
| H16 |
C6 |
H18 |
107.721 |
|
H17 |
C6 |
H18 |
107.651 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.154 |
|
|
|
| 2 |
C |
-0.090 |
|
|
|
| 3 |
C |
-0.161 |
|
|
|
| 4 |
C |
-0.001 |
|
|
|
| 5 |
C |
-0.011 |
|
|
|
| 6 |
C |
-0.191 |
|
|
|
| 7 |
H |
0.052 |
|
|
|
| 8 |
H |
0.061 |
|
|
|
| 9 |
H |
0.061 |
|
|
|
| 10 |
H |
0.066 |
|
|
|
| 11 |
H |
0.061 |
|
|
|
| 12 |
H |
0.042 |
|
|
|
| 13 |
H |
0.042 |
|
|
|
| 14 |
H |
0.032 |
|
|
|
| 15 |
H |
0.032 |
|
|
|
| 16 |
H |
0.061 |
|
|
|
| 17 |
H |
0.048 |
|
|
|
| 18 |
H |
0.048 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.204 |
0.153 |
0.000 |
0.255 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.410 |
-0.777 |
0.000 |
| y |
-0.777 |
-39.910 |
0.000 |
| z |
0.000 |
0.000 |
-41.424 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.256 |
-0.777 |
0.000 |
| y |
-0.777 |
0.507 |
0.000 |
| z |
0.000 |
0.000 |
-1.764 |
|
| Polar |
|---|
| 3z2-r2 | -3.527 |
|---|
| x2-y2 | 0.499 |
|---|
| xy | -0.777 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.731 |
-1.879 |
0.000 |
| y |
-1.879 |
12.737 |
0.000 |
| z |
0.000 |
0.000 |
8.350 |
<r2> (average value of r
2) Å
2
| <r2> |
273.578 |
| (<r2>)1/2 |
16.540 |
Jump to
S1C1
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -234.267705 |
| Energy at 298.15K | -234.280326 |
| HF Energy | -234.267705 |
| Nuclear repulsion energy | 230.801671 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3289 |
2981 |
59.90 |
|
|
|
| 2 |
A |
3261 |
2956 |
19.51 |
|
|
|
| 3 |
A |
3255 |
2950 |
20.62 |
|
|
|
| 4 |
A |
3219 |
2918 |
77.04 |
|
|
|
| 5 |
A |
3214 |
2913 |
62.86 |
|
|
|
| 6 |
A |
3205 |
2905 |
20.70 |
|
|
|
| 7 |
A |
3193 |
2894 |
31.49 |
|
|
|
| 8 |
A |
3174 |
2877 |
11.30 |
|
|
|
| 9 |
A |
3156 |
2861 |
34.03 |
|
|
|
| 10 |
A |
3154 |
2859 |
23.92 |
|
|
|
| 11 |
A |
3150 |
2856 |
63.12 |
|
|
|
| 12 |
A |
3139 |
2846 |
17.47 |
|
|
|
| 13 |
A |
1869 |
1694 |
2.99 |
|
|
|
| 14 |
A |
1628 |
1475 |
6.47 |
|
|
|
| 15 |
A |
1615 |
1463 |
2.91 |
|
|
|
| 16 |
A |
1614 |
1463 |
6.24 |
|
|
|
| 17 |
A |
1609 |
1458 |
0.77 |
|
|
|
| 18 |
A |
1605 |
1455 |
3.19 |
|
|
|
| 19 |
A |
1602 |
1452 |
3.80 |
|
|
|
| 20 |
A |
1564 |
1417 |
5.58 |
|
|
|
| 21 |
A |
1538 |
1394 |
2.47 |
|
|
|
| 22 |
A |
1531 |
1388 |
1.79 |
|
|
|
| 23 |
A |
1506 |
1365 |
0.92 |
|
|
|
| 24 |
A |
1433 |
1298 |
0.08 |
|
|
|
| 25 |
A |
1414 |
1281 |
7.17 |
|
|
|
| 26 |
A |
1397 |
1266 |
0.98 |
|
|
|
| 27 |
A |
1360 |
1232 |
0.05 |
|
|
|
| 28 |
A |
1268 |
1149 |
0.13 |
|
|
|
| 29 |
A |
1205 |
1092 |
0.87 |
|
|
|
| 30 |
A |
1172 |
1062 |
3.45 |
|
|
|
| 31 |
A |
1146 |
1039 |
1.45 |
|
|
|
| 32 |
A |
1122 |
1017 |
2.54 |
|
|
|
| 33 |
A |
1110 |
1006 |
1.55 |
|
|
|
| 34 |
A |
1056 |
957 |
4.98 |
|
|
|
| 35 |
A |
968 |
877 |
3.12 |
|
|
|
| 36 |
A |
961 |
871 |
5.71 |
|
|
|
| 37 |
A |
934 |
846 |
4.69 |
|
|
|
| 38 |
A |
803 |
728 |
4.84 |
|
|
|
| 39 |
A |
782 |
708 |
30.72 |
|
|
|
| 40 |
A |
609 |
552 |
6.28 |
|
|
|
| 41 |
A |
509 |
462 |
0.84 |
|
|
|
| 42 |
A |
394 |
357 |
0.01 |
|
|
|
| 43 |
A |
305 |
276 |
0.03 |
|
|
|
| 44 |
A |
258 |
234 |
0.00 |
|
|
|
| 45 |
A |
217 |
196 |
0.02 |
|
|
|
| 46 |
A |
134 |
121 |
0.27 |
|
|
|
| 47 |
A |
97 |
88 |
0.01 |
|
|
|
| 48 |
A |
52 |
47 |
0.11 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 38394.8 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 34801.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
2.505 |
0.930 |
-0.081 |
| C2 |
1.985 |
-0.477 |
-0.131 |
| C3 |
0.772 |
-0.913 |
0.135 |
| C4 |
-0.441 |
-0.129 |
0.551 |
| C5 |
-1.591 |
-0.240 |
-0.455 |
| C6 |
-2.849 |
0.495 |
-0.006 |
| H7 |
1.755 |
1.652 |
0.199 |
| H8 |
3.322 |
1.006 |
0.630 |
| H9 |
2.903 |
1.220 |
-1.047 |
| H10 |
2.710 |
-1.216 |
-0.424 |
| H11 |
0.610 |
-1.973 |
0.035 |
| H12 |
-0.201 |
0.913 |
0.707 |
| H13 |
-0.784 |
-0.507 |
1.511 |
| H14 |
-1.261 |
0.150 |
-1.412 |
| H15 |
-1.824 |
-1.288 |
-0.618 |
| H16 |
-3.643 |
0.401 |
-0.736 |
| H17 |
-3.217 |
0.101 |
0.935 |
| H18 |
-2.655 |
1.552 |
0.135 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 1.5004 | 2.5390 | 3.1936 | 4.2765 | 5.3721 | 1.0786 | 1.0851 | 1.0849 | 2.1826 | 3.4684 | 2.8187 | 3.9267 | 4.0702 | 4.8937 | 6.2060 | 5.8706 | 5.2017 |
C2 | 1.5004 | | 1.3162 | 2.5434 | 3.5982 | 4.9316 | 2.1669 | 2.1367 | 2.1357 | 1.0761 | 2.0387 | 2.7225 | 3.2194 | 3.5452 | 3.9243 | 5.7280 | 5.3413 | 5.0705 | C3 | 2.5390 | 1.3162 | | 1.5024 | 2.5266 | 3.8870 | 2.7483 | 3.2296 | 3.2386 | 2.0396 | 1.0768 | 2.1466 | 2.1164 | 2.7668 | 2.7286 | 4.6879 | 4.1928 | 4.2210 | C4 | 3.1936 | 2.5434 | 1.5024 | | 1.5324 | 2.5492 | 2.8498 | 3.9310 | 3.9441 | 3.4723 | 2.1836 | 1.0809 | 1.0873 | 2.1461 | 2.1504 | 3.4921 | 2.8123 | 2.8112 | C5 | 4.2765 | 3.5982 | 2.5266 | 1.5324 | | 1.5242 | 3.8998 | 5.1834 | 4.7623 | 4.4103 | 2.8434 | 2.1473 | 2.1419 | 1.0849 | 1.0858 | 2.1682 | 2.1662 | 2.1660 | C6 | 5.3721 | 4.9316 | 3.8870 | 2.5492 | 1.5242 | | 4.7517 | 6.2243 | 5.8900 | 5.8307 | 4.2485 | 2.7733 | 2.7508 | 2.1487 | 2.1458 | 1.0833 | 1.0844 | 1.0844 | H7 | 1.0786 | 2.1669 | 2.7483 | 2.8498 | 3.8998 | 4.7517 | | 1.7487 | 1.7487 | 3.0868 | 3.8055 | 2.1523 | 3.5828 | 3.7354 | 4.7037 | 5.6202 | 5.2606 | 4.4114 | H8 | 1.0851 | 2.1367 | 3.2296 | 3.9310 | 5.1834 | 6.2243 | 1.7487 | | 1.7418 | 2.5343 | 4.0722 | 3.5253 | 4.4641 | 5.0899 | 5.7705 | 7.1237 | 6.6084 | 6.0218 | H9 | 1.0849 | 2.1357 | 3.2386 | 3.9441 | 4.7623 | 5.8900 | 1.7487 | 1.7418 | | 2.5219 | 4.0770 | 3.5786 | 4.8088 | 4.3149 | 5.3680 | 6.6048 | 6.5298 | 5.6916 | H10 | 2.1826 | 1.0761 | 2.0396 | 3.4723 | 4.4103 | 5.8307 | 3.0868 | 2.5343 | 2.5219 | | 2.2787 | 3.7796 | 4.0563 | 4.3139 | 4.5382 | 6.5629 | 6.2216 | 6.0623 | H11 | 3.4684 | 2.0387 | 1.0768 | 2.1836 | 2.8434 | 4.2485 | 3.8055 | 4.0722 | 4.0770 | 2.2787 | | 3.0720 | 2.5039 | 3.1776 | 2.6109 | 4.9309 | 4.4447 | 4.8053 | H12 | 2.8187 | 2.7225 | 2.1466 | 1.0809 | 2.1473 | 2.7733 | 2.1523 | 3.5253 | 3.5786 | 3.7796 | 3.0720 | | 1.7336 | 2.4889 | 3.0387 | 3.7672 | 3.1317 | 2.5989 | H13 | 3.9267 | 3.2194 | 2.1164 | 1.0873 | 2.1419 | 2.7508 | 3.5828 | 4.4641 | 4.8088 | 4.0563 | 2.5039 | 1.7336 | | 3.0338 | 2.4950 | 3.7481 | 2.5731 | 3.1041 | H14 | 4.0702 | 3.5452 | 2.7668 | 2.1461 | 1.0849 | 2.1487 | 3.7354 | 5.0899 | 4.3149 | 4.3139 | 3.1776 | 2.4889 | 3.0338 | | 1.7359 | 2.4889 | 3.0557 | 2.5102 | H15 | 4.8937 | 3.9243 | 2.7286 | 2.1504 | 1.0858 | 2.1458 | 4.7037 | 5.7705 | 5.3680 | 4.5382 | 2.6109 | 3.0387 | 2.4950 | 1.7359 | | 2.4851 | 2.5069 | 3.0538 | H16 | 6.2060 | 5.7280 | 4.6879 | 3.4921 | 2.1682 | 1.0833 | 5.6202 | 7.1237 | 6.6048 | 6.5629 | 4.9309 | 3.7672 | 3.7481 | 2.4889 | 2.4851 | | 1.7504 | 1.7500 | H17 | 5.8706 | 5.3413 | 4.1928 | 2.8123 | 2.1662 | 1.0844 | 5.2606 | 6.6084 | 6.5298 | 6.2216 | 4.4447 | 3.1317 | 2.5731 | 3.0557 | 2.5069 | 1.7504 | | 1.7503 | H18 | 5.2017 | 5.0705 | 4.2210 | 2.8112 | 2.1660 | 1.0844 | 4.4114 | 6.0218 | 5.6916 | 6.0623 | 4.8053 | 2.5989 | 3.1041 | 2.5102 | 3.0538 | 1.7500 | 1.7503 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
128.576 |
|
C1 |
C2 |
H10 |
114.800 |
| C2 |
C1 |
H7 |
113.312 |
|
C2 |
C1 |
H8 |
110.436 |
| C2 |
C1 |
H9 |
110.359 |
|
C2 |
C3 |
C4 |
128.820 |
| C2 |
C3 |
H11 |
116.492 |
|
C3 |
C2 |
H10 |
116.624 |
| C3 |
C4 |
C5 |
112.720 |
|
C3 |
C4 |
H12 |
111.350 |
| C3 |
C4 |
H13 |
108.554 |
|
C4 |
C3 |
H11 |
114.685 |
| C4 |
C5 |
C6 |
113.021 |
|
C4 |
C5 |
H14 |
108.963 |
| C4 |
C5 |
H15 |
109.243 |
|
C5 |
C4 |
H12 |
109.290 |
| C5 |
C4 |
H13 |
108.496 |
|
C5 |
C6 |
H16 |
111.387 |
| C5 |
C6 |
H17 |
111.158 |
|
C5 |
C6 |
H18 |
111.141 |
| C6 |
C5 |
H14 |
109.736 |
|
C6 |
C5 |
H15 |
109.447 |
| H7 |
C1 |
H8 |
107.848 |
|
H7 |
C1 |
H9 |
107.856 |
| H8 |
C1 |
H9 |
106.771 |
|
H12 |
C4 |
H13 |
106.178 |
| H14 |
C5 |
H16 |
93.862 |
|
H16 |
C6 |
H17 |
107.700 |
| H16 |
C6 |
H18 |
107.661 |
|
H17 |
C6 |
H18 |
107.614 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.150 |
|
|
|
| 2 |
C |
-0.094 |
|
|
|
| 3 |
C |
-0.142 |
|
|
|
| 4 |
C |
-0.019 |
|
|
|
| 5 |
C |
-0.011 |
|
|
|
| 6 |
C |
-0.188 |
|
|
|
| 7 |
H |
0.053 |
|
|
|
| 8 |
H |
0.062 |
|
|
|
| 9 |
H |
0.062 |
|
|
|
| 10 |
H |
0.064 |
|
|
|
| 11 |
H |
0.061 |
|
|
|
| 12 |
H |
0.038 |
|
|
|
| 13 |
H |
0.046 |
|
|
|
| 14 |
H |
0.035 |
|
|
|
| 15 |
H |
0.028 |
|
|
|
| 16 |
H |
0.060 |
|
|
|
| 17 |
H |
0.047 |
|
|
|
| 18 |
H |
0.047 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.069 |
0.215 |
-0.018 |
0.227 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.529 |
0.897 |
-0.595 |
| y |
0.897 |
-39.620 |
-0.005 |
| z |
-0.595 |
-0.005 |
-41.639 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.101 |
0.897 |
-0.595 |
| y |
0.897 |
0.964 |
-0.005 |
| z |
-0.595 |
-0.005 |
-2.065 |
|
| Polar |
|---|
| 3z2-r2 | -4.129 |
|---|
| x2-y2 | 0.091 |
|---|
| xy | 0.897 |
|---|
| xz | -0.595 |
|---|
| yz | -0.005 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.571 |
0.648 |
-0.498 |
| y |
0.648 |
9.641 |
-0.028 |
| z |
-0.498 |
-0.028 |
8.658 |
<r2> (average value of r
2) Å
2
| <r2> |
262.542 |
| (<r2>)1/2 |
16.203 |
