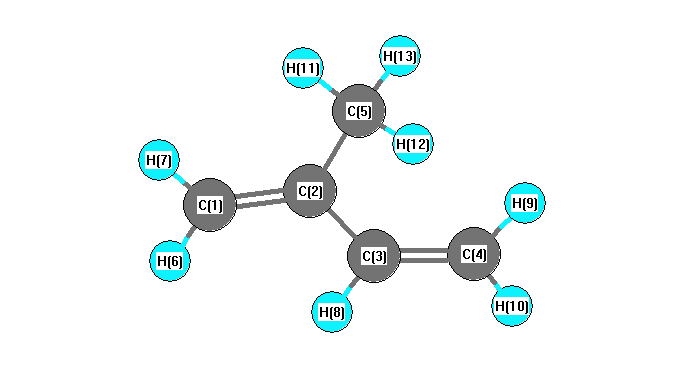
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3375 |
3059 |
17.40 |
|
|
|
| 2 |
A' |
3365 |
3050 |
14.77 |
|
|
|
| 3 |
A' |
3306 |
2996 |
6.05 |
|
|
|
| 4 |
A' |
3292 |
2984 |
14.39 |
|
|
|
| 5 |
A' |
3286 |
2978 |
3.80 |
|
|
|
| 6 |
A' |
3247 |
2943 |
25.02 |
|
|
|
| 7 |
A' |
3165 |
2868 |
26.24 |
|
|
|
| 8 |
A' |
1855 |
1681 |
1.76 |
|
|
|
| 9 |
A' |
1794 |
1626 |
42.46 |
|
|
|
| 10 |
A' |
1622 |
1470 |
6.38 |
|
|
|
| 11 |
A' |
1581 |
1433 |
1.52 |
|
|
|
| 12 |
A' |
1552 |
1406 |
2.74 |
|
|
|
| 13 |
A' |
1531 |
1388 |
2.75 |
|
|
|
| 14 |
A' |
1433 |
1299 |
0.65 |
|
|
|
| 15 |
A' |
1430 |
1296 |
0.84 |
|
|
|
| 16 |
A' |
1179 |
1069 |
4.21 |
|
|
|
| 17 |
A' |
1094 |
991 |
0.09 |
|
|
|
| 18 |
A' |
1031 |
934 |
0.48 |
|
|
|
| 19 |
A' |
829 |
751 |
0.35 |
|
|
|
| 20 |
A' |
570 |
516 |
0.01 |
|
|
|
| 21 |
A' |
459 |
416 |
2.11 |
|
|
|
| 22 |
A' |
299 |
271 |
1.40 |
|
|
|
| 23 |
A" |
3211 |
2911 |
24.61 |
|
|
|
| 24 |
A" |
1601 |
1451 |
7.55 |
|
|
|
| 25 |
A" |
1166 |
1057 |
0.00 |
|
|
|
| 26 |
A" |
1134 |
1028 |
12.99 |
|
|
|
| 27 |
A" |
1066 |
967 |
63.54 |
|
|
|
| 28 |
A" |
1053 |
954 |
42.03 |
|
|
|
| 29 |
A" |
859 |
779 |
0.63 |
|
|
|
| 30 |
A" |
707 |
641 |
0.20 |
|
|
|
| 31 |
A" |
443 |
402 |
12.01 |
|
|
|
| 32 |
A" |
231 |
209 |
0.22 |
|
|
|
| 33 |
A" |
158 |
144 |
0.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26459.7 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 23983.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.113 |
|
|
|
| 2 |
C |
0.008 |
|
|
|
| 3 |
C |
-0.109 |
|
|
|
| 4 |
C |
-0.143 |
|
|
|
| 5 |
C |
-0.165 |
|
|
|
| 6 |
H |
0.058 |
|
|
|
| 7 |
H |
0.060 |
|
|
|
| 8 |
H |
0.066 |
|
|
|
| 9 |
H |
0.058 |
|
|
|
| 10 |
H |
0.076 |
|
|
|
| 11 |
H |
0.068 |
|
|
|
| 12 |
H |
0.068 |
|
|
|
| 13 |
H |
0.068 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.259 |
-0.030 |
0.000 |
0.261 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-29.827 |
-0.005 |
0.000 |
| y |
-0.005 |
-30.309 |
0.000 |
| z |
0.000 |
0.000 |
-35.673 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.163 |
-0.005 |
0.000 |
| y |
-0.005 |
2.441 |
0.000 |
| z |
0.000 |
0.000 |
-5.604 |
|
| Polar |
|---|
| 3z2-r2 | -11.209 |
|---|
| x2-y2 | 0.482 |
|---|
| xy | -0.005 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.390 |
-1.249 |
0.000 |
| y |
-1.249 |
13.293 |
0.000 |
| z |
0.000 |
0.000 |
5.995 |
<r2> (average value of r
2) Å
2
| <r2> |
130.698 |
| (<r2>)1/2 |
11.432 |