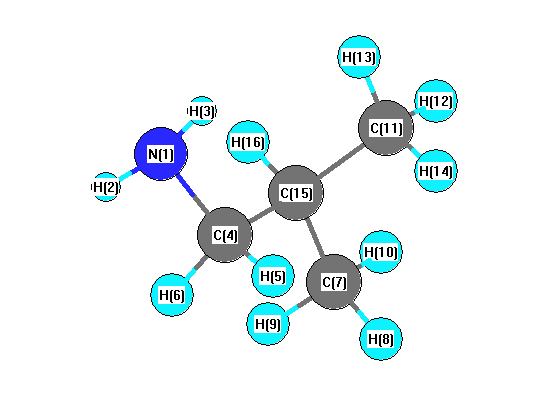
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3814 |
3457 |
2.02 |
|
|
|
| 2 |
A |
3740 |
3390 |
0.27 |
|
|
|
| 3 |
A |
3221 |
2920 |
52.22 |
|
|
|
| 4 |
A |
3214 |
2913 |
87.81 |
|
|
|
| 5 |
A |
3210 |
2909 |
88.59 |
|
|
|
| 6 |
A |
3199 |
2900 |
16.82 |
|
|
|
| 7 |
A |
3187 |
2888 |
28.67 |
|
|
|
| 8 |
A |
3158 |
2862 |
29.20 |
|
|
|
| 9 |
A |
3151 |
2856 |
21.77 |
|
|
|
| 10 |
A |
3145 |
2851 |
33.90 |
|
|
|
| 11 |
A |
3085 |
2797 |
66.65 |
|
|
|
| 12 |
A |
1796 |
1628 |
35.70 |
|
|
|
| 13 |
A |
1635 |
1482 |
9.57 |
|
|
|
| 14 |
A |
1628 |
1475 |
3.96 |
|
|
|
| 15 |
A |
1621 |
1469 |
4.54 |
|
|
|
| 16 |
A |
1611 |
1460 |
1.72 |
|
|
|
| 17 |
A |
1604 |
1454 |
0.91 |
|
|
|
| 18 |
A |
1562 |
1416 |
10.13 |
|
|
|
| 19 |
A |
1540 |
1396 |
1.89 |
|
|
|
| 20 |
A |
1529 |
1386 |
6.22 |
|
|
|
| 21 |
A |
1492 |
1352 |
1.30 |
|
|
|
| 22 |
A |
1460 |
1324 |
0.17 |
|
|
|
| 23 |
A |
1424 |
1290 |
6.92 |
|
|
|
| 24 |
A |
1327 |
1203 |
0.03 |
|
|
|
| 25 |
A |
1288 |
1167 |
1.31 |
|
|
|
| 26 |
A |
1263 |
1144 |
6.61 |
|
|
|
| 27 |
A |
1162 |
1053 |
1.57 |
|
|
|
| 28 |
A |
1156 |
1047 |
26.67 |
|
|
|
| 29 |
A |
1036 |
939 |
0.19 |
|
|
|
| 30 |
A |
1004 |
910 |
0.58 |
|
|
|
| 31 |
A |
998 |
905 |
22.93 |
|
|
|
| 32 |
A |
994 |
901 |
16.23 |
|
|
|
| 33 |
A |
901 |
817 |
101.48 |
|
|
|
| 34 |
A |
842 |
763 |
4.08 |
|
|
|
| 35 |
A |
514 |
466 |
6.19 |
|
|
|
| 36 |
A |
444 |
402 |
0.59 |
|
|
|
| 37 |
A |
387 |
351 |
0.29 |
|
|
|
| 38 |
A |
290 |
263 |
18.12 |
|
|
|
| 39 |
A |
285 |
258 |
2.22 |
|
|
|
| 40 |
A |
247 |
224 |
16.64 |
|
|
|
| 41 |
A |
228 |
207 |
12.72 |
|
|
|
| 42 |
A |
128 |
116 |
3.77 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 34758.7 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 31505.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.401 |
|
|
|
| 2 |
H |
0.135 |
|
|
|
| 3 |
H |
0.135 |
|
|
|
| 4 |
C |
0.009 |
|
|
|
| 5 |
H |
0.038 |
|
|
|
| 6 |
H |
0.044 |
|
|
|
| 7 |
C |
-0.212 |
|
|
|
| 8 |
H |
0.047 |
|
|
|
| 9 |
H |
0.057 |
|
|
|
| 10 |
H |
0.058 |
|
|
|
| 11 |
C |
-0.230 |
|
|
|
| 12 |
H |
0.059 |
|
|
|
| 13 |
H |
0.061 |
|
|
|
| 14 |
H |
0.049 |
|
|
|
| 15 |
C |
0.153 |
|
|
|
| 16 |
H |
-0.002 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.170 |
-0.169 |
1.106 |
1.132 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.957 |
0.652 |
-3.141 |
| y |
0.652 |
-33.400 |
0.489 |
| z |
-3.141 |
0.489 |
-35.706 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.597 |
0.652 |
-3.141 |
| y |
0.652 |
0.931 |
0.489 |
| z |
-3.141 |
0.489 |
-2.528 |
|
| Polar |
|---|
| 3z2-r2 | -5.056 |
|---|
| x2-y2 | 0.444 |
|---|
| xy | 0.652 |
|---|
| xz | -3.141 |
|---|
| yz | 0.489 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.935 |
-0.068 |
0.020 |
| y |
-0.068 |
8.309 |
-0.022 |
| z |
0.020 |
-0.022 |
7.337 |
<r2> (average value of r
2) Å
2
| <r2> |
149.977 |
| (<r2>)1/2 |
12.247 |